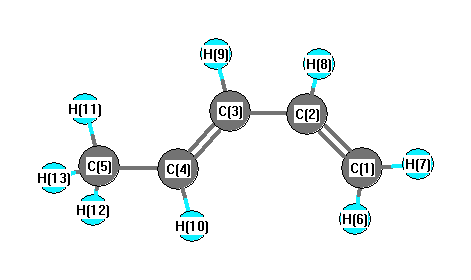
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3239 |
3126 |
17.30 |
|
|
|
| 2 |
A' |
3177 |
3066 |
25.66 |
|
|
|
| 3 |
A' |
3165 |
3054 |
16.50 |
|
|
|
| 4 |
A' |
3155 |
3044 |
6.76 |
|
|
|
| 5 |
A' |
3144 |
3034 |
3.82 |
|
|
|
| 6 |
A' |
3130 |
3020 |
3.66 |
|
|
|
| 7 |
A' |
3037 |
2930 |
22.71 |
|
|
|
| 8 |
A' |
1714 |
1654 |
6.83 |
|
|
|
| 9 |
A' |
1672 |
1614 |
1.91 |
|
|
|
| 10 |
A' |
1555 |
1500 |
9.47 |
|
|
|
| 11 |
A' |
1517 |
1463 |
8.59 |
|
|
|
| 12 |
A' |
1466 |
1414 |
8.27 |
|
|
|
| 13 |
A' |
1449 |
1398 |
0.63 |
|
|
|
| 14 |
A' |
1371 |
1323 |
0.38 |
|
|
|
| 15 |
A' |
1332 |
1285 |
1.35 |
|
|
|
| 16 |
A' |
1211 |
1169 |
3.72 |
|
|
|
| 17 |
A' |
1097 |
1058 |
0.54 |
|
|
|
| 18 |
A' |
975 |
940 |
11.28 |
|
|
|
| 19 |
A' |
889 |
858 |
2.93 |
|
|
|
| 20 |
A' |
636 |
613 |
5.75 |
|
|
|
| 21 |
A' |
404 |
390 |
3.87 |
|
|
|
| 22 |
A' |
229 |
221 |
0.19 |
|
|
|
| 23 |
A" |
3074 |
2966 |
19.42 |
|
|
|
| 24 |
A" |
1553 |
1498 |
8.72 |
|
|
|
| 25 |
A" |
1099 |
1060 |
0.01 |
|
|
|
| 26 |
A" |
1056 |
1019 |
14.47 |
|
|
|
| 27 |
A" |
1013 |
978 |
0.82 |
|
|
|
| 28 |
A" |
955 |
921 |
62.56 |
|
|
|
| 29 |
A" |
814 |
785 |
8.76 |
|
|
|
| 30 |
A" |
650 |
627 |
35.40 |
|
|
|
| 31 |
A" |
359 |
347 |
0.19 |
|
|
|
| 32 |
A" |
149 |
144 |
0.01 |
|
|
|
| 33 |
A" |
118 |
114 |
0.70 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25200.9 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 24316.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.372 |
|
|
|
| 2 |
C |
-0.198 |
|
|
|
| 3 |
C |
-0.186 |
|
|
|
| 4 |
C |
-0.163 |
|
|
|
| 5 |
C |
-0.610 |
|
|
|
| 6 |
H |
0.182 |
|
|
|
| 7 |
H |
0.186 |
|
|
|
| 8 |
H |
0.186 |
|
|
|
| 9 |
H |
0.180 |
|
|
|
| 10 |
H |
0.184 |
|
|
|
| 11 |
H |
0.201 |
|
|
|
| 12 |
H |
0.206 |
|
|
|
| 13 |
H |
0.206 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.465 |
-0.205 |
0.000 |
0.508 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.660 |
0.225 |
0.000 |
| y |
0.225 |
-29.319 |
0.000 |
| z |
0.000 |
0.000 |
-34.961 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.480 |
0.225 |
0.000 |
| y |
0.225 |
2.492 |
0.000 |
| z |
0.000 |
0.000 |
-5.972 |
|
| Polar |
|---|
| 3z2-r2 | -11.943 |
|---|
| x2-y2 | 0.658 |
|---|
| xy | 0.225 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.584 |
0.818 |
0.000 |
| y |
0.818 |
6.755 |
0.000 |
| z |
0.000 |
0.000 |
3.160 |
<r2> (average value of r
2) Å
2
| <r2> |
156.154 |
| (<r2>)1/2 |
12.496 |