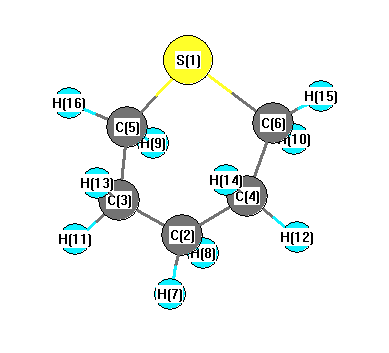
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3147 |
3037 |
15.86 |
|
|
|
| 2 |
A |
3101 |
2992 |
56.48 |
|
|
|
| 3 |
A |
3084 |
2976 |
9.56 |
|
|
|
| 4 |
A |
3077 |
2969 |
23.83 |
|
|
|
| 5 |
A |
3042 |
2935 |
20.17 |
|
|
|
| 6 |
A |
3032 |
2926 |
10.13 |
|
|
|
| 7 |
A |
1551 |
1497 |
7.97 |
|
|
|
| 8 |
A |
1541 |
1487 |
10.70 |
|
|
|
| 9 |
A |
1525 |
1472 |
12.47 |
|
|
|
| 10 |
A |
1410 |
1361 |
0.24 |
|
|
|
| 11 |
A |
1371 |
1323 |
3.96 |
|
|
|
| 12 |
A |
1285 |
1240 |
4.09 |
|
|
|
| 13 |
A |
1266 |
1222 |
2.53 |
|
|
|
| 14 |
A |
1090 |
1052 |
1.20 |
|
|
|
| 15 |
A |
1031 |
995 |
1.74 |
|
|
|
| 16 |
A |
990 |
955 |
10.85 |
|
|
|
| 17 |
A |
854 |
825 |
5.58 |
|
|
|
| 18 |
A |
802 |
774 |
2.65 |
|
|
|
| 19 |
A |
603 |
582 |
5.84 |
|
|
|
| 20 |
A |
506 |
488 |
1.32 |
|
|
|
| 21 |
A |
359 |
346 |
0.59 |
|
|
|
| 22 |
A |
332 |
321 |
0.26 |
|
|
|
| 23 |
A |
168 |
162 |
2.19 |
|
|
|
| 24 |
A |
3147 |
3037 |
6.36 |
|
|
|
| 25 |
A |
3095 |
2986 |
18.47 |
|
|
|
| 26 |
A |
3080 |
2972 |
6.53 |
|
|
|
| 27 |
A |
3038 |
2931 |
38.75 |
|
|
|
| 28 |
A |
1532 |
1479 |
3.77 |
|
|
|
| 29 |
A |
1522 |
1469 |
2.66 |
|
|
|
| 30 |
A |
1418 |
1368 |
0.79 |
|
|
|
| 31 |
A |
1406 |
1357 |
0.06 |
|
|
|
| 32 |
A |
1328 |
1281 |
7.88 |
|
|
|
| 33 |
A |
1323 |
1277 |
14.41 |
|
|
|
| 34 |
A |
1198 |
1156 |
0.04 |
|
|
|
| 35 |
A |
1144 |
1104 |
2.30 |
|
|
|
| 36 |
A |
1079 |
1041 |
0.09 |
|
|
|
| 37 |
A |
942 |
909 |
0.03 |
|
|
|
| 38 |
A |
915 |
883 |
9.20 |
|
|
|
| 39 |
A |
810 |
781 |
0.14 |
|
|
|
| 40 |
A |
634 |
612 |
2.86 |
|
|
|
| 41 |
A |
402 |
387 |
0.13 |
|
|
|
| 42 |
A |
237 |
228 |
0.70 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31709.6 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 30596.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.221 |
|
|
-0.254 |
| 2 |
C |
-0.363 |
|
|
-0.071 |
| 3 |
C |
-0.370 |
|
|
-0.009 |
| 4 |
C |
-0.370 |
|
|
-0.009 |
| 5 |
C |
-0.598 |
|
|
-0.245 |
| 6 |
C |
-0.598 |
|
|
-0.245 |
| 7 |
H |
0.194 |
|
|
0.028 |
| 8 |
H |
0.184 |
|
|
0.064 |
| 9 |
H |
0.219 |
|
|
0.128 |
| 10 |
H |
0.219 |
|
|
0.128 |
| 11 |
H |
0.197 |
|
|
0.021 |
| 12 |
H |
0.197 |
|
|
0.021 |
| 13 |
H |
0.206 |
|
|
0.060 |
| 14 |
H |
0.206 |
|
|
0.060 |
| 15 |
H |
0.229 |
|
|
0.162 |
| 16 |
H |
0.229 |
|
|
0.162 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.443 |
-2.180 |
0.000 |
2.225 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.510 |
-2.158 |
0.000 |
2.218 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.400 |
-0.130 |
0.000 |
| y |
-0.130 |
-50.063 |
0.000 |
| z |
0.000 |
0.000 |
-42.118 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.309 |
-0.130 |
0.000 |
| y |
-0.130 |
-5.804 |
0.000 |
| z |
0.000 |
0.000 |
6.113 |
|
| Polar |
|---|
| 3z2-r2 | 12.226 |
|---|
| x2-y2 | 3.664 |
|---|
| xy | -0.130 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.820 |
0.031 |
-0.533 |
| y |
0.031 |
10.432 |
-0.128 |
| z |
-0.533 |
-0.128 |
7.676 |
<r2> (average value of r
2) Å
2
| <r2> |
202.565 |
| (<r2>)1/2 |
14.233 |