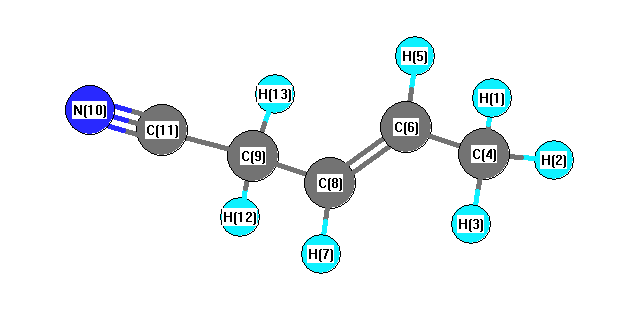
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3172 |
3061 |
16.05 |
|
|
|
| 2 |
A |
3145 |
3034 |
17.51 |
|
|
|
| 3 |
A |
3118 |
3009 |
5.56 |
|
|
|
| 4 |
A |
3089 |
2981 |
1.62 |
|
|
|
| 5 |
A |
3084 |
2976 |
16.76 |
|
|
|
| 6 |
A |
3043 |
2936 |
5.93 |
|
|
|
| 7 |
A |
3040 |
2934 |
18.41 |
|
|
|
| 8 |
A |
2349 |
2267 |
3.11 |
|
|
|
| 9 |
A |
1743 |
1682 |
0.88 |
|
|
|
| 10 |
A |
1556 |
1501 |
11.46 |
|
|
|
| 11 |
A |
1550 |
1495 |
9.64 |
|
|
|
| 12 |
A |
1518 |
1465 |
14.58 |
|
|
|
| 13 |
A |
1466 |
1414 |
4.80 |
|
|
|
| 14 |
A |
1395 |
1346 |
1.98 |
|
|
|
| 15 |
A |
1371 |
1323 |
1.10 |
|
|
|
| 16 |
A |
1351 |
1303 |
1.42 |
|
|
|
| 17 |
A |
1286 |
1240 |
0.25 |
|
|
|
| 18 |
A |
1155 |
1114 |
2.02 |
|
|
|
| 19 |
A |
1109 |
1070 |
0.53 |
|
|
|
| 20 |
A |
1053 |
1016 |
1.05 |
|
|
|
| 21 |
A |
1014 |
979 |
41.70 |
|
|
|
| 22 |
A |
990 |
955 |
11.60 |
|
|
|
| 23 |
A |
946 |
913 |
3.08 |
|
|
|
| 24 |
A |
905 |
873 |
2.40 |
|
|
|
| 25 |
A |
787 |
760 |
0.66 |
|
|
|
| 26 |
A |
614 |
593 |
1.74 |
|
|
|
| 27 |
A |
463 |
447 |
0.49 |
|
|
|
| 28 |
A |
447 |
431 |
0.04 |
|
|
|
| 29 |
A |
316 |
305 |
1.27 |
|
|
|
| 30 |
A |
271 |
262 |
4.12 |
|
|
|
| 31 |
A |
204 |
197 |
1.28 |
|
|
|
| 32 |
A |
146 |
141 |
2.63 |
|
|
|
| 33 |
A |
69 |
67 |
1.61 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23882.5 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 23044.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.207 |
|
|
|
| 2 |
H |
0.211 |
|
|
|
| 3 |
H |
0.202 |
|
|
|
| 4 |
C |
-0.602 |
|
|
|
| 5 |
H |
0.184 |
|
|
|
| 6 |
C |
-0.153 |
|
|
|
| 7 |
H |
0.197 |
|
|
|
| 8 |
C |
-0.131 |
|
|
|
| 9 |
C |
-0.509 |
|
|
|
| 10 |
N |
-0.477 |
|
|
|
| 11 |
C |
0.361 |
|
|
|
| 12 |
H |
0.252 |
|
|
|
| 13 |
H |
0.259 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.308 |
2.125 |
0.217 |
3.938 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.562 |
-6.262 |
-1.553 |
| y |
-6.262 |
-37.121 |
-1.257 |
| z |
-1.553 |
-1.257 |
-34.728 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.637 |
-6.262 |
-1.553 |
| y |
-6.262 |
2.524 |
-1.257 |
| z |
-1.553 |
-1.257 |
6.113 |
|
| Polar |
|---|
| 3z2-r2 | 12.227 |
|---|
| x2-y2 | -7.441 |
|---|
| xy | -6.262 |
|---|
| xz | -1.553 |
|---|
| yz | -1.257 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.668 |
0.323 |
-0.897 |
| y |
0.323 |
5.941 |
-0.365 |
| z |
-0.897 |
-0.365 |
5.981 |
<r2> (average value of r
2) Å
2
| <r2> |
234.258 |
| (<r2>)1/2 |
15.305 |