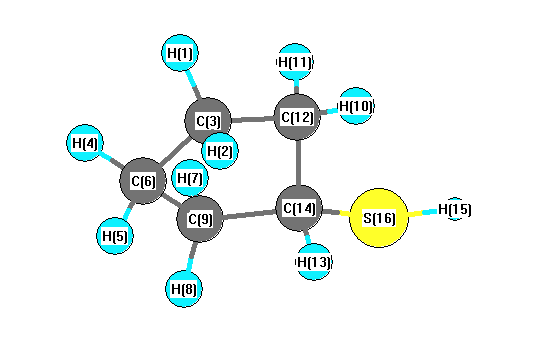
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3146 |
3036 |
52.55 |
|
|
|
| 2 |
A |
3132 |
3022 |
15.01 |
|
|
|
| 3 |
A |
3129 |
3019 |
3.99 |
|
|
|
| 4 |
A |
3110 |
3001 |
12.71 |
|
|
|
| 5 |
A |
3106 |
2997 |
3.93 |
|
|
|
| 6 |
A |
3089 |
2980 |
33.02 |
|
|
|
| 7 |
A |
3078 |
2970 |
19.73 |
|
|
|
| 8 |
A |
3075 |
2967 |
14.13 |
|
|
|
| 9 |
A |
3072 |
2965 |
12.90 |
|
|
|
| 10 |
A |
2458 |
2371 |
38.11 |
|
|
|
| 11 |
A |
1571 |
1516 |
1.31 |
|
|
|
| 12 |
A |
1552 |
1498 |
9.09 |
|
|
|
| 13 |
A |
1548 |
1493 |
4.87 |
|
|
|
| 14 |
A |
1540 |
1486 |
2.00 |
|
|
|
| 15 |
A |
1382 |
1334 |
7.62 |
|
|
|
| 16 |
A |
1375 |
1327 |
2.21 |
|
|
|
| 17 |
A |
1362 |
1315 |
0.49 |
|
|
|
| 18 |
A |
1355 |
1307 |
0.79 |
|
|
|
| 19 |
A |
1338 |
1291 |
0.72 |
|
|
|
| 20 |
A |
1302 |
1257 |
14.32 |
|
|
|
| 21 |
A |
1294 |
1248 |
3.33 |
|
|
|
| 22 |
A |
1247 |
1203 |
10.37 |
|
|
|
| 23 |
A |
1226 |
1183 |
1.46 |
|
|
|
| 24 |
A |
1195 |
1153 |
6.05 |
|
|
|
| 25 |
A |
1100 |
1062 |
0.37 |
|
|
|
| 26 |
A |
1044 |
1008 |
3.32 |
|
|
|
| 27 |
A |
1017 |
981 |
1.46 |
|
|
|
| 28 |
A |
982 |
948 |
0.35 |
|
|
|
| 29 |
A |
946 |
913 |
1.01 |
|
|
|
| 30 |
A |
926 |
894 |
2.49 |
|
|
|
| 31 |
A |
876 |
845 |
3.36 |
|
|
|
| 32 |
A |
852 |
822 |
5.64 |
|
|
|
| 33 |
A |
817 |
788 |
1.72 |
|
|
|
| 34 |
A |
810 |
782 |
8.59 |
|
|
|
| 35 |
A |
740 |
714 |
5.26 |
|
|
|
| 36 |
A |
639 |
617 |
0.80 |
|
|
|
| 37 |
A |
463 |
447 |
0.19 |
|
|
|
| 38 |
A |
351 |
338 |
3.01 |
|
|
|
| 39 |
A |
243 |
235 |
3.46 |
|
|
|
| 40 |
A |
201 |
194 |
27.03 |
|
|
|
| 41 |
A |
159 |
153 |
1.05 |
|
|
|
| 42 |
A |
40 |
39 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30942.7 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 29856.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.196 |
|
|
0.037 |
| 2 |
H |
0.195 |
|
|
0.030 |
| 3 |
C |
-0.378 |
|
|
-0.016 |
| 4 |
H |
0.198 |
|
|
0.069 |
| 5 |
H |
0.197 |
|
|
0.076 |
| 6 |
C |
-0.376 |
|
|
-0.198 |
| 7 |
H |
0.212 |
|
|
0.110 |
| 8 |
H |
0.204 |
|
|
0.059 |
| 9 |
C |
-0.359 |
|
|
-0.070 |
| 10 |
H |
0.190 |
|
|
0.045 |
| 11 |
H |
0.213 |
|
|
0.058 |
| 12 |
C |
-0.355 |
|
|
-0.091 |
| 13 |
H |
0.231 |
|
|
0.110 |
| 14 |
C |
-0.456 |
|
|
-0.029 |
| 15 |
H |
0.069 |
|
|
0.213 |
| 16 |
S |
0.020 |
|
|
-0.404 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.797 |
0.931 |
0.831 |
2.188 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-1.840 |
0.904 |
0.886 |
2.234 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.128 |
3.141 |
1.715 |
| y |
3.141 |
-45.117 |
0.958 |
| z |
1.715 |
0.958 |
-45.957 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.591 |
3.141 |
1.715 |
| y |
3.141 |
2.426 |
0.958 |
| z |
1.715 |
0.958 |
1.165 |
|
| Polar |
|---|
| 3z2-r2 | 2.331 |
|---|
| x2-y2 | -4.011 |
|---|
| xy | 3.141 |
|---|
| xz | 1.715 |
|---|
| yz | 0.958 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.564 |
0.374 |
-0.108 |
| y |
0.374 |
8.816 |
0.446 |
| z |
-0.108 |
0.446 |
7.502 |
<r2> (average value of r
2) Å
2
| <r2> |
236.742 |
| (<r2>)1/2 |
15.386 |