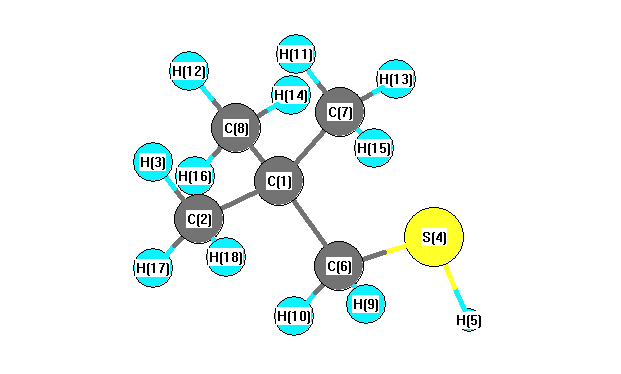
Vibrational Frequencies calculated at BLYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3050 |
3033 |
40.34 |
|
|
|
| 2 |
A' |
3035 |
3019 |
66.57 |
|
|
|
| 3 |
A' |
3028 |
3011 |
0.19 |
|
|
|
| 4 |
A' |
3009 |
2992 |
17.13 |
|
|
|
| 5 |
A' |
2976 |
2960 |
10.51 |
|
|
|
| 6 |
A' |
2965 |
2948 |
21.25 |
|
|
|
| 7 |
A' |
2384 |
2371 |
47.49 |
|
|
|
| 8 |
A' |
1547 |
1538 |
11.70 |
|
|
|
| 9 |
A' |
1536 |
1528 |
12.29 |
|
|
|
| 10 |
A' |
1517 |
1508 |
0.13 |
|
|
|
| 11 |
A' |
1490 |
1482 |
3.78 |
|
|
|
| 12 |
A' |
1440 |
1432 |
7.00 |
|
|
|
| 13 |
A' |
1412 |
1404 |
9.38 |
|
|
|
| 14 |
A' |
1276 |
1269 |
26.23 |
|
|
|
| 15 |
A' |
1251 |
1244 |
12.89 |
|
|
|
| 16 |
A' |
1196 |
1190 |
15.57 |
|
|
|
| 17 |
A' |
1049 |
1043 |
0.37 |
|
|
|
| 18 |
A' |
934 |
928 |
0.79 |
|
|
|
| 19 |
A' |
895 |
890 |
3.87 |
|
|
|
| 20 |
A' |
819 |
815 |
9.75 |
|
|
|
| 21 |
A' |
700 |
697 |
8.34 |
|
|
|
| 22 |
A' |
644 |
640 |
2.81 |
|
|
|
| 23 |
A' |
438 |
435 |
2.40 |
|
|
|
| 24 |
A' |
380 |
378 |
0.05 |
|
|
|
| 25 |
A' |
304 |
303 |
1.14 |
|
|
|
| 26 |
A' |
258 |
257 |
0.08 |
|
|
|
| 27 |
A' |
190 |
189 |
2.50 |
|
|
|
| 28 |
A" |
3070 |
3054 |
21.02 |
|
|
|
| 29 |
A" |
3045 |
3028 |
2.72 |
|
|
|
| 30 |
A" |
3030 |
3014 |
36.33 |
|
|
|
| 31 |
A" |
3026 |
3009 |
0.79 |
|
|
|
| 32 |
A" |
2970 |
2954 |
29.53 |
|
|
|
| 33 |
A" |
1535 |
1526 |
6.51 |
|
|
|
| 34 |
A" |
1517 |
1509 |
0.33 |
|
|
|
| 35 |
A" |
1511 |
1502 |
0.10 |
|
|
|
| 36 |
A" |
1416 |
1408 |
10.95 |
|
|
|
| 37 |
A" |
1283 |
1276 |
3.27 |
|
|
|
| 38 |
A" |
1136 |
1129 |
2.97 |
|
|
|
| 39 |
A" |
1044 |
1038 |
1.05 |
|
|
|
| 40 |
A" |
979 |
974 |
0.00 |
|
|
|
| 41 |
A" |
925 |
919 |
0.01 |
|
|
|
| 42 |
A" |
854 |
849 |
2.34 |
|
|
|
| 43 |
A" |
404 |
402 |
0.10 |
|
|
|
| 44 |
A" |
335 |
333 |
0.16 |
|
|
|
| 45 |
A" |
273 |
272 |
0.04 |
|
|
|
| 46 |
A" |
225 |
223 |
0.01 |
|
|
|
| 47 |
A" |
140 |
139 |
6.36 |
|
|
|
| 48 |
A" |
25 |
25 |
19.47 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34231.8 cm
-1
Scaled (by 0.9945) Zero Point Vibrational Energy (zpe) 34043.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.125 |
|
|
|
| 2 |
C |
-0.470 |
|
|
|
| 3 |
H |
0.179 |
|
|
|
| 4 |
S |
-0.025 |
|
|
|
| 5 |
H |
0.064 |
|
|
|
| 6 |
C |
-0.499 |
|
|
|
| 7 |
C |
-0.482 |
|
|
|
| 8 |
C |
-0.482 |
|
|
|
| 9 |
H |
0.207 |
|
|
|
| 10 |
H |
0.207 |
|
|
|
| 11 |
H |
0.171 |
|
|
|
| 12 |
H |
0.171 |
|
|
|
| 13 |
H |
0.199 |
|
|
|
| 14 |
H |
0.199 |
|
|
|
| 15 |
H |
0.170 |
|
|
|
| 16 |
H |
0.170 |
|
|
|
| 17 |
H |
0.172 |
|
|
|
| 18 |
H |
0.172 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.515 |
1.223 |
0.000 |
1.947 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.324 |
3.487 |
0.000 |
| y |
3.487 |
-48.304 |
0.000 |
| z |
0.000 |
0.000 |
-49.028 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.342 |
3.487 |
0.000 |
| y |
3.487 |
-0.628 |
0.000 |
| z |
0.000 |
0.000 |
-1.714 |
|
| Polar |
|---|
| 3z2-r2 | -3.427 |
|---|
| x2-y2 | 1.980 |
|---|
| xy | 3.487 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
253.062 |
| (<r2>)1/2 |
15.908 |