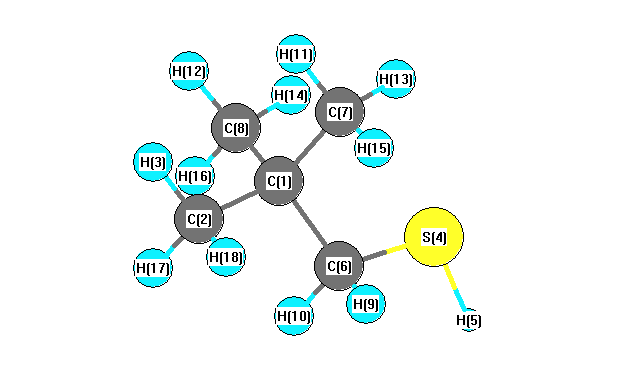
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3129 |
3020 |
35.83 |
|
|
|
| 2 |
A' |
3115 |
3005 |
60.49 |
|
|
|
| 3 |
A' |
3108 |
2999 |
0.49 |
|
|
|
| 4 |
A' |
3089 |
2981 |
15.80 |
|
|
|
| 5 |
A' |
3052 |
2945 |
9.11 |
|
|
|
| 6 |
A' |
3041 |
2934 |
19.22 |
|
|
|
| 7 |
A' |
2469 |
2383 |
41.70 |
|
|
|
| 8 |
A' |
1582 |
1527 |
14.11 |
|
|
|
| 9 |
A' |
1572 |
1517 |
14.44 |
|
|
|
| 10 |
A' |
1551 |
1497 |
0.05 |
|
|
|
| 11 |
A' |
1527 |
1473 |
4.88 |
|
|
|
| 12 |
A' |
1479 |
1427 |
9.48 |
|
|
|
| 13 |
A' |
1451 |
1400 |
13.36 |
|
|
|
| 14 |
A' |
1323 |
1277 |
21.63 |
|
|
|
| 15 |
A' |
1296 |
1250 |
14.79 |
|
|
|
| 16 |
A' |
1240 |
1196 |
15.73 |
|
|
|
| 17 |
A' |
1082 |
1044 |
0.36 |
|
|
|
| 18 |
A' |
969 |
935 |
0.62 |
|
|
|
| 19 |
A' |
934 |
901 |
3.80 |
|
|
|
| 20 |
A' |
853 |
823 |
10.95 |
|
|
|
| 21 |
A' |
734 |
709 |
8.21 |
|
|
|
| 22 |
A' |
673 |
649 |
1.99 |
|
|
|
| 23 |
A' |
454 |
438 |
2.14 |
|
|
|
| 24 |
A' |
392 |
378 |
0.06 |
|
|
|
| 25 |
A' |
317 |
306 |
1.06 |
|
|
|
| 26 |
A' |
268 |
258 |
0.07 |
|
|
|
| 27 |
A' |
200 |
193 |
2.45 |
|
|
|
| 28 |
A" |
3152 |
3041 |
16.75 |
|
|
|
| 29 |
A" |
3124 |
3015 |
2.79 |
|
|
|
| 30 |
A" |
3110 |
3000 |
33.54 |
|
|
|
| 31 |
A" |
3106 |
2997 |
0.94 |
|
|
|
| 32 |
A" |
3046 |
2939 |
26.85 |
|
|
|
| 33 |
A" |
1571 |
1516 |
7.92 |
|
|
|
| 34 |
A" |
1552 |
1497 |
0.32 |
|
|
|
| 35 |
A" |
1546 |
1492 |
0.13 |
|
|
|
| 36 |
A" |
1455 |
1404 |
15.36 |
|
|
|
| 37 |
A" |
1328 |
1282 |
3.11 |
|
|
|
| 38 |
A" |
1173 |
1132 |
2.92 |
|
|
|
| 39 |
A" |
1077 |
1039 |
0.90 |
|
|
|
| 40 |
A" |
1007 |
971 |
0.00 |
|
|
|
| 41 |
A" |
958 |
924 |
0.10 |
|
|
|
| 42 |
A" |
884 |
853 |
2.41 |
|
|
|
| 43 |
A" |
416 |
401 |
0.10 |
|
|
|
| 44 |
A" |
345 |
333 |
0.17 |
|
|
|
| 45 |
A" |
283 |
274 |
0.03 |
|
|
|
| 46 |
A" |
235 |
226 |
0.02 |
|
|
|
| 47 |
A" |
138 |
133 |
4.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35199.9 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 33964.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.168 |
|
|
0.655 |
| 2 |
C |
-0.495 |
|
|
-0.552 |
| 3 |
H |
0.191 |
|
|
0.143 |
| 4 |
S |
-0.012 |
|
|
-0.373 |
| 5 |
H |
0.069 |
|
|
0.237 |
| 6 |
C |
-0.539 |
|
|
-0.569 |
| 7 |
C |
-0.507 |
|
|
-0.517 |
| 8 |
C |
-0.507 |
|
|
-0.517 |
| 9 |
H |
0.222 |
|
|
0.213 |
| 10 |
H |
0.222 |
|
|
0.213 |
| 11 |
H |
0.184 |
|
|
0.122 |
| 12 |
H |
0.184 |
|
|
0.122 |
| 13 |
H |
0.212 |
|
|
0.160 |
| 14 |
H |
0.212 |
|
|
0.160 |
| 15 |
H |
0.182 |
|
|
0.123 |
| 16 |
H |
0.182 |
|
|
0.123 |
| 17 |
H |
0.184 |
|
|
0.129 |
| 18 |
H |
0.184 |
|
|
0.129 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.567 |
1.193 |
0.000 |
1.969 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-1.611 |
1.180 |
0.000 |
1.997 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.326 |
3.596 |
0.000 |
| y |
3.596 |
-48.175 |
0.000 |
| z |
0.000 |
0.000 |
-48.969 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.246 |
3.596 |
0.000 |
| y |
3.596 |
-0.527 |
0.000 |
| z |
0.000 |
0.000 |
-1.719 |
|
| Polar |
|---|
| 3z2-r2 | -3.438 |
|---|
| x2-y2 | 1.849 |
|---|
| xy | 3.596 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.147 |
0.011 |
0.000 |
| y |
0.011 |
11.823 |
0.000 |
| z |
0.000 |
0.000 |
8.152 |
<r2> (average value of r
2) Å
2
| <r2> |
248.686 |
| (<r2>)1/2 |
15.770 |