Jump to
S1C2
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -245.898260 |
| Energy at 298.15K | -245.903941 |
| HF Energy | -245.898260 |
| Nuclear repulsion energy | 155.132737 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3150 |
3040 |
15.24 |
|
|
|
| 2 |
A' |
3003 |
2898 |
43.16 |
|
|
|
| 3 |
A' |
2991 |
2886 |
18.85 |
|
|
|
| 4 |
A' |
2367 |
2283 |
0.55 |
|
|
|
| 5 |
A' |
1579 |
1524 |
6.10 |
|
|
|
| 6 |
A' |
1562 |
1507 |
8.05 |
|
|
|
| 7 |
A' |
1508 |
1455 |
0.90 |
|
|
|
| 8 |
A' |
1424 |
1374 |
26.89 |
|
|
|
| 9 |
A' |
1191 |
1149 |
7.99 |
|
|
|
| 10 |
A' |
1087 |
1049 |
114.93 |
|
|
|
| 11 |
A' |
958 |
924 |
3.67 |
|
|
|
| 12 |
A' |
917 |
885 |
23.59 |
|
|
|
| 13 |
A' |
584 |
564 |
1.11 |
|
|
|
| 14 |
A' |
379 |
366 |
1.77 |
|
|
|
| 15 |
A' |
185 |
178 |
0.73 |
|
|
|
| 16 |
A" |
3047 |
2940 |
55.35 |
|
|
|
| 17 |
A" |
3013 |
2907 |
31.35 |
|
|
|
| 18 |
A" |
1550 |
1496 |
10.30 |
|
|
|
| 19 |
A" |
1279 |
1234 |
0.56 |
|
|
|
| 20 |
A" |
1164 |
1123 |
1.38 |
|
|
|
| 21 |
A" |
1057 |
1020 |
1.40 |
|
|
|
| 22 |
A" |
461 |
445 |
0.09 |
|
|
|
| 23 |
A" |
241 |
233 |
5.33 |
|
|
|
| 24 |
A" |
93 |
89 |
0.76 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17394.6 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 16784.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.150 |
-0.440 |
0.000 |
| O2 |
-0.704 |
-0.647 |
0.000 |
| C3 |
0.000 |
0.627 |
0.000 |
| C4 |
1.431 |
0.341 |
0.000 |
| N5 |
2.570 |
0.110 |
0.000 |
| H6 |
-2.586 |
-1.440 |
0.000 |
| H7 |
-2.485 |
0.106 |
0.894 |
| H8 |
-2.485 |
0.106 |
-0.894 |
| H9 |
-0.248 |
1.229 |
-0.889 |
| H10 |
-0.248 |
1.229 |
0.889 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4607 | 2.4000 | 3.6649 | 4.7519 | 1.0907 | 1.0997 | 1.0997 | 2.6817 | 2.6817 |
O2 | 1.4607 | | 1.4556 | 2.3523 | 3.3604 | 2.0422 | 2.1303 | 2.1303 | 2.1255 | 2.1255 | C3 | 2.4000 | 1.4556 | | 1.4595 | 2.6218 | 3.3103 | 2.6918 | 2.6918 | 1.1018 | 1.1018 | C4 | 3.6649 | 2.3523 | 1.4595 | | 1.1623 | 4.3938 | 4.0236 | 4.0236 | 2.0974 | 2.0974 | N5 | 4.7519 | 3.3604 | 2.6218 | 1.1623 | | 5.3839 | 5.1336 | 5.1336 | 3.1601 | 3.1601 | H6 | 1.0907 | 2.0422 | 3.3103 | 4.3938 | 5.3839 | | 1.7882 | 1.7882 | 3.6575 | 3.6575 | H7 | 1.0997 | 2.1303 | 2.6918 | 4.0236 | 5.1336 | 1.7882 | | 1.7878 | 3.0730 | 2.5030 | H8 | 1.0997 | 2.1303 | 2.6918 | 4.0236 | 5.1336 | 1.7882 | 1.7878 | | 2.5030 | 3.0730 | H9 | 2.6817 | 2.1255 | 1.1018 | 2.0974 | 3.1601 | 3.6575 | 3.0730 | 2.5030 | | 1.7777 | H10 | 2.6817 | 2.1255 | 1.1018 | 2.0974 | 3.1601 | 3.6575 | 2.5030 | 3.0730 | 1.7777 | |
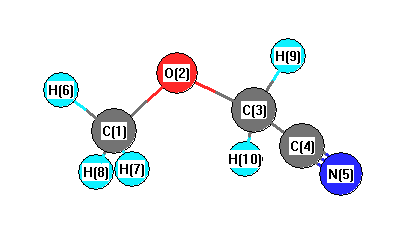 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
110.761 |
|
O2 |
C1 |
H6 |
105.419 |
| O2 |
C1 |
H7 |
111.848 |
|
O2 |
C1 |
H8 |
111.848 |
| O2 |
C3 |
C4 |
107.593 |
|
O2 |
C3 |
H9 |
111.678 |
| O2 |
C3 |
H10 |
111.678 |
|
C3 |
C4 |
N5 |
179.864 |
| C4 |
C3 |
H9 |
109.151 |
|
C4 |
C3 |
H10 |
109.151 |
| H6 |
C1 |
H7 |
109.450 |
|
H6 |
C1 |
H8 |
109.450 |
| H7 |
C1 |
H8 |
108.754 |
|
H9 |
C3 |
H10 |
107.551 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.324 |
|
|
|
| 2 |
O |
-0.461 |
|
|
|
| 3 |
C |
-0.139 |
|
|
|
| 4 |
C |
0.343 |
|
|
|
| 5 |
N |
-0.468 |
|
|
|
| 6 |
H |
0.223 |
|
|
|
| 7 |
H |
0.184 |
|
|
|
| 8 |
H |
0.184 |
|
|
|
| 9 |
H |
0.229 |
|
|
|
| 10 |
H |
0.229 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.208 |
1.873 |
0.000 |
4.606 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.762 |
0.726 |
0.000 |
| y |
0.726 |
-28.908 |
0.000 |
| z |
0.000 |
0.000 |
-29.132 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.742 |
0.726 |
0.000 |
| y |
0.726 |
3.039 |
0.000 |
| z |
0.000 |
0.000 |
2.703 |
|
| Polar |
|---|
| 3z2-r2 | 5.405 |
|---|
| x2-y2 | -5.854 |
|---|
| xy | 0.726 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.538 |
0.136 |
0.000 |
| y |
0.136 |
4.240 |
0.000 |
| z |
0.000 |
0.000 |
3.860 |
<r2> (average value of r
2) Å
2
| <r2> |
144.648 |
| (<r2>)1/2 |
12.027 |
Jump to
S1C1
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -245.902071 |
| Energy at 298.15K | -245.907832 |
| HF Energy | -245.902071 |
| Nuclear repulsion energy | 157.972721 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3153 |
3042 |
16.14 |
|
|
|
| 2 |
A |
3114 |
3004 |
3.68 |
|
|
|
| 3 |
A |
3069 |
2962 |
38.81 |
|
|
|
| 4 |
A |
3014 |
2909 |
38.46 |
|
|
|
| 5 |
A |
3008 |
2903 |
22.75 |
|
|
|
| 6 |
A |
2345 |
2263 |
0.74 |
|
|
|
| 7 |
A |
1576 |
1520 |
10.72 |
|
|
|
| 8 |
A |
1550 |
1496 |
11.23 |
|
|
|
| 9 |
A |
1532 |
1478 |
6.72 |
|
|
|
| 10 |
A |
1505 |
1453 |
0.67 |
|
|
|
| 11 |
A |
1397 |
1348 |
6.36 |
|
|
|
| 12 |
A |
1323 |
1276 |
3.13 |
|
|
|
| 13 |
A |
1198 |
1156 |
2.70 |
|
|
|
| 14 |
A |
1164 |
1123 |
1.27 |
|
|
|
| 15 |
A |
1092 |
1054 |
88.09 |
|
|
|
| 16 |
A |
1034 |
998 |
13.55 |
|
|
|
| 17 |
A |
907 |
875 |
26.49 |
|
|
|
| 18 |
A |
877 |
846 |
14.87 |
|
|
|
| 19 |
A |
620 |
599 |
1.74 |
|
|
|
| 20 |
A |
443 |
428 |
0.55 |
|
|
|
| 21 |
A |
386 |
372 |
4.02 |
|
|
|
| 22 |
A |
262 |
253 |
10.60 |
|
|
|
| 23 |
A |
190 |
184 |
3.82 |
|
|
|
| 24 |
A |
124 |
119 |
9.86 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17441.1 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 16828.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.577 |
-0.790 |
0.143 |
| O2 |
1.095 |
0.462 |
-0.447 |
| C3 |
-0.098 |
0.950 |
0.220 |
| C4 |
-1.254 |
0.059 |
0.044 |
| N5 |
-2.145 |
-0.673 |
-0.110 |
| H6 |
2.475 |
-1.050 |
-0.419 |
| H7 |
0.837 |
-1.594 |
0.047 |
| H8 |
1.831 |
-0.663 |
1.205 |
| H9 |
-0.313 |
1.922 |
-0.232 |
| H10 |
0.068 |
1.093 |
1.300 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4658 | 2.4164 | 2.9578 | 3.7323 | 1.0907 | 1.0972 | 1.0993 | 3.3272 | 2.6762 |
O2 | 1.4658 | | 1.4516 | 2.4337 | 3.4497 | 2.0468 | 2.1305 | 2.1296 | 2.0402 | 2.1225 | C3 | 2.4164 | 1.4516 | | 1.4698 | 2.6330 | 3.3206 | 2.7157 | 2.6998 | 1.0937 | 1.1012 | C4 | 2.9578 | 2.4337 | 1.4698 | | 1.1635 | 3.9181 | 2.6659 | 3.3746 | 2.1049 | 2.0961 | N5 | 3.7323 | 3.4497 | 2.6330 | 1.1635 | | 4.6452 | 3.1244 | 4.1875 | 3.1790 | 3.1628 | H6 | 1.0907 | 2.0468 | 3.3206 | 3.9181 | 4.6452 | | 1.7881 | 1.7897 | 4.0793 | 3.6524 | H7 | 1.0972 | 2.1305 | 2.7157 | 2.6659 | 3.1244 | 1.7881 | | 1.7882 | 3.7101 | 3.0630 | H8 | 1.0993 | 2.1296 | 2.6998 | 3.3746 | 4.1875 | 1.7897 | 1.7882 | | 3.6525 | 2.4899 | H9 | 3.3272 | 2.0402 | 1.0937 | 2.1049 | 3.1790 | 4.0793 | 3.7101 | 3.6525 | | 1.7822 | H10 | 2.6762 | 2.1225 | 1.1012 | 2.0961 | 3.1628 | 3.6524 | 3.0630 | 2.4899 | 1.7822 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
111.841 |
|
O2 |
C1 |
H6 |
105.442 |
| O2 |
C1 |
H7 |
111.648 |
|
O2 |
C1 |
H8 |
111.447 |
| O2 |
C3 |
C4 |
112.826 |
|
O2 |
C3 |
H9 |
105.696 |
| O2 |
C3 |
H10 |
111.753 |
|
C3 |
C4 |
N5 |
178.024 |
| C4 |
C3 |
H9 |
109.513 |
|
C4 |
C3 |
H10 |
108.372 |
| H6 |
C1 |
H7 |
109.625 |
|
H6 |
C1 |
H8 |
109.615 |
| H7 |
C1 |
H8 |
109.000 |
|
H9 |
C3 |
H10 |
108.575 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.339 |
|
|
|
| 2 |
O |
-0.453 |
|
|
|
| 3 |
C |
-0.171 |
|
|
|
| 4 |
C |
0.328 |
|
|
|
| 5 |
N |
-0.468 |
|
|
|
| 6 |
H |
0.221 |
|
|
|
| 7 |
H |
0.205 |
|
|
|
| 8 |
H |
0.183 |
|
|
|
| 9 |
H |
0.264 |
|
|
|
| 10 |
H |
0.229 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.119 |
1.003 |
1.555 |
2.813 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
4.201 |
1.906 |
0.000 |
4.613 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.495 |
-5.945 |
0.682 |
| y |
-5.945 |
-27.816 |
0.585 |
| z |
0.682 |
0.585 |
-29.592 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.791 |
-5.945 |
0.682 |
| y |
-5.945 |
4.727 |
0.585 |
| z |
0.682 |
0.585 |
2.064 |
|
| Polar |
|---|
| 3z2-r2 | 4.127 |
|---|
| x2-y2 | -7.678 |
|---|
| xy | -5.945 |
|---|
| xz | 0.682 |
|---|
| yz | 0.585 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.253 |
0.336 |
0.160 |
| y |
0.336 |
5.057 |
0.191 |
| z |
0.160 |
0.191 |
3.894 |
<r2> (average value of r
2) Å
2
| <r2> |
124.353 |
| (<r2>)1/2 |
11.151 |
