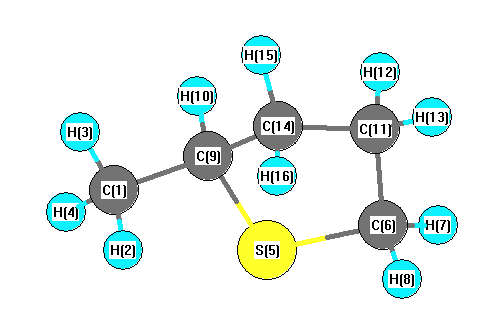
Vibrational Frequencies calculated at BLYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3088 |
3071 |
15.34 |
|
|
|
| 2 |
A |
3055 |
3039 |
35.79 |
|
|
|
| 3 |
A |
3034 |
3017 |
42.64 |
|
|
|
| 4 |
A |
3030 |
3013 |
22.23 |
|
|
|
| 5 |
A |
3025 |
3009 |
5.80 |
|
|
|
| 6 |
A |
3017 |
3000 |
5.55 |
|
|
|
| 7 |
A |
3005 |
2989 |
7.33 |
|
|
|
| 8 |
A |
2982 |
2966 |
21.09 |
|
|
|
| 9 |
A |
2973 |
2957 |
32.11 |
|
|
|
| 10 |
A |
2966 |
2950 |
16.21 |
|
|
|
| 11 |
A |
1529 |
1521 |
7.46 |
|
|
|
| 12 |
A |
1518 |
1510 |
5.49 |
|
|
|
| 13 |
A |
1512 |
1503 |
0.26 |
|
|
|
| 14 |
A |
1504 |
1496 |
6.58 |
|
|
|
| 15 |
A |
1499 |
1491 |
2.35 |
|
|
|
| 16 |
A |
1424 |
1416 |
5.08 |
|
|
|
| 17 |
A |
1360 |
1352 |
2.70 |
|
|
|
| 18 |
A |
1353 |
1345 |
0.63 |
|
|
|
| 19 |
A |
1324 |
1317 |
0.98 |
|
|
|
| 20 |
A |
1277 |
1270 |
19.02 |
|
|
|
| 21 |
A |
1267 |
1260 |
10.20 |
|
|
|
| 22 |
A |
1217 |
1211 |
6.00 |
|
|
|
| 23 |
A |
1175 |
1169 |
13.50 |
|
|
|
| 24 |
A |
1124 |
1118 |
1.38 |
|
|
|
| 25 |
A |
1077 |
1071 |
0.72 |
|
|
|
| 26 |
A |
1064 |
1058 |
1.60 |
|
|
|
| 27 |
A |
1014 |
1009 |
6.63 |
|
|
|
| 28 |
A |
978 |
973 |
0.66 |
|
|
|
| 29 |
A |
971 |
965 |
4.34 |
|
|
|
| 30 |
A |
929 |
924 |
4.07 |
|
|
|
| 31 |
A |
864 |
860 |
1.03 |
|
|
|
| 32 |
A |
846 |
842 |
2.16 |
|
|
|
| 33 |
A |
814 |
810 |
3.24 |
|
|
|
| 34 |
A |
608 |
605 |
4.09 |
|
|
|
| 35 |
A |
564 |
561 |
5.26 |
|
|
|
| 36 |
A |
506 |
503 |
1.64 |
|
|
|
| 37 |
A |
436 |
433 |
0.27 |
|
|
|
| 38 |
A |
391 |
389 |
1.15 |
|
|
|
| 39 |
A |
276 |
275 |
1.75 |
|
|
|
| 40 |
A |
253 |
252 |
0.29 |
|
|
|
| 41 |
A |
232 |
230 |
0.17 |
|
|
|
| 42 |
A |
92 |
91 |
1.49 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30585.7 cm
-1
Scaled (by 0.9945) Zero Point Vibrational Energy (zpe) 30417.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.505 |
|
|
|
| 2 |
H |
0.192 |
|
|
|
| 3 |
H |
0.179 |
|
|
|
| 4 |
H |
0.191 |
|
|
|
| 5 |
S |
0.174 |
|
|
|
| 6 |
C |
-0.548 |
|
|
|
| 7 |
H |
0.212 |
|
|
|
| 8 |
H |
0.210 |
|
|
|
| 9 |
C |
-0.391 |
|
|
|
| 10 |
H |
0.210 |
|
|
|
| 11 |
C |
-0.349 |
|
|
|
| 12 |
H |
0.191 |
|
|
|
| 13 |
H |
0.187 |
|
|
|
| 14 |
C |
-0.326 |
|
|
|
| 15 |
H |
0.185 |
|
|
|
| 16 |
H |
0.188 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.314 |
2.386 |
0.046 |
2.407 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.289 |
-0.225 |
0.940 |
| y |
-0.225 |
-48.887 |
-0.371 |
| z |
0.940 |
-0.371 |
-46.279 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.294 |
-0.225 |
0.940 |
| y |
-0.225 |
-4.603 |
-0.371 |
| z |
0.940 |
-0.371 |
-0.691 |
|
| Polar |
|---|
| 3z2-r2 | -1.382 |
|---|
| x2-y2 | 6.598 |
|---|
| xy | -0.225 |
|---|
| xz | 0.940 |
|---|
| yz | -0.371 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
213.180 |
| (<r2>)1/2 |
14.601 |