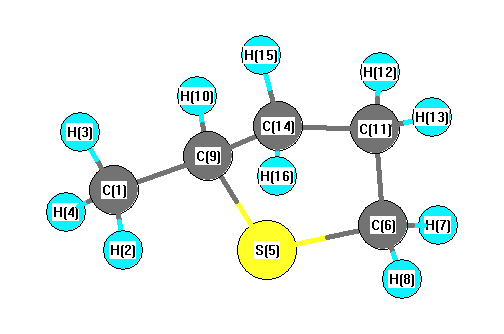
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3171 |
3060 |
11.49 |
|
|
|
| 2 |
A |
3136 |
3026 |
33.78 |
|
|
|
| 3 |
A |
3117 |
3008 |
32.10 |
|
|
|
| 4 |
A |
3112 |
3003 |
20.75 |
|
|
|
| 5 |
A |
3107 |
2998 |
7.14 |
|
|
|
| 6 |
A |
3101 |
2992 |
4.61 |
|
|
|
| 7 |
A |
3091 |
2982 |
6.68 |
|
|
|
| 8 |
A |
3064 |
2956 |
17.75 |
|
|
|
| 9 |
A |
3055 |
2948 |
24.53 |
|
|
|
| 10 |
A |
3046 |
2939 |
17.92 |
|
|
|
| 11 |
A |
1564 |
1509 |
9.04 |
|
|
|
| 12 |
A |
1554 |
1500 |
6.59 |
|
|
|
| 13 |
A |
1548 |
1494 |
0.38 |
|
|
|
| 14 |
A |
1540 |
1486 |
8.43 |
|
|
|
| 15 |
A |
1535 |
1481 |
3.68 |
|
|
|
| 16 |
A |
1464 |
1413 |
7.41 |
|
|
|
| 17 |
A |
1399 |
1350 |
2.62 |
|
|
|
| 18 |
A |
1391 |
1342 |
0.77 |
|
|
|
| 19 |
A |
1363 |
1315 |
2.20 |
|
|
|
| 20 |
A |
1317 |
1271 |
15.60 |
|
|
|
| 21 |
A |
1308 |
1262 |
12.59 |
|
|
|
| 22 |
A |
1256 |
1212 |
6.41 |
|
|
|
| 23 |
A |
1217 |
1174 |
11.76 |
|
|
|
| 24 |
A |
1162 |
1122 |
1.05 |
|
|
|
| 25 |
A |
1114 |
1075 |
0.40 |
|
|
|
| 26 |
A |
1100 |
1062 |
1.86 |
|
|
|
| 27 |
A |
1051 |
1014 |
7.25 |
|
|
|
| 28 |
A |
1017 |
981 |
0.57 |
|
|
|
| 29 |
A |
1001 |
965 |
4.87 |
|
|
|
| 30 |
A |
961 |
928 |
4.89 |
|
|
|
| 31 |
A |
894 |
863 |
1.08 |
|
|
|
| 32 |
A |
872 |
841 |
2.04 |
|
|
|
| 33 |
A |
843 |
814 |
3.53 |
|
|
|
| 34 |
A |
640 |
618 |
4.20 |
|
|
|
| 35 |
A |
596 |
575 |
5.16 |
|
|
|
| 36 |
A |
527 |
508 |
1.36 |
|
|
|
| 37 |
A |
454 |
438 |
0.12 |
|
|
|
| 38 |
A |
406 |
392 |
0.99 |
|
|
|
| 39 |
A |
288 |
278 |
1.65 |
|
|
|
| 40 |
A |
260 |
250 |
0.26 |
|
|
|
| 41 |
A |
240 |
231 |
0.18 |
|
|
|
| 42 |
A |
95 |
91 |
1.63 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31487.4 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 30382.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.535 |
|
|
-0.552 |
| 2 |
H |
0.205 |
|
|
0.144 |
| 3 |
H |
0.192 |
|
|
0.129 |
| 4 |
H |
0.203 |
|
|
0.191 |
| 5 |
S |
0.208 |
|
|
-0.341 |
| 6 |
C |
-0.594 |
|
|
-0.038 |
| 7 |
H |
0.228 |
|
|
0.106 |
| 8 |
H |
0.226 |
|
|
0.100 |
| 9 |
C |
-0.439 |
|
|
0.231 |
| 10 |
H |
0.227 |
|
|
0.054 |
| 11 |
C |
-0.380 |
|
|
-0.328 |
| 12 |
H |
0.206 |
|
|
0.127 |
| 13 |
H |
0.203 |
|
|
0.120 |
| 14 |
C |
-0.353 |
|
|
-0.112 |
| 15 |
H |
0.201 |
|
|
0.069 |
| 16 |
H |
0.203 |
|
|
0.098 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.334 |
2.419 |
0.044 |
2.443 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.342 |
2.407 |
0.028 |
2.432 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.269 |
-0.269 |
0.943 |
| y |
-0.269 |
-48.953 |
-0.380 |
| z |
0.943 |
-0.380 |
-46.184 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.299 |
-0.269 |
0.943 |
| y |
-0.269 |
-4.726 |
-0.380 |
| z |
0.943 |
-0.380 |
-0.573 |
|
| Polar |
|---|
| 3z2-r2 | -1.146 |
|---|
| x2-y2 | 6.684 |
|---|
| xy | -0.269 |
|---|
| xz | 0.943 |
|---|
| yz | -0.380 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.986 |
-0.141 |
0.410 |
| y |
-0.141 |
9.935 |
-0.040 |
| z |
0.410 |
-0.040 |
7.437 |
<r2> (average value of r
2) Å
2
| <r2> |
209.730 |
| (<r2>)1/2 |
14.482 |