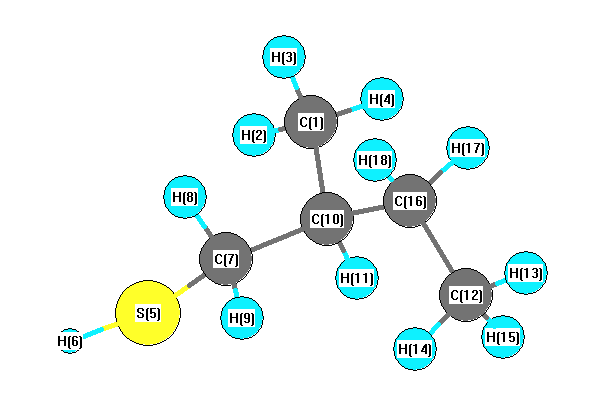
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3164 |
3053 |
13.50 |
|
|
|
| 2 |
A |
3126 |
3016 |
23.42 |
|
|
|
| 3 |
A |
3118 |
3008 |
65.37 |
|
|
|
| 4 |
A |
3112 |
3002 |
36.59 |
|
|
|
| 5 |
A |
3108 |
2999 |
8.99 |
|
|
|
| 6 |
A |
3095 |
2987 |
11.41 |
|
|
|
| 7 |
A |
3072 |
2965 |
7.57 |
|
|
|
| 8 |
A |
3054 |
2947 |
21.76 |
|
|
|
| 9 |
A |
3050 |
2943 |
10.61 |
|
|
|
| 10 |
A |
3046 |
2939 |
16.83 |
|
|
|
| 11 |
A |
3037 |
2930 |
11.27 |
|
|
|
| 12 |
A |
2464 |
2378 |
42.61 |
|
|
|
| 13 |
A |
1574 |
1519 |
10.65 |
|
|
|
| 14 |
A |
1567 |
1512 |
10.57 |
|
|
|
| 15 |
A |
1565 |
1510 |
7.67 |
|
|
|
| 16 |
A |
1564 |
1509 |
2.57 |
|
|
|
| 17 |
A |
1547 |
1492 |
2.90 |
|
|
|
| 18 |
A |
1528 |
1474 |
7.71 |
|
|
|
| 19 |
A |
1465 |
1414 |
1.82 |
|
|
|
| 20 |
A |
1463 |
1412 |
17.09 |
|
|
|
| 21 |
A |
1413 |
1364 |
2.41 |
|
|
|
| 22 |
A |
1392 |
1343 |
1.52 |
|
|
|
| 23 |
A |
1367 |
1319 |
0.61 |
|
|
|
| 24 |
A |
1340 |
1293 |
10.18 |
|
|
|
| 25 |
A |
1300 |
1254 |
14.60 |
|
|
|
| 26 |
A |
1261 |
1217 |
6.07 |
|
|
|
| 27 |
A |
1198 |
1156 |
0.40 |
|
|
|
| 28 |
A |
1160 |
1119 |
3.06 |
|
|
|
| 29 |
A |
1096 |
1057 |
2.51 |
|
|
|
| 30 |
A |
1045 |
1009 |
3.23 |
|
|
|
| 31 |
A |
1026 |
990 |
2.87 |
|
|
|
| 32 |
A |
1014 |
978 |
9.03 |
|
|
|
| 33 |
A |
932 |
899 |
0.63 |
|
|
|
| 34 |
A |
885 |
854 |
6.41 |
|
|
|
| 35 |
A |
872 |
842 |
1.79 |
|
|
|
| 36 |
A |
816 |
788 |
2.45 |
|
|
|
| 37 |
A |
764 |
738 |
11.81 |
|
|
|
| 38 |
A |
687 |
663 |
6.01 |
|
|
|
| 39 |
A |
460 |
444 |
0.29 |
|
|
|
| 40 |
A |
393 |
379 |
0.11 |
|
|
|
| 41 |
A |
382 |
369 |
1.57 |
|
|
|
| 42 |
A |
257 |
248 |
0.10 |
|
|
|
| 43 |
A |
236 |
227 |
1.59 |
|
|
|
| 44 |
A |
213 |
206 |
0.12 |
|
|
|
| 45 |
A |
197 |
191 |
2.35 |
|
|
|
| 46 |
A |
146 |
141 |
20.85 |
|
|
|
| 47 |
A |
103 |
99 |
0.03 |
|
|
|
| 48 |
A |
70 |
67 |
11.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35371.1 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 34129.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.537 |
|
|
-0.577 |
| 2 |
H |
0.216 |
|
|
0.197 |
| 3 |
H |
0.184 |
|
|
0.140 |
| 4 |
H |
0.187 |
|
|
0.132 |
| 5 |
S |
-0.007 |
|
|
-0.377 |
| 6 |
H |
0.068 |
|
|
0.229 |
| 7 |
C |
-0.571 |
|
|
-0.490 |
| 8 |
H |
0.223 |
|
|
0.203 |
| 9 |
H |
0.231 |
|
|
0.207 |
| 10 |
C |
-0.240 |
|
|
0.354 |
| 11 |
H |
0.207 |
|
|
0.043 |
| 12 |
C |
-0.559 |
|
|
-0.274 |
| 13 |
H |
0.191 |
|
|
0.095 |
| 14 |
H |
0.187 |
|
|
0.065 |
| 15 |
H |
0.191 |
|
|
0.071 |
| 16 |
C |
-0.351 |
|
|
-0.137 |
| 17 |
H |
0.194 |
|
|
0.060 |
| 18 |
H |
0.185 |
|
|
0.060 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.655 |
-0.789 |
0.998 |
2.087 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
1.666 |
-0.814 |
1.018 |
2.115 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-50.481 |
2.574 |
-2.095 |
| y |
2.574 |
-46.070 |
-1.621 |
| z |
-2.095 |
-1.621 |
-48.202 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.345 |
2.574 |
-2.095 |
| y |
2.574 |
3.271 |
-1.621 |
| z |
-2.095 |
-1.621 |
0.074 |
|
| Polar |
|---|
| 3z2-r2 | 0.147 |
|---|
| x2-y2 | -4.410 |
|---|
| xy | 2.574 |
|---|
| xz | -2.095 |
|---|
| yz | -1.621 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.048 |
0.537 |
0.245 |
| y |
0.537 |
9.380 |
-0.460 |
| z |
0.245 |
-0.460 |
7.931 |
<r2> (average value of r
2) Å
2
| <r2> |
293.896 |
| (<r2>)1/2 |
17.143 |