Jump to
S1C2
Vibrational Frequencies calculated at B3LYP/3-21G
Geometric Data calculated at B3LYP/3-21G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -234.569285 |
| Energy at 298.15K | -234.582735 |
| Nuclear repulsion energy | 252.831497 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3153 |
3042 |
54.17 |
|
|
|
| 2 |
A' |
3125 |
3016 |
13.55 |
|
|
|
| 3 |
A' |
3105 |
2996 |
40.20 |
|
|
|
| 4 |
A' |
3100 |
2991 |
18.11 |
|
|
|
| 5 |
A' |
3092 |
2983 |
17.92 |
|
|
|
| 6 |
A' |
3078 |
2970 |
9.14 |
|
|
|
| 7 |
A' |
3037 |
2930 |
26.12 |
|
|
|
| 8 |
A' |
3031 |
2925 |
23.78 |
|
|
|
| 9 |
A' |
1567 |
1512 |
4.68 |
|
|
|
| 10 |
A' |
1556 |
1502 |
4.99 |
|
|
|
| 11 |
A' |
1550 |
1496 |
2.42 |
|
|
|
| 12 |
A' |
1532 |
1478 |
1.35 |
|
|
|
| 13 |
A' |
1464 |
1413 |
2.66 |
|
|
|
| 14 |
A' |
1447 |
1396 |
11.96 |
|
|
|
| 15 |
A' |
1336 |
1289 |
6.30 |
|
|
|
| 16 |
A' |
1299 |
1254 |
11.11 |
|
|
|
| 17 |
A' |
1253 |
1209 |
0.78 |
|
|
|
| 18 |
A' |
1214 |
1172 |
0.74 |
|
|
|
| 19 |
A' |
1017 |
981 |
1.08 |
|
|
|
| 20 |
A' |
996 |
961 |
0.23 |
|
|
|
| 21 |
A' |
956 |
922 |
0.43 |
|
|
|
| 22 |
A' |
906 |
874 |
0.74 |
|
|
|
| 23 |
A' |
874 |
843 |
0.49 |
|
|
|
| 24 |
A' |
704 |
680 |
2.07 |
|
|
|
| 25 |
A' |
609 |
588 |
1.00 |
|
|
|
| 26 |
A' |
390 |
376 |
0.21 |
|
|
|
| 27 |
A' |
347 |
335 |
0.08 |
|
|
|
| 28 |
A' |
135 |
130 |
0.01 |
|
|
|
| 29 |
A" |
3132 |
3022 |
8.33 |
|
|
|
| 30 |
A" |
3097 |
2989 |
50.60 |
|
|
|
| 31 |
A" |
3090 |
2982 |
2.65 |
|
|
|
| 32 |
A" |
3073 |
2966 |
51.07 |
|
|
|
| 33 |
A" |
1572 |
1516 |
7.75 |
|
|
|
| 34 |
A" |
1550 |
1496 |
0.09 |
|
|
|
| 35 |
A" |
1526 |
1472 |
5.28 |
|
|
|
| 36 |
A" |
1296 |
1250 |
0.66 |
|
|
|
| 37 |
A" |
1281 |
1236 |
0.01 |
|
|
|
| 38 |
A" |
1246 |
1202 |
0.69 |
|
|
|
| 39 |
A" |
1194 |
1152 |
6.37 |
|
|
|
| 40 |
A" |
1119 |
1080 |
0.28 |
|
|
|
| 41 |
A" |
1027 |
991 |
0.00 |
|
|
|
| 42 |
A" |
912 |
880 |
1.40 |
|
|
|
| 43 |
A" |
867 |
836 |
0.03 |
|
|
|
| 44 |
A" |
798 |
770 |
0.20 |
|
|
|
| 45 |
A" |
384 |
371 |
0.37 |
|
|
|
| 46 |
A" |
309 |
298 |
0.01 |
|
|
|
| 47 |
A" |
274 |
264 |
0.02 |
|
|
|
| 48 |
A" |
234 |
226 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36927.8 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 35631.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.091 |
0.417 |
0.000 |
| C2 |
0.325 |
-1.760 |
0.000 |
| C3 |
-0.091 |
-0.717 |
1.102 |
| C4 |
-0.091 |
-0.717 |
-1.102 |
| C5 |
1.234 |
1.194 |
0.000 |
| C6 |
-1.297 |
1.360 |
0.000 |
| H7 |
1.402 |
-1.944 |
0.000 |
| H8 |
-0.212 |
-2.711 |
0.000 |
| H9 |
0.596 |
-0.577 |
1.940 |
| H10 |
0.596 |
-0.577 |
-1.940 |
| H11 |
-1.103 |
-0.908 |
1.475 |
| H12 |
-1.103 |
-0.908 |
-1.475 |
| H13 |
2.089 |
0.508 |
0.000 |
| H14 |
-2.234 |
0.792 |
0.000 |
| H15 |
1.304 |
1.832 |
-0.890 |
| H16 |
1.304 |
1.832 |
0.890 |
| H17 |
-1.283 |
2.003 |
0.889 |
| H18 |
-1.283 |
2.003 |
-0.889 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 2.2160 | 1.5814 | 1.5814 | 1.5364 | 1.5307 | 2.7932 | 3.1294 | 2.2857 | 2.2857 | 2.2251 | 2.2251 | 2.1826 | 2.1754 | 2.1774 | 2.1774 | 2.1742 | 2.1742 |
C2 | 2.2160 | | 1.5732 | 1.5732 | 3.0905 | 3.5162 | 1.0928 | 1.0914 | 2.2882 | 2.2882 | 2.2223 | 2.2223 | 2.8737 | 3.6137 | 3.8277 | 3.8277 | 4.1876 | 4.1876 | C3 | 1.5814 | 1.5732 | | 2.2045 | 2.5738 | 2.6427 | 2.2247 | 2.2807 | 1.0929 | 3.1221 | 1.0942 | 2.7748 | 2.7336 | 2.8432 | 3.5233 | 2.9138 | 2.9777 | 3.5759 | C4 | 1.5814 | 1.5732 | 2.2045 | | 2.5738 | 2.6427 | 2.2247 | 2.2807 | 3.1221 | 1.0929 | 2.7748 | 1.0942 | 2.7336 | 2.8432 | 2.9138 | 3.5233 | 3.5759 | 2.9777 | C5 | 1.5364 | 3.0905 | 2.5738 | 2.5738 | | 2.5362 | 3.1421 | 4.1633 | 2.7031 | 2.7031 | 3.4713 | 3.4713 | 1.0962 | 3.4914 | 1.0972 | 1.0972 | 2.7890 | 2.7890 | C6 | 1.5307 | 3.5162 | 2.6427 | 2.6427 | 2.5362 | | 4.2660 | 4.2127 | 3.3318 | 3.3318 | 2.7120 | 2.7120 | 3.4916 | 1.0962 | 2.7888 | 2.7888 | 1.0976 | 1.0976 | H7 | 2.7932 | 1.0928 | 2.2247 | 2.2247 | 3.1421 | 4.2660 | | 1.7864 | 2.5060 | 2.5060 | 3.0856 | 3.0856 | 2.5466 | 4.5502 | 3.8804 | 3.8804 | 4.8556 | 4.8556 | H8 | 3.1294 | 1.0914 | 2.2807 | 2.2807 | 4.1633 | 4.2127 | 1.7864 | | 2.9944 | 2.9944 | 2.4938 | 2.4938 | 3.9565 | 4.0442 | 4.8704 | 4.8704 | 4.9150 | 4.9150 | H9 | 2.2857 | 2.2882 | 1.0929 | 3.1221 | 2.7031 | 3.3318 | 2.5060 | 2.9944 | | 3.8799 | 1.7921 | 3.8283 | 2.6778 | 3.6945 | 3.7832 | 2.7216 | 3.3607 | 4.2654 | H10 | 2.2857 | 2.2882 | 3.1221 | 1.0929 | 2.7031 | 3.3318 | 2.5060 | 2.9944 | 3.8799 | | 3.8283 | 1.7921 | 2.6778 | 3.6945 | 2.7216 | 3.7832 | 4.2654 | 3.3607 | H11 | 2.2251 | 2.2223 | 1.0942 | 2.7748 | 3.4713 | 2.7120 | 3.0856 | 2.4938 | 1.7921 | 3.8283 | | 2.9495 | 3.7904 | 2.5186 | 4.3459 | 3.6928 | 2.9747 | 3.7542 | H12 | 2.2251 | 2.2223 | 2.7748 | 1.0942 | 3.4713 | 2.7120 | 3.0856 | 2.4938 | 3.8283 | 1.7921 | 2.9495 | | 3.7904 | 2.5186 | 3.6928 | 4.3459 | 3.7542 | 2.9747 | H13 | 2.1826 | 2.8737 | 2.7336 | 2.7336 | 1.0962 | 3.4916 | 2.5466 | 3.9565 | 2.6778 | 2.6778 | 3.7904 | 3.7904 | | 4.3327 | 1.7779 | 1.7779 | 3.7941 | 3.7941 | H14 | 2.1754 | 3.6137 | 2.8432 | 2.8432 | 3.4914 | 1.0962 | 4.5502 | 4.0442 | 3.6945 | 3.6945 | 2.5186 | 2.5186 | 4.3327 | | 3.7936 | 3.7936 | 1.7789 | 1.7789 | H15 | 2.1774 | 3.8277 | 3.5233 | 2.9138 | 1.0972 | 2.7888 | 3.8804 | 4.8704 | 3.7832 | 2.7216 | 4.3459 | 3.6928 | 1.7779 | 3.7936 | | 1.7799 | 3.1438 | 2.5920 | H16 | 2.1774 | 3.8277 | 2.9138 | 3.5233 | 1.0972 | 2.7888 | 3.8804 | 4.8704 | 2.7216 | 3.7832 | 3.6928 | 4.3459 | 1.7779 | 3.7936 | 1.7799 | | 2.5920 | 3.1438 | H17 | 2.1742 | 4.1876 | 2.9777 | 3.5759 | 2.7890 | 1.0976 | 4.8556 | 4.9150 | 3.3607 | 4.2654 | 2.9747 | 3.7542 | 3.7941 | 1.7789 | 3.1438 | 2.5920 | | 1.7783 | H18 | 2.1742 | 4.1876 | 3.5759 | 2.9777 | 2.7890 | 1.0976 | 4.8556 | 4.9150 | 4.2654 | 3.3607 | 3.7542 | 2.9747 | 3.7941 | 1.7789 | 2.5920 | 3.1438 | 1.7783 | |
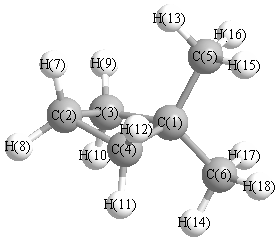 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C2 |
89.247 |
|
C1 |
C3 |
H9 |
116.251 |
| C1 |
C3 |
H11 |
111.221 |
|
C1 |
C4 |
C2 |
89.247 |
| C1 |
C4 |
H10 |
116.251 |
|
C1 |
C4 |
H12 |
111.221 |
| C1 |
C5 |
H13 |
110.899 |
|
C1 |
C5 |
H15 |
110.428 |
| C1 |
C5 |
H16 |
110.428 |
|
C1 |
C6 |
H14 |
110.721 |
| C1 |
C6 |
H17 |
110.547 |
|
C1 |
C6 |
H18 |
110.547 |
| C2 |
C3 |
H9 |
117.099 |
|
C2 |
C3 |
H11 |
111.578 |
| C2 |
C4 |
H10 |
117.099 |
|
C2 |
C4 |
H12 |
111.578 |
| C3 |
C1 |
C4 |
88.371 |
|
C3 |
C1 |
C5 |
111.270 |
| C3 |
C1 |
C6 |
116.227 |
|
C3 |
C2 |
C4 |
88.957 |
| C3 |
C2 |
H7 |
111.856 |
|
C3 |
C2 |
H8 |
116.562 |
| C4 |
C1 |
C5 |
111.270 |
|
C4 |
C1 |
C6 |
116.227 |
| C4 |
C2 |
H7 |
111.856 |
|
C4 |
C2 |
H8 |
116.562 |
| C5 |
C1 |
C6 |
111.565 |
|
H7 |
C2 |
H8 |
109.743 |
| H9 |
C3 |
H11 |
110.050 |
|
H10 |
C4 |
H12 |
110.050 |
| H13 |
C5 |
H15 |
108.302 |
|
H13 |
C5 |
H16 |
108.302 |
| H14 |
C6 |
H17 |
108.362 |
|
H14 |
C6 |
H18 |
108.362 |
| H15 |
C5 |
H16 |
108.399 |
|
H17 |
C6 |
H18 |
108.218 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.105 |
|
|
|
| 2 |
C |
-0.384 |
|
|
|
| 3 |
C |
-0.335 |
|
|
|
| 4 |
C |
-0.335 |
|
|
|
| 5 |
C |
-0.514 |
|
|
|
| 6 |
C |
-0.532 |
|
|
|
| 7 |
H |
0.192 |
|
|
|
| 8 |
H |
0.183 |
|
|
|
| 9 |
H |
0.178 |
|
|
|
| 10 |
H |
0.178 |
|
|
|
| 11 |
H |
0.188 |
|
|
|
| 12 |
H |
0.188 |
|
|
|
| 13 |
H |
0.184 |
|
|
|
| 14 |
H |
0.180 |
|
|
|
| 15 |
H |
0.182 |
|
|
|
| 16 |
H |
0.182 |
|
|
|
| 17 |
H |
0.183 |
|
|
|
| 18 |
H |
0.183 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.015 |
0.056 |
0.000 |
0.058 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.502 |
0.238 |
0.000 |
| y |
0.238 |
-40.364 |
0.000 |
| z |
0.000 |
0.000 |
-40.187 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.773 |
0.238 |
0.000 |
| y |
0.238 |
-0.520 |
0.000 |
| z |
0.000 |
0.000 |
-0.253 |
|
| Polar |
|---|
| 3z2-r2 | -0.507 |
|---|
| x2-y2 | 0.862 |
|---|
| xy | 0.238 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.377 |
-0.075 |
0.000 |
| y |
-0.075 |
9.117 |
0.000 |
| z |
0.000 |
0.000 |
8.273 |
<r2> (average value of r
2) Å
2
| <r2> |
170.336 |
| (<r2>)1/2 |
13.051 |
