Jump to
S1C2
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -133.209632 |
| Energy at 298.15K | -133.215128 |
| HF Energy | -133.209632 |
| Nuclear repulsion energy | 70.493565 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3268 |
3153 |
17.90 |
|
|
|
| 2 |
A' |
3155 |
3044 |
9.64 |
|
|
|
| 3 |
A' |
3044 |
2937 |
9.65 |
|
|
|
| 4 |
A' |
2993 |
2888 |
66.24 |
|
|
|
| 5 |
A' |
1706 |
1646 |
31.76 |
|
|
|
| 6 |
A' |
1534 |
1480 |
10.76 |
|
|
|
| 7 |
A' |
1448 |
1398 |
13.99 |
|
|
|
| 8 |
A' |
1426 |
1376 |
31.96 |
|
|
|
| 9 |
A' |
1297 |
1251 |
62.36 |
|
|
|
| 10 |
A' |
1078 |
1040 |
27.46 |
|
|
|
| 11 |
A' |
916 |
884 |
13.77 |
|
|
|
| 12 |
A' |
490 |
472 |
16.34 |
|
|
|
| 13 |
A" |
3091 |
2982 |
15.21 |
|
|
|
| 14 |
A" |
1549 |
1495 |
11.88 |
|
|
|
| 15 |
A" |
1145 |
1105 |
5.83 |
|
|
|
| 16 |
A" |
1113 |
1074 |
8.26 |
|
|
|
| 17 |
A" |
703 |
678 |
73.25 |
|
|
|
| 18 |
A" |
199 |
192 |
0.58 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15076.8 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 14547.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.043 |
-0.631 |
0.000 |
| C2 |
0.000 |
0.459 |
0.000 |
| N3 |
1.247 |
0.176 |
0.000 |
| H4 |
-0.541 |
-1.601 |
0.000 |
| H5 |
-1.686 |
-0.548 |
0.886 |
| H6 |
-1.686 |
-0.548 |
-0.886 |
| H7 |
-0.410 |
1.478 |
0.000 |
| H8 |
1.847 |
1.023 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5082 | 2.4278 | 1.0919 | 1.0976 | 1.0976 | 2.2023 | 3.3297 |
C2 | 1.5082 | | 1.2790 | 2.1297 | 2.1537 | 2.1537 | 1.0992 | 1.9314 | N3 | 2.4278 | 1.2790 | | 2.5210 | 3.1479 | 3.1479 | 2.1083 | 1.0383 | H4 | 1.0919 | 2.1297 | 2.5210 | | 1.7898 | 1.7898 | 3.0824 | 3.5484 | H5 | 1.0976 | 2.1537 | 3.1479 | 1.7898 | | 1.7710 | 2.5527 | 3.9664 | H6 | 1.0976 | 2.1537 | 3.1479 | 1.7898 | 1.7710 | | 2.5527 | 3.9664 | H7 | 2.2023 | 1.0992 | 2.1083 | 3.0824 | 2.5527 | 2.5527 | | 2.3028 | H8 | 3.3297 | 1.9314 | 1.0383 | 3.5484 | 3.9664 | 3.9664 | 2.3028 | |
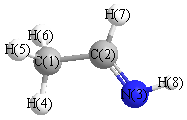 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
120.947 |
|
C1 |
C2 |
H7 |
114.353 |
| C2 |
C1 |
H4 |
108.938 |
|
C2 |
C1 |
H5 |
110.498 |
| C2 |
C1 |
H6 |
110.498 |
|
C2 |
N3 |
H8 |
112.504 |
| N3 |
C2 |
H7 |
124.700 |
|
H4 |
C1 |
H5 |
109.661 |
| H4 |
C1 |
H6 |
109.661 |
|
H5 |
C1 |
H6 |
107.571 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.606 |
|
|
|
| 2 |
C |
0.070 |
|
|
|
| 3 |
N |
-0.511 |
|
|
|
| 4 |
H |
0.223 |
|
|
|
| 5 |
H |
0.204 |
|
|
|
| 6 |
H |
0.204 |
|
|
|
| 7 |
H |
0.165 |
|
|
|
| 8 |
H |
0.249 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.165 |
1.496 |
0.000 |
1.896 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.134 |
2.697 |
0.000 |
| y |
2.697 |
-17.729 |
0.000 |
| z |
0.000 |
0.000 |
-19.615 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.462 |
2.697 |
0.000 |
| y |
2.697 |
1.645 |
0.000 |
| z |
0.000 |
0.000 |
-1.183 |
|
| Polar |
|---|
| 3z2-r2 | -2.366 |
|---|
| x2-y2 | -1.405 |
|---|
| xy | 2.697 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.347 |
0.370 |
0.000 |
| y |
0.370 |
3.748 |
0.000 |
| z |
0.000 |
0.000 |
2.380 |
<r2> (average value of r
2) Å
2
| <r2> |
50.558 |
| (<r2>)1/2 |
7.110 |
Jump to
S1C1
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -133.209727 |
| Energy at 298.15K | -133.215169 |
| HF Energy | -133.209727 |
| Nuclear repulsion energy | 70.318015 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3224 |
3110 |
28.77 |
|
|
|
| 2 |
A' |
3122 |
3012 |
29.64 |
|
|
|
| 3 |
A' |
3086 |
2978 |
19.33 |
|
|
|
| 4 |
A' |
3039 |
2932 |
7.70 |
|
|
|
| 5 |
A' |
1692 |
1633 |
23.89 |
|
|
|
| 6 |
A' |
1539 |
1485 |
26.04 |
|
|
|
| 7 |
A' |
1449 |
1398 |
9.69 |
|
|
|
| 8 |
A' |
1445 |
1394 |
27.28 |
|
|
|
| 9 |
A' |
1299 |
1254 |
102.12 |
|
|
|
| 10 |
A' |
1081 |
1043 |
24.73 |
|
|
|
| 11 |
A' |
892 |
861 |
3.25 |
|
|
|
| 12 |
A' |
488 |
471 |
6.79 |
|
|
|
| 13 |
A" |
3092 |
2983 |
11.01 |
|
|
|
| 14 |
A" |
1541 |
1487 |
10.75 |
|
|
|
| 15 |
A" |
1167 |
1126 |
52.44 |
|
|
|
| 16 |
A" |
1110 |
1071 |
10.99 |
|
|
|
| 17 |
A" |
708 |
683 |
6.43 |
|
|
|
| 18 |
A" |
165 |
159 |
1.72 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15069.5 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 14540.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.032 |
-0.628 |
0.000 |
| C2 |
0.000 |
0.480 |
0.000 |
| N3 |
1.275 |
0.388 |
0.000 |
| H4 |
-0.555 |
-1.614 |
0.000 |
| H5 |
-1.677 |
-0.546 |
0.884 |
| H6 |
-1.677 |
-0.546 |
-0.884 |
| H7 |
-0.420 |
1.489 |
0.000 |
| H8 |
1.597 |
-0.604 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5146 | 2.5212 | 1.0953 | 1.0977 | 1.0977 | 2.2040 | 2.6297 |
C2 | 1.5146 | | 1.2785 | 2.1665 | 2.1561 | 2.1561 | 1.0927 | 1.9302 | N3 | 2.5212 | 1.2785 | | 2.7123 | 3.2205 | 3.2205 | 2.0212 | 1.0424 | H4 | 1.0953 | 2.1665 | 2.7123 | | 1.7839 | 1.7839 | 3.1060 | 2.3777 | H5 | 1.0977 | 2.1561 | 3.2205 | 1.7839 | | 1.7685 | 2.5505 | 3.3925 | H6 | 1.0977 | 2.1561 | 3.2205 | 1.7839 | 1.7685 | | 2.5505 | 3.3925 | H7 | 2.2040 | 1.0927 | 2.0212 | 3.1060 | 2.5505 | 2.5505 | | 2.9063 | H8 | 2.6297 | 1.9302 | 1.0424 | 2.3777 | 3.3925 | 3.3925 | 2.9063 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.822 |
|
C1 |
C2 |
H7 |
114.447 |
| C2 |
C1 |
H4 |
111.203 |
|
C2 |
C1 |
H5 |
110.233 |
| C2 |
C1 |
H6 |
110.233 |
|
C2 |
N3 |
H8 |
112.144 |
| N3 |
C2 |
H7 |
116.731 |
|
H4 |
C1 |
H5 |
108.871 |
| H4 |
C1 |
H6 |
108.871 |
|
H5 |
C1 |
H6 |
107.333 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.628 |
|
|
|
| 2 |
C |
0.072 |
|
|
|
| 3 |
N |
-0.509 |
|
|
|
| 4 |
H |
0.195 |
|
|
|
| 5 |
H |
0.215 |
|
|
|
| 6 |
H |
0.215 |
|
|
|
| 7 |
H |
0.199 |
|
|
|
| 8 |
H |
0.241 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.819 |
-1.545 |
0.000 |
2.387 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.813 |
-3.309 |
0.000 |
| y |
-3.309 |
-19.040 |
0.000 |
| z |
0.000 |
0.000 |
-19.588 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.499 |
-3.309 |
0.000 |
| y |
-3.309 |
1.161 |
0.000 |
| z |
0.000 |
0.000 |
0.339 |
|
| Polar |
|---|
| 3z2-r2 | 0.677 |
|---|
| x2-y2 | -1.774 |
|---|
| xy | -3.309 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.141 |
-0.221 |
0.000 |
| y |
-0.221 |
3.794 |
0.000 |
| z |
0.000 |
0.000 |
2.372 |
<r2> (average value of r
2) Å
2
| <r2> |
50.964 |
| (<r2>)1/2 |
7.139 |
