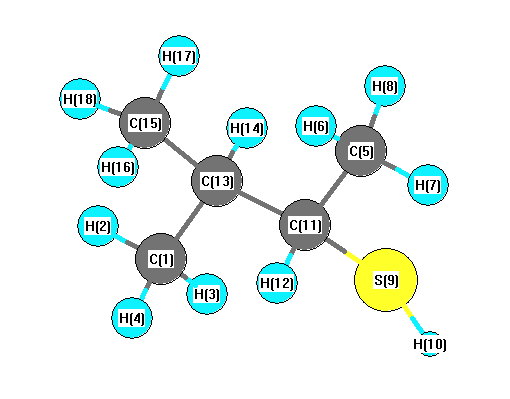
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3135 |
3025 |
20.30 |
|
|
|
| 2 |
A |
3134 |
3024 |
22.73 |
|
|
|
| 3 |
A |
3131 |
3021 |
31.14 |
|
|
|
| 4 |
A |
3115 |
3006 |
51.44 |
|
|
|
| 5 |
A |
3113 |
3004 |
21.32 |
|
|
|
| 6 |
A |
3105 |
2996 |
0.55 |
|
|
|
| 7 |
A |
3095 |
2986 |
6.67 |
|
|
|
| 8 |
A |
3057 |
2949 |
14.98 |
|
|
|
| 9 |
A |
3056 |
2948 |
15.13 |
|
|
|
| 10 |
A |
3049 |
2942 |
11.31 |
|
|
|
| 11 |
A |
3047 |
2940 |
10.42 |
|
|
|
| 12 |
A |
2464 |
2377 |
43.96 |
|
|
|
| 13 |
A |
1585 |
1529 |
4.52 |
|
|
|
| 14 |
A |
1572 |
1517 |
10.22 |
|
|
|
| 15 |
A |
1567 |
1512 |
21.64 |
|
|
|
| 16 |
A |
1558 |
1503 |
2.58 |
|
|
|
| 17 |
A |
1557 |
1502 |
3.79 |
|
|
|
| 18 |
A |
1551 |
1497 |
2.12 |
|
|
|
| 19 |
A |
1474 |
1423 |
4.27 |
|
|
|
| 20 |
A |
1466 |
1415 |
21.40 |
|
|
|
| 21 |
A |
1453 |
1402 |
7.18 |
|
|
|
| 22 |
A |
1404 |
1355 |
0.28 |
|
|
|
| 23 |
A |
1373 |
1325 |
0.65 |
|
|
|
| 24 |
A |
1358 |
1310 |
1.14 |
|
|
|
| 25 |
A |
1268 |
1224 |
28.97 |
|
|
|
| 26 |
A |
1219 |
1176 |
4.36 |
|
|
|
| 27 |
A |
1203 |
1161 |
17.67 |
|
|
|
| 28 |
A |
1151 |
1110 |
1.66 |
|
|
|
| 29 |
A |
1096 |
1058 |
5.39 |
|
|
|
| 30 |
A |
1057 |
1020 |
10.92 |
|
|
|
| 31 |
A |
1013 |
978 |
5.75 |
|
|
|
| 32 |
A |
988 |
953 |
0.15 |
|
|
|
| 33 |
A |
969 |
935 |
1.21 |
|
|
|
| 34 |
A |
917 |
884 |
4.10 |
|
|
|
| 35 |
A |
873 |
843 |
4.35 |
|
|
|
| 36 |
A |
780 |
753 |
6.05 |
|
|
|
| 37 |
A |
660 |
637 |
6.16 |
|
|
|
| 38 |
A |
478 |
462 |
0.67 |
|
|
|
| 39 |
A |
412 |
397 |
0.59 |
|
|
|
| 40 |
A |
369 |
356 |
0.28 |
|
|
|
| 41 |
A |
364 |
351 |
0.45 |
|
|
|
| 42 |
A |
331 |
320 |
2.48 |
|
|
|
| 43 |
A |
259 |
250 |
2.52 |
|
|
|
| 44 |
A |
246 |
237 |
1.00 |
|
|
|
| 45 |
A |
238 |
230 |
0.13 |
|
|
|
| 46 |
A |
211 |
203 |
3.99 |
|
|
|
| 47 |
A |
170 |
164 |
21.02 |
|
|
|
| 48 |
A |
65 |
63 |
2.56 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35376.8 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 34135.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.536 |
|
|
-0.553 |
| 2 |
H |
0.186 |
|
|
0.146 |
| 3 |
H |
0.219 |
|
|
0.170 |
| 4 |
H |
0.185 |
|
|
0.131 |
| 5 |
C |
-0.540 |
|
|
-0.307 |
| 6 |
H |
0.201 |
|
|
0.106 |
| 7 |
H |
0.192 |
|
|
0.107 |
| 8 |
H |
0.210 |
|
|
0.099 |
| 9 |
S |
-0.005 |
|
|
-0.414 |
| 10 |
H |
0.063 |
|
|
0.228 |
| 11 |
C |
-0.411 |
|
|
-0.131 |
| 12 |
H |
0.225 |
|
|
0.141 |
| 13 |
C |
-0.234 |
|
|
0.339 |
| 14 |
H |
0.206 |
|
|
0.051 |
| 15 |
C |
-0.531 |
|
|
-0.519 |
| 16 |
H |
0.186 |
|
|
0.143 |
| 17 |
H |
0.192 |
|
|
0.130 |
| 18 |
H |
0.194 |
|
|
0.133 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.146 |
1.640 |
0.771 |
2.144 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-1.177 |
1.672 |
0.816 |
2.201 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.300 |
3.089 |
2.062 |
| y |
3.089 |
-48.327 |
0.569 |
| z |
2.062 |
0.569 |
-48.185 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.956 |
3.089 |
2.062 |
| y |
3.089 |
-0.584 |
0.569 |
| z |
2.062 |
0.569 |
-0.372 |
|
| Polar |
|---|
| 3z2-r2 | -0.743 |
|---|
| x2-y2 | 1.027 |
|---|
| xy | 3.089 |
|---|
| xz | 2.062 |
|---|
| yz | 0.569 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.753 |
-0.287 |
0.167 |
| y |
-0.287 |
9.841 |
0.449 |
| z |
0.167 |
0.449 |
7.721 |
<r2> (average value of r
2) Å
2
| <r2> |
255.411 |
| (<r2>)1/2 |
15.982 |