Jump to
S1C2
Vibrational Frequencies calculated at B3LYP/3-21G
Geometric Data calculated at B3LYP/3-21G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -192.396299 |
| Energy at 298.15K | -192.400303 |
| HF Energy | -192.396299 |
| Nuclear repulsion energy | 147.990890 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3285 |
3170 |
0.08 |
|
|
|
| 2 |
A1 |
3273 |
3158 |
7.68 |
|
|
|
| 3 |
A1 |
3243 |
3129 |
1.00 |
|
|
|
| 4 |
A1 |
1566 |
1511 |
0.34 |
|
|
|
| 5 |
A1 |
1428 |
1378 |
0.44 |
|
|
|
| 6 |
A1 |
1163 |
1123 |
0.17 |
|
|
|
| 7 |
A1 |
1092 |
1054 |
11.26 |
|
|
|
| 8 |
A1 |
1020 |
984 |
8.82 |
|
|
|
| 9 |
A1 |
880 |
849 |
0.16 |
|
|
|
| 10 |
A2 |
928 |
896 |
0.00 |
|
|
|
| 11 |
A2 |
818 |
790 |
0.00 |
|
|
|
| 12 |
A2 |
537 |
518 |
0.00 |
|
|
|
| 13 |
B1 |
902 |
871 |
8.26 |
|
|
|
| 14 |
B1 |
736 |
710 |
0.65 |
|
|
|
| 15 |
B1 |
689 |
665 |
118.69 |
|
|
|
| 16 |
B1 |
509 |
491 |
13.67 |
|
|
|
| 17 |
B2 |
3258 |
3143 |
10.71 |
|
|
|
| 18 |
B2 |
3244 |
3130 |
4.61 |
|
|
|
| 19 |
B2 |
1403 |
1354 |
42.11 |
|
|
|
| 20 |
B2 |
1334 |
1287 |
0.00 |
|
|
|
| 21 |
B2 |
1230 |
1187 |
1.14 |
|
|
|
| 22 |
B2 |
975 |
941 |
2.25 |
|
|
|
| 23 |
B2 |
914 |
882 |
0.45 |
|
|
|
| 24 |
B2 |
30 |
29 |
5.76 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17228.0 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 16623.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
1.229 |
| C2 |
0.000 |
1.133 |
0.399 |
| C3 |
0.000 |
0.681 |
-1.014 |
| C4 |
0.000 |
-0.681 |
-1.014 |
| C5 |
0.000 |
-1.133 |
0.399 |
| H6 |
0.000 |
0.000 |
2.307 |
| H7 |
0.000 |
2.164 |
0.720 |
| H8 |
0.000 |
1.337 |
-1.870 |
| H9 |
0.000 |
-1.337 |
-1.870 |
| H10 |
0.000 |
-2.164 |
0.720 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4043 | 2.3443 | 2.3443 | 1.4043 | 1.0780 | 2.2236 | 3.3755 | 3.3755 | 2.2236 |
C2 | 1.4043 | | 1.4835 | 2.2990 | 2.2653 | 2.2190 | 1.0805 | 2.2784 | 3.3540 | 3.3126 | C3 | 2.3443 | 1.4835 | | 1.3617 | 2.2990 | 3.3903 | 2.2818 | 1.0788 | 2.1922 | 3.3319 | C4 | 2.3443 | 2.2990 | 1.3617 | | 1.4835 | 3.3903 | 3.3319 | 2.1922 | 1.0788 | 2.2818 | C5 | 1.4043 | 2.2653 | 2.2990 | 1.4835 | | 2.2190 | 3.3126 | 3.3540 | 2.2784 | 1.0805 | H6 | 1.0780 | 2.2190 | 3.3903 | 3.3903 | 2.2190 | | 2.6842 | 4.3861 | 4.3861 | 2.6842 | H7 | 2.2236 | 1.0805 | 2.2818 | 3.3319 | 3.3126 | 2.6842 | | 2.7187 | 4.3554 | 4.3289 | H8 | 3.3755 | 2.2784 | 1.0788 | 2.1922 | 3.3540 | 4.3861 | 2.7187 | | 2.6745 | 4.3554 | H9 | 3.3755 | 3.3540 | 2.1922 | 1.0788 | 2.2784 | 4.3861 | 4.3554 | 2.6745 | | 2.7187 | H10 | 2.2236 | 3.3126 | 3.3319 | 2.2818 | 1.0805 | 2.6842 | 4.3289 | 4.3554 | 2.7187 | |
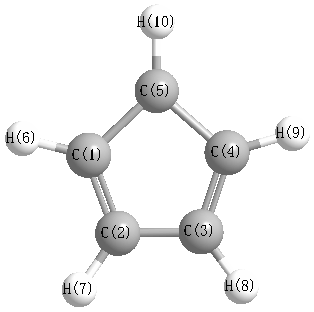 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
108.511 |
|
C1 |
C2 |
H7 |
126.496 |
| C1 |
C5 |
C4 |
108.511 |
|
C1 |
C5 |
H10 |
126.496 |
| C2 |
C1 |
C5 |
107.520 |
|
C2 |
C1 |
H6 |
126.240 |
| C2 |
C3 |
C4 |
107.730 |
|
C2 |
C3 |
H8 |
124.792 |
| C3 |
C2 |
H7 |
124.993 |
|
C3 |
C4 |
C5 |
107.730 |
| C3 |
C4 |
H9 |
127.478 |
|
C4 |
C3 |
H8 |
127.478 |
| C4 |
C5 |
H10 |
124.993 |
|
C5 |
C1 |
H6 |
126.240 |
| C5 |
C4 |
H9 |
124.792 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.213 |
|
|
|
| 2 |
C |
-0.164 |
|
|
|
| 3 |
C |
-0.195 |
|
|
|
| 4 |
C |
-0.195 |
|
|
|
| 5 |
C |
-0.164 |
|
|
|
| 6 |
H |
0.180 |
|
|
|
| 7 |
H |
0.191 |
|
|
|
| 8 |
H |
0.183 |
|
|
|
| 9 |
H |
0.183 |
|
|
|
| 10 |
H |
0.191 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.056 |
0.056 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.450 |
0.000 |
0.000 |
| y |
0.000 |
-25.180 |
0.000 |
| z |
0.000 |
0.000 |
-27.055 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.333 |
0.000 |
0.000 |
| y |
0.000 |
5.073 |
0.000 |
| z |
0.000 |
0.000 |
2.260 |
|
| Polar |
|---|
| 3z2-r2 | 4.520 |
|---|
| x2-y2 | -8.270 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.018 |
0.000 |
0.000 |
| y |
0.000 |
8.298 |
0.000 |
| z |
0.000 |
0.000 |
7.397 |
<r2> (average value of r
2) Å
2
| <r2> |
88.411 |
| (<r2>)1/2 |
9.403 |
