Jump to
S2C1
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -76.834134 |
| Energy at 298.15K | -76.834005 |
| Nuclear repulsion energy | 23.737622 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3141 |
3031 |
44.06 |
|
|
|
| 2 |
A1 |
1688 |
1629 |
79.51 |
|
|
|
| 3 |
A1 |
1293 |
1248 |
28.89 |
|
|
|
| 4 |
B1 |
822 |
793 |
133.58 |
|
|
|
| 5 |
B2 |
3207 |
3094 |
19.11 |
|
|
|
| 6 |
B2 |
504 |
487 |
0.65 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5327.5 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 5140.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-0.479 |
| C2 |
0.000 |
0.000 |
0.827 |
| H3 |
0.000 |
0.931 |
-1.044 |
| H4 |
0.000 |
-0.931 |
-1.044 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
| C1 | | 1.3062 | 1.0892 | 1.0892 |
C2 | 1.3062 | | 2.0903 | 2.0903 | H3 | 1.0892 | 2.0903 | | 1.8619 | H4 | 1.0892 | 2.0903 | 1.8619 | |
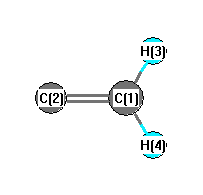 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C2 |
C1 |
H3 |
121.270 |
|
C2 |
C1 |
H4 |
121.270 |
| H3 |
C1 |
H4 |
117.460 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.580 |
|
|
|
| 2 |
C |
0.048 |
|
|
|
| 3 |
H |
0.266 |
|
|
|
| 4 |
H |
0.266 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.291 |
2.291 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-13.917 |
0.000 |
0.000 |
| y |
0.000 |
-10.061 |
0.000 |
| z |
0.000 |
0.000 |
-13.903 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.935 |
0.000 |
0.000 |
| y |
0.000 |
3.849 |
0.000 |
| z |
0.000 |
0.000 |
-1.914 |
|
| Polar |
|---|
| 3z2-r2 | -3.828 |
|---|
| x2-y2 | -3.856 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.167 |
0.000 |
0.000 |
| y |
0.000 |
2.322 |
0.000 |
| z |
0.000 |
0.000 |
3.573 |
<r2> (average value of r
2) Å
2
| <r2> |
17.283 |
| (<r2>)1/2 |
4.157 |
Jump to
S1C1
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -76.764883 |
| Energy at 298.15K | -76.764975 |
| HF Energy | -76.764883 |
| Nuclear repulsion energy | 23.562222 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3039 |
2933 |
1.69 |
|
|
|
| 2 |
A1 |
1585 |
1529 |
3.98 |
|
|
|
| 3 |
A1 |
1464 |
1413 |
3.04 |
|
|
|
| 4 |
B1 |
826 |
797 |
16.95 |
|
|
|
| 5 |
B2 |
3100 |
2991 |
8.19 |
|
|
|
| 6 |
B2 |
1046 |
1009 |
6.82 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5530.1 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 5336.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-0.483 |
| C2 |
0.000 |
0.000 |
0.835 |
| H3 |
0.000 |
0.930 |
-1.057 |
| H4 |
0.000 |
-0.930 |
-1.057 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
| C1 | | 1.3175 | 1.0935 | 1.0935 |
C2 | 1.3175 | | 2.1084 | 2.1084 | H3 | 1.0935 | 2.1084 | | 1.8610 | H4 | 1.0935 | 2.1084 | 1.8610 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.440 |
|
|
|
| 2 |
C |
0.009 |
|
|
|
| 3 |
H |
0.215 |
|
|
|
| 4 |
H |
0.215 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.576 |
0.576 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-13.653 |
0.000 |
0.000 |
| y |
0.000 |
-11.850 |
0.000 |
| z |
0.000 |
0.000 |
-11.028 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.214 |
0.000 |
0.000 |
| y |
0.000 |
0.491 |
0.000 |
| z |
0.000 |
0.000 |
1.724 |
|
| Polar |
|---|
| 3z2-r2 | 3.447 |
|---|
| x2-y2 | -1.804 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.359 |
0.000 |
0.000 |
| y |
0.000 |
1.917 |
0.000 |
| z |
0.000 |
0.000 |
3.693 |
<r2> (average value of r
2) Å
2
| <r2> |
17.152 |
| (<r2>)1/2 |
4.141 |
