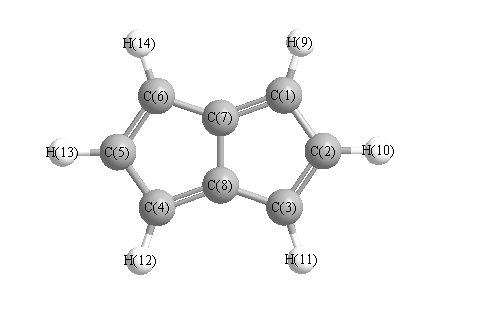
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3273 |
3158 |
0.00 |
|
|
|
| 2 |
Ag |
3251 |
3136 |
0.00 |
|
|
|
| 3 |
Ag |
3241 |
3127 |
0.00 |
|
|
|
| 4 |
Ag |
1644 |
1587 |
0.00 |
|
|
|
| 5 |
Ag |
1548 |
1494 |
0.00 |
|
|
|
| 6 |
Ag |
1372 |
1324 |
0.00 |
|
|
|
| 7 |
Ag |
1314 |
1267 |
0.00 |
|
|
|
| 8 |
Ag |
1187 |
1145 |
0.00 |
|
|
|
| 9 |
Ag |
1143 |
1103 |
0.00 |
|
|
|
| 10 |
Ag |
917 |
884 |
0.00 |
|
|
|
| 11 |
Ag |
843 |
813 |
0.00 |
|
|
|
| 12 |
Ag |
738 |
712 |
0.00 |
|
|
|
| 13 |
Ag |
655 |
632 |
0.00 |
|
|
|
| 14 |
Au |
952 |
918 |
2.11 |
|
|
|
| 15 |
Au |
897 |
865 |
24.63 |
|
|
|
| 16 |
Au |
738 |
712 |
107.27 |
|
|
|
| 17 |
Au |
523 |
505 |
11.59 |
|
|
|
| 18 |
Au |
299 |
288 |
1.58 |
|
|
|
| 19 |
Au |
157 |
152 |
9.27 |
|
|
|
| 20 |
Bg |
951 |
917 |
0.00 |
|
|
|
| 21 |
Bg |
916 |
884 |
0.00 |
|
|
|
| 22 |
Bg |
782 |
755 |
0.00 |
|
|
|
| 23 |
Bg |
665 |
642 |
0.00 |
|
|
|
| 24 |
Bg |
560 |
540 |
0.00 |
|
|
|
| 25 |
Bu |
3273 |
3158 |
23.15 |
|
|
|
| 26 |
Bu |
3251 |
3137 |
10.64 |
|
|
|
| 27 |
Bu |
3242 |
3128 |
6.67 |
|
|
|
| 28 |
Bu |
1644 |
1586 |
36.68 |
|
|
|
| 29 |
Bu |
1535 |
1481 |
10.58 |
|
|
|
| 30 |
Bu |
1360 |
1312 |
15.86 |
|
|
|
| 31 |
Bu |
1289 |
1243 |
15.43 |
|
|
|
| 32 |
Bu |
1150 |
1109 |
35.92 |
|
|
|
| 33 |
Bu |
1046 |
1009 |
25.97 |
|
|
|
| 34 |
Bu |
912 |
880 |
1.87 |
|
|
|
| 35 |
Bu |
886 |
855 |
9.48 |
|
|
|
| 36 |
Bu |
444 |
428 |
3.94 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24296.7 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 23443.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.212 |
|
|
|
| 2 |
C |
-0.201 |
|
|
|
| 3 |
C |
-0.167 |
|
|
|
| 4 |
C |
-0.212 |
|
|
|
| 5 |
C |
-0.201 |
|
|
|
| 6 |
C |
-0.167 |
|
|
|
| 7 |
C |
0.022 |
|
|
|
| 8 |
C |
0.022 |
|
|
|
| 9 |
H |
0.188 |
|
|
|
| 10 |
H |
0.186 |
|
|
|
| 11 |
H |
0.184 |
|
|
|
| 12 |
H |
0.188 |
|
|
|
| 13 |
H |
0.186 |
|
|
|
| 14 |
H |
0.184 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.695 |
1.167 |
0.000 |
| y |
1.167 |
-39.939 |
0.000 |
| z |
0.000 |
0.000 |
-51.500 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.024 |
1.167 |
0.000 |
| y |
1.167 |
5.658 |
0.000 |
| z |
0.000 |
0.000 |
-11.682 |
|
| Polar |
|---|
| 3z2-r2 | -23.365 |
|---|
| x2-y2 | 0.244 |
|---|
| xy | 1.167 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
16.414 |
1.187 |
0.000 |
| y |
1.187 |
11.211 |
0.000 |
| z |
0.000 |
0.000 |
2.990 |
<r2> (average value of r
2) Å
2
| <r2> |
219.964 |
| (<r2>)1/2 |
14.831 |