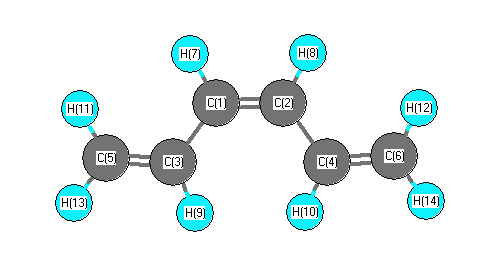
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3243 |
3129 |
29.11 |
|
|
|
| 2 |
A1 |
3183 |
3072 |
22.19 |
|
|
|
| 3 |
A1 |
3168 |
3056 |
7.12 |
|
|
|
| 4 |
A1 |
3156 |
3046 |
1.28 |
|
|
|
| 5 |
A1 |
1689 |
1630 |
0.01 |
|
|
|
| 6 |
A1 |
1638 |
1580 |
0.00 |
|
|
|
| 7 |
A1 |
1487 |
1435 |
0.17 |
|
|
|
| 8 |
A1 |
1392 |
1343 |
3.38 |
|
|
|
| 9 |
A1 |
1329 |
1282 |
0.63 |
|
|
|
| 10 |
A1 |
1136 |
1096 |
1.47 |
|
|
|
| 11 |
A1 |
898 |
866 |
0.05 |
|
|
|
| 12 |
A1 |
410 |
395 |
0.00 |
|
|
|
| 13 |
A1 |
171 |
165 |
0.03 |
|
|
|
| 14 |
A2 |
1054 |
1017 |
0.00 |
|
|
|
| 15 |
A2 |
1004 |
969 |
0.00 |
|
|
|
| 16 |
A2 |
953 |
919 |
0.00 |
|
|
|
| 17 |
A2 |
743 |
717 |
0.00 |
|
|
|
| 18 |
A2 |
335 |
323 |
0.00 |
|
|
|
| 19 |
A2 |
153 |
148 |
0.00 |
|
|
|
| 20 |
B1 |
1043 |
1006 |
33.90 |
|
|
|
| 21 |
B1 |
958 |
925 |
126.89 |
|
|
|
| 22 |
B1 |
853 |
823 |
2.92 |
|
|
|
| 23 |
B1 |
608 |
586 |
34.06 |
|
|
|
| 24 |
B1 |
102 |
98 |
0.28 |
|
|
|
| 25 |
B2 |
3242 |
3128 |
0.40 |
|
|
|
| 26 |
B2 |
3167 |
3056 |
0.30 |
|
|
|
| 27 |
B2 |
3157 |
3046 |
8.99 |
|
|
|
| 28 |
B2 |
3145 |
3035 |
0.01 |
|
|
|
| 29 |
B2 |
1687 |
1628 |
9.13 |
|
|
|
| 30 |
B2 |
1531 |
1478 |
7.80 |
|
|
|
| 31 |
B2 |
1440 |
1390 |
0.00 |
|
|
|
| 32 |
B2 |
1355 |
1307 |
0.35 |
|
|
|
| 33 |
B2 |
1234 |
1191 |
3.52 |
|
|
|
| 34 |
B2 |
988 |
953 |
4.91 |
|
|
|
| 35 |
B2 |
711 |
686 |
0.15 |
|
|
|
| 36 |
B2 |
364 |
351 |
10.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26361.8 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 25436.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.187 |
|
|
-0.189 |
| 2 |
C |
-0.187 |
|
|
-0.189 |
| 3 |
C |
-0.195 |
|
|
-0.013 |
| 4 |
C |
-0.195 |
|
|
-0.013 |
| 5 |
C |
-0.368 |
|
|
-0.462 |
| 6 |
C |
-0.368 |
|
|
-0.462 |
| 7 |
H |
0.186 |
|
|
0.154 |
| 8 |
H |
0.186 |
|
|
0.154 |
| 9 |
H |
0.189 |
|
|
0.131 |
| 10 |
H |
0.189 |
|
|
0.131 |
| 11 |
H |
0.185 |
|
|
0.196 |
| 12 |
H |
0.185 |
|
|
0.196 |
| 13 |
H |
0.190 |
|
|
0.183 |
| 14 |
H |
0.190 |
|
|
0.183 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.049 |
0.049 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.000 |
0.000 |
-0.032 |
0.032 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.931 |
0.000 |
0.000 |
| y |
0.000 |
-32.885 |
0.000 |
| z |
0.000 |
0.000 |
-33.073 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.952 |
0.000 |
0.000 |
| y |
0.000 |
4.617 |
0.000 |
| z |
0.000 |
0.000 |
4.336 |
|
| Polar |
|---|
| 3z2-r2 | 8.671 |
|---|
| x2-y2 | -9.046 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.431 |
0.000 |
0.000 |
| y |
0.000 |
20.759 |
0.000 |
| z |
0.000 |
0.000 |
7.843 |
<r2> (average value of r
2) Å
2
| <r2> |
233.183 |
| (<r2>)1/2 |
15.270 |