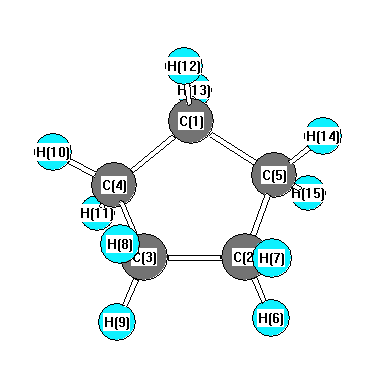
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3135 |
3025 |
80.43 |
|
|
|
| 2 |
A |
3118 |
3008 |
10.53 |
|
|
|
| 3 |
A |
3115 |
3006 |
19.33 |
|
|
|
| 4 |
A |
3109 |
3000 |
11.42 |
|
|
|
| 5 |
A |
3101 |
2992 |
29.80 |
|
|
|
| 6 |
A |
3082 |
2974 |
43.24 |
|
|
|
| 7 |
A |
3065 |
2958 |
37.73 |
|
|
|
| 8 |
A |
3065 |
2958 |
0.97 |
|
|
|
| 9 |
A |
3056 |
2949 |
28.88 |
|
|
|
| 10 |
A |
3052 |
2945 |
17.66 |
|
|
|
| 11 |
A |
1574 |
1519 |
0.26 |
|
|
|
| 12 |
A |
1552 |
1498 |
7.27 |
|
|
|
| 13 |
A |
1550 |
1496 |
3.46 |
|
|
|
| 14 |
A |
1546 |
1492 |
4.70 |
|
|
|
| 15 |
A |
1541 |
1487 |
0.66 |
|
|
|
| 16 |
A |
1376 |
1328 |
0.23 |
|
|
|
| 17 |
A |
1376 |
1327 |
0.13 |
|
|
|
| 18 |
A |
1369 |
1321 |
0.32 |
|
|
|
| 19 |
A |
1355 |
1308 |
0.46 |
|
|
|
| 20 |
A |
1351 |
1304 |
1.39 |
|
|
|
| 21 |
A |
1318 |
1272 |
1.45 |
|
|
|
| 22 |
A |
1273 |
1228 |
0.13 |
|
|
|
| 23 |
A |
1249 |
1205 |
0.61 |
|
|
|
| 24 |
A |
1233 |
1189 |
0.26 |
|
|
|
| 25 |
A |
1188 |
1147 |
0.00 |
|
|
|
| 26 |
A |
1045 |
1009 |
0.10 |
|
|
|
| 27 |
A |
1043 |
1007 |
0.05 |
|
|
|
| 28 |
A |
1027 |
991 |
0.84 |
|
|
|
| 29 |
A |
1002 |
966 |
1.54 |
|
|
|
| 30 |
A |
900 |
868 |
2.39 |
|
|
|
| 31 |
A |
893 |
861 |
2.69 |
|
|
|
| 32 |
A |
873 |
842 |
0.94 |
|
|
|
| 33 |
A |
871 |
841 |
0.88 |
|
|
|
| 34 |
A |
831 |
802 |
0.22 |
|
|
|
| 35 |
A |
779 |
752 |
1.61 |
|
|
|
| 36 |
A |
629 |
607 |
0.18 |
|
|
|
| 37 |
A |
558 |
539 |
2.02 |
|
|
|
| 38 |
A |
266 |
256 |
0.00 |
|
|
|
| 39 |
A |
13 |
13 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31239.2 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 30142.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.373 |
|
|
|
| 2 |
C |
-0.373 |
|
|
|
| 3 |
C |
-0.373 |
|
|
|
| 4 |
C |
-0.373 |
|
|
|
| 5 |
C |
-0.373 |
|
|
|
| 6 |
H |
0.189 |
|
|
|
| 7 |
H |
0.185 |
|
|
|
| 8 |
H |
0.189 |
|
|
|
| 9 |
H |
0.189 |
|
|
|
| 10 |
H |
0.189 |
|
|
|
| 11 |
H |
0.185 |
|
|
|
| 12 |
H |
0.183 |
|
|
|
| 13 |
H |
0.186 |
|
|
|
| 14 |
H |
0.186 |
|
|
|
| 15 |
H |
0.183 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.002 |
-0.000 |
0.001 |
0.002 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.021 |
0.005 |
0.001 |
0.021 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.529 |
-0.000 |
0.003 |
| y |
-0.000 |
-33.514 |
-0.012 |
| z |
0.003 |
-0.012 |
-32.622 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.461 |
-0.000 |
0.003 |
| y |
-0.000 |
-0.438 |
-0.012 |
| z |
0.003 |
-0.012 |
0.899 |
|
| Polar |
|---|
| 3z2-r2 | 1.798 |
|---|
| x2-y2 | -0.015 |
|---|
| xy | -0.000 |
|---|
| xz | 0.003 |
|---|
| yz | -0.012 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.296 |
-0.000 |
-0.000 |
| y |
-0.000 |
7.301 |
0.005 |
| z |
-0.000 |
0.005 |
6.677 |
<r2> (average value of r
2) Å
2
| <r2> |
116.962 |
| (<r2>)1/2 |
10.815 |