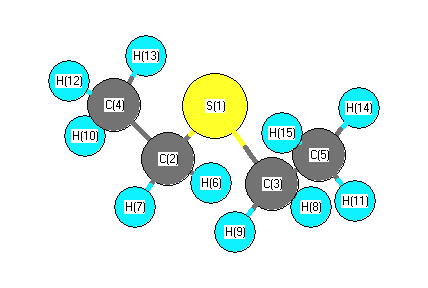
Vibrational Frequencies calculated at BLYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3037 |
3020 |
43.13 |
|
|
|
| 2 |
A1 |
3001 |
2984 |
30.59 |
|
|
|
| 3 |
A1 |
2979 |
2962 |
1.63 |
|
|
|
| 4 |
A1 |
1536 |
1527 |
5.47 |
|
|
|
| 5 |
A1 |
1504 |
1496 |
0.01 |
|
|
|
| 6 |
A1 |
1425 |
1418 |
6.15 |
|
|
|
| 7 |
A1 |
1290 |
1283 |
12.25 |
|
|
|
| 8 |
A1 |
1077 |
1071 |
0.28 |
|
|
|
| 9 |
A1 |
953 |
948 |
4.43 |
|
|
|
| 10 |
A1 |
610 |
607 |
0.18 |
|
|
|
| 11 |
A1 |
299 |
297 |
1.13 |
|
|
|
| 12 |
A1 |
124 |
124 |
0.49 |
|
|
|
| 13 |
A2 |
3068 |
3051 |
0.00 |
|
|
|
| 14 |
A2 |
3040 |
3024 |
0.00 |
|
|
|
| 15 |
A2 |
1525 |
1517 |
0.00 |
|
|
|
| 16 |
A2 |
1266 |
1259 |
0.00 |
|
|
|
| 17 |
A2 |
1016 |
1011 |
0.00 |
|
|
|
| 18 |
A2 |
778 |
774 |
0.00 |
|
|
|
| 19 |
A2 |
238 |
236 |
0.00 |
|
|
|
| 20 |
A2 |
63 |
62 |
0.00 |
|
|
|
| 21 |
B1 |
3067 |
3050 |
73.67 |
|
|
|
| 22 |
B1 |
3040 |
3023 |
0.26 |
|
|
|
| 23 |
B1 |
1525 |
1516 |
15.58 |
|
|
|
| 24 |
B1 |
1271 |
1264 |
0.43 |
|
|
|
| 25 |
B1 |
1031 |
1025 |
0.28 |
|
|
|
| 26 |
B1 |
796 |
792 |
13.37 |
|
|
|
| 27 |
B1 |
237 |
236 |
0.12 |
|
|
|
| 28 |
B1 |
37 |
37 |
0.91 |
|
|
|
| 29 |
B2 |
3037 |
3021 |
5.96 |
|
|
|
| 30 |
B2 |
3001 |
2984 |
0.43 |
|
|
|
| 31 |
B2 |
2979 |
2962 |
45.93 |
|
|
|
| 32 |
B2 |
1535 |
1527 |
1.66 |
|
|
|
| 33 |
B2 |
1498 |
1490 |
10.09 |
|
|
|
| 34 |
B2 |
1425 |
1418 |
4.16 |
|
|
|
| 35 |
B2 |
1251 |
1244 |
68.61 |
|
|
|
| 36 |
B2 |
1036 |
1030 |
9.06 |
|
|
|
| 37 |
B2 |
948 |
942 |
8.28 |
|
|
|
| 38 |
B2 |
591 |
588 |
7.17 |
|
|
|
| 39 |
B2 |
325 |
323 |
1.81 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28728.8 cm
-1
Scaled (by 0.9945) Zero Point Vibrational Energy (zpe) 28570.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.194 |
|
|
|
| 2 |
C |
-0.560 |
|
|
|
| 3 |
C |
-0.560 |
|
|
|
| 4 |
C |
-0.526 |
|
|
|
| 5 |
C |
-0.526 |
|
|
|
| 6 |
H |
0.205 |
|
|
|
| 7 |
H |
0.205 |
|
|
|
| 8 |
H |
0.205 |
|
|
|
| 9 |
H |
0.205 |
|
|
|
| 10 |
H |
0.186 |
|
|
|
| 11 |
H |
0.186 |
|
|
|
| 12 |
H |
0.196 |
|
|
|
| 13 |
H |
0.196 |
|
|
|
| 14 |
H |
0.196 |
|
|
|
| 15 |
H |
0.196 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.807 |
1.807 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
239.034 |
| (<r2>)1/2 |
15.461 |