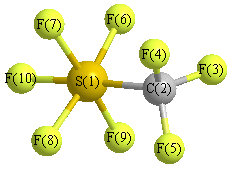
Vibrational Frequencies calculated at BLYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
1245 |
1238 |
177.46 |
|
|
|
| 2 |
A' |
1009 |
1004 |
242.42 |
|
|
|
| 3 |
A' |
814 |
809 |
171.64 |
|
|
|
| 4 |
A' |
769 |
765 |
130.93 |
|
|
|
| 5 |
A' |
661 |
657 |
76.15 |
|
|
|
| 6 |
A' |
548 |
545 |
0.20 |
|
|
|
| 7 |
A' |
545 |
542 |
19.47 |
|
|
|
| 8 |
A' |
487 |
484 |
2.07 |
|
|
|
| 9 |
A' |
433 |
430 |
47.57 |
|
|
|
| 10 |
A' |
366 |
364 |
8.34 |
|
|
|
| 11 |
A' |
279 |
277 |
0.42 |
|
|
|
| 12 |
A' |
245 |
243 |
3.91 |
|
|
|
| 13 |
A' |
240 |
238 |
3.12 |
|
|
|
| 14 |
A' |
205 |
204 |
0.97 |
|
|
|
| 15 |
A' |
140 |
139 |
0.40 |
|
|
|
| 16 |
A" |
1248 |
1241 |
171.53 |
|
|
|
| 17 |
A" |
815 |
811 |
173.08 |
|
|
|
| 18 |
A" |
486 |
483 |
2.14 |
|
|
|
| 19 |
A" |
368 |
366 |
8.61 |
|
|
|
| 20 |
A" |
306 |
305 |
0.01 |
|
|
|
| 21 |
A" |
275 |
274 |
0.56 |
|
|
|
| 22 |
A" |
236 |
235 |
2.87 |
|
|
|
| 23 |
A" |
144 |
143 |
0.40 |
|
|
|
| 24 |
A" |
28i |
28i |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5917.5 cm
-1
Scaled (by 0.9945) Zero Point Vibrational Energy (zpe) 5884.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
1.558 |
|
|
|
| 2 |
C |
0.465 |
|
|
|
| 3 |
F |
-0.191 |
|
|
|
| 4 |
F |
-0.188 |
|
|
|
| 5 |
F |
-0.188 |
|
|
|
| 6 |
F |
-0.293 |
|
|
|
| 7 |
F |
-0.298 |
|
|
|
| 8 |
F |
-0.302 |
|
|
|
| 9 |
F |
-0.298 |
|
|
|
| 10 |
F |
-0.263 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.643 |
1.908 |
0.000 |
2.014 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-59.535 |
-1.004 |
0.000 |
| y |
-1.004 |
-56.600 |
0.000 |
| z |
0.000 |
0.000 |
-59.813 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.328 |
-1.004 |
0.000 |
| y |
-1.004 |
3.073 |
0.000 |
| z |
0.000 |
0.000 |
-1.745 |
|
| Polar |
|---|
| 3z2-r2 | -3.491 |
|---|
| x2-y2 | -2.934 |
|---|
| xy | -1.004 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.371 |
-0.220 |
0.000 |
| y |
-0.220 |
6.422 |
0.000 |
| z |
0.000 |
0.000 |
5.331 |
<r2> (average value of r
2) Å
2
| <r2> |
357.949 |
| (<r2>)1/2 |
18.920 |