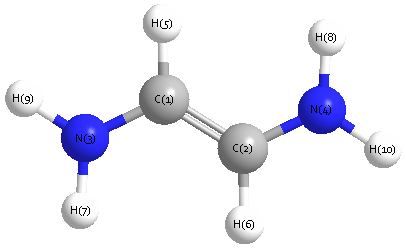
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3564 |
3439 |
0.64 |
|
|
|
| 2 |
A |
3455 |
3334 |
0.78 |
|
|
|
| 3 |
A |
3148 |
3037 |
0.12 |
|
|
|
| 4 |
A |
1741 |
1680 |
0.29 |
|
|
|
| 5 |
A |
1705 |
1645 |
14.93 |
|
|
|
| 6 |
A |
1360 |
1312 |
1.75 |
|
|
|
| 7 |
A |
1284 |
1239 |
0.02 |
|
|
|
| 8 |
A |
1021 |
985 |
0.26 |
|
|
|
| 9 |
A |
951 |
918 |
76.99 |
|
|
|
| 10 |
A |
600 |
579 |
294.56 |
|
|
|
| 11 |
A |
557 |
538 |
29.10 |
|
|
|
| 12 |
A |
352 |
340 |
2.47 |
|
|
|
| 13 |
A |
249 |
241 |
25.61 |
|
|
|
| 14 |
B |
3563 |
3438 |
1.51 |
|
|
|
| 15 |
B |
3453 |
3332 |
0.72 |
|
|
|
| 16 |
B |
3154 |
3043 |
53.95 |
|
|
|
| 17 |
B |
1708 |
1648 |
58.57 |
|
|
|
| 18 |
B |
1410 |
1360 |
3.13 |
|
|
|
| 19 |
B |
1171 |
1130 |
94.48 |
|
|
|
| 20 |
B |
1137 |
1097 |
12.80 |
|
|
|
| 21 |
B |
812 |
783 |
19.65 |
|
|
|
| 22 |
B |
526 |
508 |
368.23 |
|
|
|
| 23 |
B |
342 |
330 |
34.16 |
|
|
|
| 24 |
B |
209 |
202 |
217.69 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18735.2 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 18077.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.011 |
|
|
|
| 2 |
C |
-0.011 |
|
|
|
| 3 |
N |
-0.717 |
|
|
|
| 4 |
N |
-0.717 |
|
|
|
| 5 |
H |
0.171 |
|
|
|
| 6 |
H |
0.171 |
|
|
|
| 7 |
H |
0.279 |
|
|
|
| 8 |
H |
0.279 |
|
|
|
| 9 |
H |
0.278 |
|
|
|
| 10 |
H |
0.278 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.061 |
2.061 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.205 |
-2.289 |
0.000 |
| y |
-2.289 |
-20.482 |
0.000 |
| z |
0.000 |
0.000 |
-27.941 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.006 |
-2.289 |
0.000 |
| y |
-2.289 |
3.591 |
0.000 |
| z |
0.000 |
0.000 |
-7.597 |
|
| Polar |
|---|
| 3z2-r2 | -15.194 |
|---|
| x2-y2 | 0.277 |
|---|
| xy | -2.289 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.692 |
0.123 |
0.000 |
| y |
0.123 |
8.199 |
0.000 |
| z |
0.000 |
0.000 |
1.788 |
<r2> (average value of r
2) Å
2
| <r2> |
98.589 |
| (<r2>)1/2 |
9.929 |