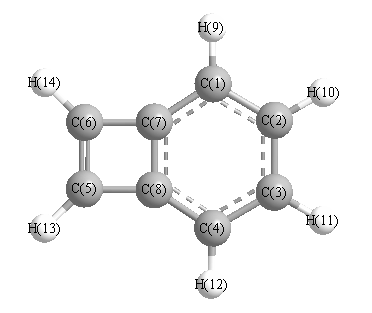
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3291 |
3176 |
10.09 |
|
|
|
| 2 |
A1 |
3222 |
3108 |
8.86 |
|
|
|
| 3 |
A1 |
3204 |
3092 |
12.22 |
|
|
|
| 4 |
A1 |
1689 |
1630 |
2.33 |
|
|
|
| 5 |
A1 |
1547 |
1493 |
1.02 |
|
|
|
| 6 |
A1 |
1475 |
1423 |
0.06 |
|
|
|
| 7 |
A1 |
1446 |
1395 |
15.71 |
|
|
|
| 8 |
A1 |
1235 |
1192 |
0.01 |
|
|
|
| 9 |
A1 |
1091 |
1052 |
0.24 |
|
|
|
| 10 |
A1 |
1072 |
1034 |
0.03 |
|
|
|
| 11 |
A1 |
963 |
929 |
2.52 |
|
|
|
| 12 |
A1 |
800 |
772 |
0.23 |
|
|
|
| 13 |
A1 |
553 |
534 |
0.06 |
|
|
|
| 14 |
A2 |
1005 |
970 |
0.00 |
|
|
|
| 15 |
A2 |
945 |
912 |
0.00 |
|
|
|
| 16 |
A2 |
905 |
873 |
0.00 |
|
|
|
| 17 |
A2 |
794 |
767 |
0.00 |
|
|
|
| 18 |
A2 |
598 |
577 |
0.00 |
|
|
|
| 19 |
A2 |
306 |
295 |
0.00 |
|
|
|
| 20 |
B1 |
945 |
912 |
10.00 |
|
|
|
| 21 |
B1 |
767 |
740 |
127.22 |
|
|
|
| 22 |
B1 |
735 |
709 |
2.57 |
|
|
|
| 23 |
B1 |
378 |
365 |
3.72 |
|
|
|
| 24 |
B1 |
230 |
222 |
13.71 |
|
|
|
| 25 |
B2 |
3258 |
3144 |
0.30 |
|
|
|
| 26 |
B2 |
3215 |
3102 |
21.48 |
|
|
|
| 27 |
B2 |
3190 |
3078 |
0.37 |
|
|
|
| 28 |
B2 |
1645 |
1588 |
0.04 |
|
|
|
| 29 |
B2 |
1483 |
1431 |
7.00 |
|
|
|
| 30 |
B2 |
1341 |
1294 |
0.27 |
|
|
|
| 31 |
B2 |
1267 |
1222 |
13.45 |
|
|
|
| 32 |
B2 |
1103 |
1064 |
2.25 |
|
|
|
| 33 |
B2 |
1069 |
1032 |
1.41 |
|
|
|
| 34 |
B2 |
870 |
839 |
0.01 |
|
|
|
| 35 |
B2 |
677 |
653 |
0.40 |
|
|
|
| 36 |
B2 |
403 |
389 |
4.49 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24357.2 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 23502.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.278 |
|
|
|
| 2 |
C |
-0.179 |
|
|
|
| 3 |
C |
-0.179 |
|
|
|
| 4 |
C |
-0.278 |
|
|
|
| 5 |
C |
-0.142 |
|
|
|
| 6 |
C |
-0.142 |
|
|
|
| 7 |
C |
0.044 |
|
|
|
| 8 |
C |
0.044 |
|
|
|
| 9 |
H |
0.184 |
|
|
|
| 10 |
H |
0.185 |
|
|
|
| 11 |
H |
0.185 |
|
|
|
| 12 |
H |
0.184 |
|
|
|
| 13 |
H |
0.186 |
|
|
|
| 14 |
H |
0.186 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.541 |
0.541 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-51.471 |
0.000 |
0.000 |
| y |
0.000 |
-40.489 |
0.000 |
| z |
0.000 |
0.000 |
-39.286 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-11.583 |
0.000 |
0.000 |
| y |
0.000 |
4.889 |
0.000 |
| z |
0.000 |
0.000 |
6.694 |
|
| Polar |
|---|
| 3z2-r2 | 13.387 |
|---|
| x2-y2 | -10.981 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.996 |
0.000 |
0.000 |
| y |
0.000 |
12.343 |
0.000 |
| z |
0.000 |
0.000 |
14.873 |
<r2> (average value of r
2) Å
2
| <r2> |
222.958 |
| (<r2>)1/2 |
14.932 |