Jump to
S1C2
Vibrational Frequencies calculated at B3LYP/3-21G
Geometric Data calculated at B3LYP/3-21G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -234.575645 |
| Energy at 298.15K | -234.587924 |
| HF Energy | -234.575645 |
| Nuclear repulsion energy | 225.637695 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3142 |
3032 |
52.92 |
|
|
|
| 2 |
A |
3128 |
3018 |
9.19 |
|
|
|
| 3 |
A |
3113 |
3004 |
50.37 |
|
|
|
| 4 |
A |
3110 |
3001 |
24.71 |
|
|
|
| 5 |
A |
3110 |
3001 |
21.48 |
|
|
|
| 6 |
A |
3085 |
2977 |
4.08 |
|
|
|
| 7 |
A |
3074 |
2966 |
23.54 |
|
|
|
| 8 |
A |
3066 |
2958 |
8.64 |
|
|
|
| 9 |
A |
3047 |
2940 |
28.15 |
|
|
|
| 10 |
A |
3044 |
2937 |
21.23 |
|
|
|
| 11 |
A |
3033 |
2926 |
30.24 |
|
|
|
| 12 |
A |
3024 |
2918 |
20.29 |
|
|
|
| 13 |
A |
1741 |
1680 |
0.32 |
|
|
|
| 14 |
A |
1572 |
1517 |
6.73 |
|
|
|
| 15 |
A |
1567 |
1512 |
7.39 |
|
|
|
| 16 |
A |
1559 |
1505 |
9.93 |
|
|
|
| 17 |
A |
1556 |
1502 |
1.97 |
|
|
|
| 18 |
A |
1551 |
1496 |
7.94 |
|
|
|
| 19 |
A |
1540 |
1486 |
2.27 |
|
|
|
| 20 |
A |
1464 |
1412 |
2.69 |
|
|
|
| 21 |
A |
1461 |
1410 |
4.73 |
|
|
|
| 22 |
A |
1397 |
1347 |
1.65 |
|
|
|
| 23 |
A |
1386 |
1337 |
0.40 |
|
|
|
| 24 |
A |
1375 |
1327 |
0.56 |
|
|
|
| 25 |
A |
1358 |
1311 |
0.06 |
|
|
|
| 26 |
A |
1337 |
1290 |
1.74 |
|
|
|
| 27 |
A |
1293 |
1248 |
0.03 |
|
|
|
| 28 |
A |
1199 |
1157 |
0.03 |
|
|
|
| 29 |
A |
1134 |
1094 |
3.19 |
|
|
|
| 30 |
A |
1107 |
1068 |
0.38 |
|
|
|
| 31 |
A |
1096 |
1057 |
6.05 |
|
|
|
| 32 |
A |
1058 |
1021 |
8.88 |
|
|
|
| 33 |
A |
1018 |
982 |
34.45 |
|
|
|
| 34 |
A |
1013 |
977 |
2.05 |
|
|
|
| 35 |
A |
950 |
916 |
3.17 |
|
|
|
| 36 |
A |
910 |
878 |
2.74 |
|
|
|
| 37 |
A |
887 |
856 |
1.65 |
|
|
|
| 38 |
A |
796 |
768 |
1.39 |
|
|
|
| 39 |
A |
764 |
737 |
3.62 |
|
|
|
| 40 |
A |
546 |
527 |
0.23 |
|
|
|
| 41 |
A |
392 |
378 |
1.35 |
|
|
|
| 42 |
A |
316 |
305 |
0.72 |
|
|
|
| 43 |
A |
292 |
282 |
1.31 |
|
|
|
| 44 |
A |
247 |
239 |
0.05 |
|
|
|
| 45 |
A |
205 |
198 |
1.02 |
|
|
|
| 46 |
A |
149 |
144 |
0.40 |
|
|
|
| 47 |
A |
93 |
90 |
0.08 |
|
|
|
| 48 |
A |
75 |
72 |
0.23 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36687.6 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 35399.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
3.182 |
0.242 |
-0.153 |
| C2 |
1.792 |
-0.340 |
-0.262 |
| C3 |
0.748 |
0.037 |
0.477 |
| C4 |
-0.648 |
-0.527 |
0.352 |
| C5 |
-1.667 |
0.542 |
-0.128 |
| C6 |
-3.101 |
-0.021 |
-0.212 |
| H7 |
3.222 |
1.023 |
0.614 |
| H8 |
3.914 |
-0.535 |
0.108 |
| H9 |
3.501 |
0.680 |
-1.108 |
| H10 |
1.661 |
-1.126 |
-1.006 |
| H11 |
0.876 |
0.825 |
1.220 |
| H12 |
-0.984 |
-0.913 |
1.325 |
| H13 |
-0.649 |
-1.367 |
-0.354 |
| H14 |
-1.350 |
0.912 |
-1.111 |
| H15 |
-1.644 |
1.395 |
0.563 |
| H16 |
-3.806 |
0.746 |
-0.553 |
| H17 |
-3.144 |
-0.863 |
-0.913 |
| H18 |
-3.432 |
-0.378 |
0.771 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.5106 | 2.5225 | 3.9394 | 4.8583 | 6.2891 | 1.0950 | 1.0982 | 1.0982 | 2.2165 | 2.7468 | 4.5695 | 4.1605 | 4.6805 | 5.0132 | 7.0176 | 6.4669 | 6.7074 |
C2 | 1.5106 | | 1.3336 | 2.5237 | 3.5724 | 4.9041 | 2.1604 | 2.1623 | 2.1622 | 1.0900 | 2.0962 | 3.2495 | 2.6506 | 3.4873 | 3.9367 | 5.7099 | 5.0066 | 5.3259 | C3 | 2.5225 | 1.3336 | | 1.5111 | 2.5404 | 3.9108 | 2.6664 | 3.2378 | 3.2410 | 2.0937 | 1.0907 | 2.1505 | 2.1482 | 2.7729 | 2.7521 | 4.7226 | 4.2298 | 4.2113 | C4 | 3.9394 | 2.5237 | 1.5111 | | 1.5526 | 2.5670 | 4.1770 | 4.5686 | 4.5612 | 2.7449 | 2.2144 | 1.1001 | 1.0976 | 2.1682 | 2.1747 | 3.5225 | 2.8180 | 2.8192 | C5 | 4.8583 | 3.5724 | 2.5404 | 1.5526 | | 1.5432 | 4.9679 | 5.6884 | 5.2617 | 3.8243 | 2.8917 | 2.1675 | 2.1755 | 1.0965 | 1.0979 | 2.1904 | 2.1846 | 2.1846 | C6 | 6.2891 | 4.9041 | 3.9108 | 2.5670 | 1.5432 | | 6.4614 | 7.0410 | 6.6994 | 4.9526 | 4.3105 | 2.7641 | 2.8009 | 2.1783 | 2.1745 | 1.0956 | 1.0965 | 1.0965 | H7 | 1.0950 | 2.1604 | 2.6664 | 4.1770 | 4.9679 | 6.4614 | | 1.7775 | 1.7778 | 3.1107 | 2.4313 | 4.6848 | 4.6513 | 4.8875 | 4.8801 | 7.1294 | 6.8126 | 6.8018 | H8 | 1.0982 | 2.1623 | 3.2378 | 4.5686 | 5.6884 | 7.0410 | 1.7775 | | 1.7676 | 2.5820 | 3.5092 | 5.0613 | 4.6614 | 5.5933 | 5.9006 | 7.8530 | 7.1390 | 7.3776 | H9 | 1.0982 | 2.1622 | 3.2410 | 4.5612 | 5.2617 | 6.6994 | 1.7778 | 1.7676 | | 2.5800 | 3.5120 | 5.3459 | 4.6887 | 4.8565 | 5.4564 | 7.3282 | 6.8247 | 7.2609 | H10 | 2.2165 | 1.0900 | 2.0937 | 2.7449 | 3.8243 | 4.9526 | 3.1107 | 2.5820 | 2.5800 | | 3.0622 | 3.5323 | 2.4125 | 3.6370 | 4.4425 | 5.7957 | 4.8131 | 5.4457 | H11 | 2.7468 | 2.0962 | 1.0907 | 2.2144 | 2.8917 | 4.3105 | 2.4313 | 3.5092 | 3.5120 | 3.0622 | | 2.5481 | 3.0999 | 3.2240 | 2.6655 | 5.0069 | 4.8535 | 4.4954 | H12 | 4.5695 | 3.2495 | 2.1505 | 1.1001 | 2.1675 | 2.7641 | 4.6848 | 5.0613 | 5.3459 | 3.5323 | 2.5481 | | 1.7716 | 3.0660 | 2.5192 | 3.7739 | 3.1106 | 2.5665 | H13 | 4.1605 | 2.6506 | 2.1482 | 1.0976 | 2.1755 | 2.8009 | 4.6513 | 4.6614 | 4.6887 | 2.4125 | 3.0999 | 1.7716 | | 2.5017 | 3.0758 | 3.8036 | 2.6060 | 3.1605 | H14 | 4.6805 | 3.4873 | 2.7729 | 2.1682 | 1.0965 | 2.1783 | 4.8875 | 5.5933 | 4.8565 | 3.6370 | 3.2240 | 3.0660 | 2.5017 | | 1.7665 | 2.5239 | 2.5315 | 3.0887 | H15 | 5.0132 | 3.9367 | 2.7521 | 2.1747 | 1.0979 | 2.1745 | 4.8801 | 5.9006 | 5.4564 | 4.4425 | 2.6655 | 2.5192 | 3.0758 | 1.7665 | | 2.5185 | 3.0865 | 2.5272 | H16 | 7.0176 | 5.7099 | 4.7226 | 3.5225 | 2.1904 | 1.0956 | 7.1294 | 7.8530 | 7.3282 | 5.7957 | 5.0069 | 3.7739 | 3.8036 | 2.5239 | 2.5185 | | 1.7758 | 1.7762 | H17 | 6.4669 | 5.0066 | 4.2298 | 2.8180 | 2.1846 | 1.0965 | 6.8126 | 7.1390 | 6.8247 | 4.8131 | 4.8535 | 3.1106 | 2.6060 | 2.5315 | 3.0865 | 1.7758 | | 1.7752 | H18 | 6.7074 | 5.3259 | 4.2113 | 2.8192 | 2.1846 | 1.0965 | 6.8018 | 7.3776 | 7.2609 | 5.4457 | 4.4954 | 2.5665 | 3.1605 | 3.0887 | 2.5272 | 1.7762 | 1.7752 | |
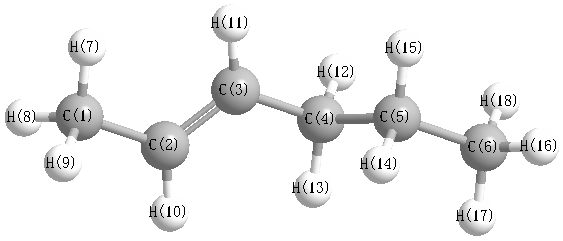 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
124.854 |
|
C1 |
C2 |
H10 |
115.978 |
| C2 |
C1 |
H7 |
111.012 |
|
C2 |
C1 |
H8 |
110.969 |
| C2 |
C1 |
H9 |
110.959 |
|
C2 |
C3 |
C4 |
124.916 |
| C2 |
C3 |
H11 |
119.352 |
|
C3 |
C2 |
H10 |
119.167 |
| C3 |
C4 |
C5 |
112.027 |
|
C3 |
C4 |
H12 |
109.882 |
| C3 |
C4 |
H13 |
109.854 |
|
C4 |
C3 |
H11 |
115.716 |
| C4 |
C5 |
C6 |
112.036 |
|
C4 |
C5 |
H14 |
108.642 |
| C4 |
C5 |
H15 |
109.059 |
|
C5 |
C4 |
H12 |
108.380 |
| C5 |
C4 |
H13 |
109.142 |
|
C5 |
C6 |
H16 |
111.075 |
| C5 |
C6 |
H17 |
110.566 |
|
C5 |
C6 |
H18 |
110.563 |
| C6 |
C5 |
H14 |
110.064 |
|
C6 |
C5 |
H15 |
109.692 |
| H7 |
C1 |
H8 |
108.277 |
|
H7 |
C1 |
H9 |
108.307 |
| H8 |
C1 |
H9 |
107.176 |
|
H12 |
C4 |
H13 |
107.434 |
| H14 |
C5 |
H15 |
107.218 |
|
H16 |
C6 |
H17 |
108.205 |
| H16 |
C6 |
H18 |
108.239 |
|
H17 |
C6 |
H18 |
108.091 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.599 |
|
|
|
| 2 |
C |
-0.167 |
|
|
|
| 3 |
C |
-0.145 |
|
|
|
| 4 |
C |
-0.418 |
|
|
|
| 5 |
C |
-0.368 |
|
|
|
| 6 |
C |
-0.553 |
|
|
|
| 7 |
H |
0.194 |
|
|
|
| 8 |
H |
0.199 |
|
|
|
| 9 |
H |
0.199 |
|
|
|
| 10 |
H |
0.172 |
|
|
|
| 11 |
H |
0.169 |
|
|
|
| 12 |
H |
0.193 |
|
|
|
| 13 |
H |
0.188 |
|
|
|
| 14 |
H |
0.194 |
|
|
|
| 15 |
H |
0.185 |
|
|
|
| 16 |
H |
0.186 |
|
|
|
| 17 |
H |
0.186 |
|
|
|
| 18 |
H |
0.185 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.024 |
0.056 |
-0.061 |
0.086 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.563 |
0.212 |
-0.708 |
| y |
0.212 |
-40.051 |
1.329 |
| z |
-0.708 |
1.329 |
-39.648 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.287 |
0.212 |
-0.708 |
| y |
0.212 |
-0.946 |
1.329 |
| z |
-0.708 |
1.329 |
-0.341 |
|
| Polar |
|---|
| 3z2-r2 | -0.682 |
|---|
| x2-y2 | 1.488 |
|---|
| xy | 0.212 |
|---|
| xz | -0.708 |
|---|
| yz | 1.329 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.457 |
-0.088 |
-1.023 |
| y |
-0.088 |
7.444 |
0.747 |
| z |
-1.023 |
0.747 |
7.692 |
<r2> (average value of r
2) Å
2
| <r2> |
295.914 |
| (<r2>)1/2 |
17.202 |
