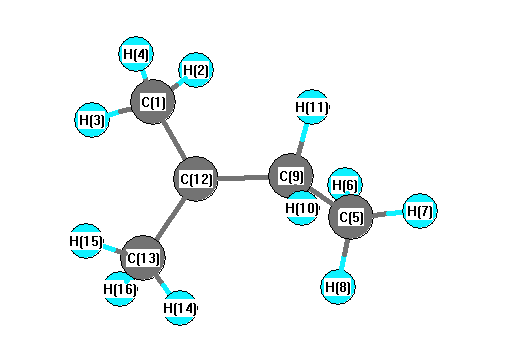
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3125 |
3015 |
37.62 |
|
|
|
| 2 |
A |
3114 |
3005 |
36.30 |
|
|
|
| 3 |
A |
3100 |
2992 |
35.02 |
|
|
|
| 4 |
A |
3095 |
2986 |
20.47 |
|
|
|
| 5 |
A |
3070 |
2962 |
10.43 |
|
|
|
| 6 |
A |
3056 |
2949 |
31.46 |
|
|
|
| 7 |
A |
3051 |
2944 |
21.62 |
|
|
|
| 8 |
A |
3048 |
2941 |
19.52 |
|
|
|
| 9 |
A |
2978 |
2873 |
40.67 |
|
|
|
| 10 |
A |
2956 |
2852 |
42.01 |
|
|
|
| 11 |
A |
2948 |
2845 |
27.17 |
|
|
|
| 12 |
A |
1571 |
1516 |
5.65 |
|
|
|
| 13 |
A |
1568 |
1513 |
6.28 |
|
|
|
| 14 |
A |
1560 |
1505 |
5.66 |
|
|
|
| 15 |
A |
1552 |
1498 |
7.66 |
|
|
|
| 16 |
A |
1547 |
1493 |
6.37 |
|
|
|
| 17 |
A |
1538 |
1484 |
3.65 |
|
|
|
| 18 |
A |
1537 |
1483 |
1.01 |
|
|
|
| 19 |
A |
1469 |
1418 |
1.88 |
|
|
|
| 20 |
A |
1454 |
1403 |
6.61 |
|
|
|
| 21 |
A |
1447 |
1396 |
5.05 |
|
|
|
| 22 |
A |
1388 |
1339 |
2.14 |
|
|
|
| 23 |
A |
1344 |
1297 |
2.42 |
|
|
|
| 24 |
A |
1296 |
1251 |
9.22 |
|
|
|
| 25 |
A |
1259 |
1215 |
4.04 |
|
|
|
| 26 |
A |
1118 |
1078 |
4.27 |
|
|
|
| 27 |
A |
1078 |
1040 |
6.77 |
|
|
|
| 28 |
A |
1049 |
1012 |
2.42 |
|
|
|
| 29 |
A |
1016 |
981 |
1.17 |
|
|
|
| 30 |
A |
1008 |
973 |
1.60 |
|
|
|
| 31 |
A |
979 |
944 |
0.62 |
|
|
|
| 32 |
A |
927 |
894 |
0.10 |
|
|
|
| 33 |
A |
804 |
776 |
3.81 |
|
|
|
| 34 |
A |
720 |
695 |
1.20 |
|
|
|
| 35 |
A |
438 |
422 |
1.47 |
|
|
|
| 36 |
A |
376 |
363 |
0.03 |
|
|
|
| 37 |
A |
338 |
326 |
1.38 |
|
|
|
| 38 |
A |
236 |
228 |
0.34 |
|
|
|
| 39 |
A |
193 |
186 |
2.04 |
|
|
|
| 40 |
A |
129 |
125 |
0.02 |
|
|
|
| 41 |
A |
108 |
105 |
0.10 |
|
|
|
| 42 |
A |
36 |
35 |
0.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32312.2 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 31178.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.595 |
|
|
-0.415 |
| 2 |
H |
0.196 |
|
|
0.110 |
| 3 |
H |
0.192 |
|
|
0.121 |
| 4 |
H |
0.190 |
|
|
0.123 |
| 5 |
C |
-0.542 |
|
|
-0.330 |
| 6 |
H |
0.188 |
|
|
0.096 |
| 7 |
H |
0.182 |
|
|
0.088 |
| 8 |
H |
0.191 |
|
|
0.083 |
| 9 |
C |
-0.431 |
|
|
0.004 |
| 10 |
H |
0.189 |
|
|
0.044 |
| 11 |
H |
0.189 |
|
|
0.032 |
| 12 |
C |
0.070 |
|
|
0.144 |
| 13 |
C |
-0.592 |
|
|
-0.552 |
| 14 |
H |
0.192 |
|
|
0.147 |
| 15 |
H |
0.190 |
|
|
0.146 |
| 16 |
H |
0.191 |
|
|
0.160 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.100 |
0.005 |
0.205 |
0.228 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.120 |
0.011 |
0.221 |
0.252 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.788 |
-0.051 |
0.306 |
| y |
-0.051 |
-33.534 |
0.054 |
| z |
0.306 |
0.054 |
-34.590 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.274 |
-0.051 |
0.306 |
| y |
-0.051 |
0.655 |
0.054 |
| z |
0.306 |
0.054 |
-0.929 |
|
| Polar |
|---|
| 3z2-r2 | -1.858 |
|---|
| x2-y2 | -0.254 |
|---|
| xy | -0.051 |
|---|
| xz | 0.306 |
|---|
| yz | 0.054 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.553 |
-0.030 |
0.167 |
| y |
-0.030 |
7.614 |
0.041 |
| z |
0.167 |
0.041 |
6.413 |
<r2> (average value of r
2) Å
2
| <r2> |
156.876 |
| (<r2>)1/2 |
12.525 |