Jump to
S1C2
Energy calculated at MP4=FULL/3-21G
| | hartrees |
|---|
| Energy at 0K | -186.270173 |
| Energy at 298.15K | -186.273531 |
| HF Energy | -185.847633 |
| Nuclear repulsion energy | 101.226677 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3631 |
3554 |
0.00 |
|
|
|
| 2 |
A |
3525 |
3450 |
0.00 |
|
|
|
| 3 |
A |
2354 |
2304 |
0.00 |
|
|
|
| 4 |
A |
1733 |
1697 |
0.00 |
|
|
|
| 5 |
A |
1196 |
1171 |
0.00 |
|
|
|
| 6 |
A |
816 |
799 |
0.00 |
|
|
|
| 7 |
A |
537 |
525 |
0.00 |
|
|
|
| 8 |
A |
525 |
514 |
0.00 |
|
|
|
| 9 |
A |
253 |
247 |
0.00 |
|
|
|
| 10 |
A |
98 |
96 |
0.00 |
|
|
|
| 11 |
B |
3631 |
3554 |
0.00 |
|
|
|
| 12 |
B |
3531 |
3456 |
0.00 |
|
|
|
| 13 |
B |
1738 |
1701 |
0.00 |
|
|
|
| 14 |
B |
1392 |
1362 |
0.00 |
|
|
|
| 15 |
B |
1196 |
1171 |
0.00 |
|
|
|
| 16 |
B |
536 |
525 |
0.00 |
|
|
|
| 17 |
B |
254 |
248 |
0.00 |
|
|
|
| 18 |
B |
105 |
103 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13525.4 cm
-1
Scaled (by 0.9788) Zero Point Vibrational Energy (zpe) 13238.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/3-21G
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.001 |
0.613 |
-0.001 |
| C2 |
-0.001 |
-0.613 |
-0.001 |
| N3 |
-0.001 |
1.966 |
-0.011 |
| N4 |
0.001 |
-1.966 |
-0.011 |
| H5 |
-0.557 |
2.492 |
0.653 |
| H6 |
0.665 |
2.485 |
-0.571 |
| H7 |
0.557 |
-2.492 |
0.653 |
| H8 |
-0.665 |
-2.485 |
-0.571 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2264 | 1.3528 | 2.5791 | 2.0664 | 2.0661 | 3.2219 | 3.2196 |
C2 | 1.2264 | | 2.5791 | 1.3528 | 3.2219 | 3.2196 | 2.0664 | 2.0661 | N3 | 1.3528 | 2.5791 | | 3.9318 | 1.0132 | 1.0132 | 4.5416 | 4.5348 | N4 | 2.5791 | 1.3528 | 3.9318 | | 4.5416 | 4.5348 | 1.0132 | 1.0132 | H5 | 2.0664 | 3.2219 | 1.0132 | 4.5416 | | 1.7295 | 5.1076 | 5.1264 | H6 | 2.0661 | 3.2196 | 1.0132 | 4.5348 | 1.7295 | | 5.1264 | 5.1448 | H7 | 3.2219 | 2.0664 | 4.5416 | 1.0132 | 5.1076 | 5.1264 | | 1.7295 | H8 | 3.2196 | 2.0661 | 4.5348 | 1.0132 | 5.1264 | 5.1448 | 1.7295 | |
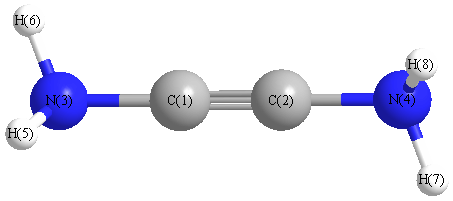 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N4 |
179.575 |
|
C1 |
N3 |
H5 |
121.039 |
| C1 |
N3 |
H6 |
121.010 |
|
C2 |
C1 |
N3 |
179.575 |
| C2 |
N4 |
H7 |
121.039 |
|
C2 |
N4 |
H8 |
121.010 |
| H5 |
N3 |
H6 |
117.183 |
|
H7 |
N4 |
H8 |
117.183 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP4=FULL/3-21G
| | hartrees |
|---|
| Energy at 0K | -186.270168 |
| Energy at 298.15K | |
| HF Energy | -185.847854 |
| Nuclear repulsion energy | 101.246898 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
2353 |
2303 |
|
|
|
|
| 2 |
A1 |
817 |
800 |
|
|
|
|
| 3 |
B1 |
529 |
518 |
|
|
|
|
| 4 |
B2 |
1395 |
1365 |
|
|
|
|
| 5 |
E |
3636 |
3559 |
|
|
|
|
| 6 |
E |
3636 |
3558 |
|
|
|
|
| 7 |
E |
3536 |
3461 |
|
|
|
|
| 8 |
E |
3530 |
3455 |
|
|
|
|
| 9 |
E |
1737 |
1700 |
|
|
|
|
| 9 |
E |
1731 |
1694 |
|
|
|
|
| 10 |
E |
1192 |
1166 |
|
|
|
|
| 10 |
E |
1191 |
1165 |
|
|
|
|
| 11 |
E |
538 |
526 |
|
|
|
|
| 11 |
E |
538 |
526 |
|
|
|
|
| 12 |
E |
253 |
248 |
|
|
|
|
| 12 |
E |
251 |
246 |
|
|
|
|
| 13 |
E |
76i |
74i |
|
|
|
|
| 13 |
E |
76i |
74i |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13354.8 cm
-1
Scaled (by 0.9788) Zero Point Vibrational Energy (zpe) 13071.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/3-21G
Point Group is D2d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.613 |
| C2 |
0.000 |
0.000 |
-0.613 |
| N3 |
0.000 |
0.000 |
1.965 |
| N4 |
0.000 |
0.000 |
-1.965 |
| H5 |
0.000 |
0.865 |
2.491 |
| H6 |
0.000 |
-0.865 |
2.491 |
| H7 |
0.865 |
0.000 |
-2.491 |
| H8 |
-0.865 |
0.000 |
-2.491 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2267 | 1.3518 | 2.5785 | 2.0677 | 2.0677 | 3.2229 | 3.2229 |
C2 | 1.2267 | | 2.5785 | 1.3518 | 3.2229 | 3.2229 | 2.0677 | 2.0677 | N3 | 1.3518 | 2.5785 | | 3.9304 | 1.0128 | 1.0128 | 4.5397 | 4.5397 | N4 | 2.5785 | 1.3518 | 3.9304 | | 4.5397 | 4.5397 | 1.0128 | 1.0128 | H5 | 2.0677 | 3.2229 | 1.0128 | 4.5397 | | 1.7310 | 5.1306 | 5.1306 | H6 | 2.0677 | 3.2229 | 1.0128 | 4.5397 | 1.7310 | | 5.1306 | 5.1306 | H7 | 3.2229 | 2.0677 | 4.5397 | 1.0128 | 5.1306 | 5.1306 | | 1.7310 | H8 | 3.2229 | 2.0677 | 4.5397 | 1.0128 | 5.1306 | 5.1306 | 1.7310 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N4 |
180.000 |
|
C1 |
N3 |
H5 |
121.291 |
| C1 |
N3 |
H6 |
121.291 |
|
C2 |
C1 |
N3 |
180.000 |
| C2 |
N4 |
H7 |
121.291 |
|
C2 |
N4 |
H8 |
121.291 |
| H5 |
N3 |
H6 |
117.418 |
|
H7 |
N4 |
H8 |
117.418 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
