Jump to
S1C2
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -187.010829 |
| Energy at 298.15K | -187.015331 |
| HF Energy | -187.010829 |
| Nuclear repulsion energy | 102.196216 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3569 |
3443 |
10.12 |
|
|
|
| 2 |
A |
3483 |
3361 |
0.00 |
|
|
|
| 3 |
A |
2378 |
2295 |
0.00 |
|
|
|
| 4 |
A |
1702 |
1643 |
0.00 |
|
|
|
| 5 |
A |
1191 |
1149 |
2.61 |
|
|
|
| 6 |
A |
860 |
830 |
0.00 |
|
|
|
| 7 |
A |
731 |
706 |
45.48 |
|
|
|
| 8 |
A |
587 |
566 |
0.00 |
|
|
|
| 9 |
A |
401 |
387 |
326.93 |
|
|
|
| 10 |
A |
280 |
270 |
43.08 |
|
|
|
| 11 |
B |
3569 |
3443 |
10.13 |
|
|
|
| 12 |
B |
3491 |
3369 |
2.05 |
|
|
|
| 13 |
B |
1712 |
1652 |
31.64 |
|
|
|
| 14 |
B |
1461 |
1410 |
116.38 |
|
|
|
| 15 |
B |
1191 |
1149 |
2.61 |
|
|
|
| 16 |
B |
732 |
706 |
45.48 |
|
|
|
| 17 |
B |
401 |
387 |
326.89 |
|
|
|
| 18 |
B |
280 |
270 |
43.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14009.3 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 13517.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.003 |
0.606 |
-0.014 |
| C2 |
-0.003 |
-0.606 |
-0.014 |
| N3 |
-0.003 |
1.964 |
-0.031 |
| N4 |
0.003 |
-1.964 |
-0.031 |
| H5 |
-0.412 |
2.461 |
0.762 |
| H6 |
0.785 |
2.448 |
-0.464 |
| H7 |
0.412 |
-2.461 |
0.762 |
| H8 |
-0.785 |
-2.448 |
-0.464 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2129 | 1.3577 | 2.5706 | 2.0530 | 2.0509 | 3.1908 | 3.1864 |
C2 | 1.2129 | | 2.5706 | 1.3577 | 3.1908 | 3.1864 | 2.0530 | 2.0509 | N3 | 1.3577 | 2.5706 | | 3.9282 | 1.0211 | 1.0214 | 4.5149 | 4.5018 | N4 | 2.5706 | 1.3577 | 3.9282 | | 4.5149 | 4.5018 | 1.0211 | 1.0214 | H5 | 2.0530 | 3.1908 | 1.0211 | 4.5149 | | 1.7138 | 4.9910 | 5.0739 | H6 | 2.0509 | 3.1864 | 1.0214 | 4.5018 | 1.7138 | | 5.0739 | 5.1416 | H7 | 3.1908 | 2.0530 | 4.5149 | 1.0211 | 4.9910 | 5.0739 | | 1.7138 | H8 | 3.1864 | 2.0509 | 4.5018 | 1.0214 | 5.0739 | 5.1416 | 1.7138 | |
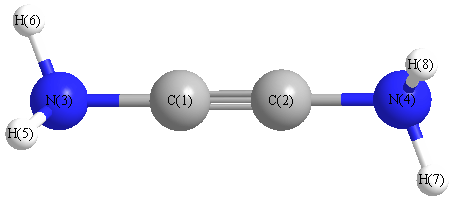 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N4 |
179.151 |
|
C1 |
N3 |
H5 |
118.626 |
| C1 |
N3 |
H6 |
118.403 |
|
C2 |
C1 |
N3 |
179.151 |
| C2 |
N4 |
H7 |
118.626 |
|
C2 |
N4 |
H8 |
118.403 |
| H5 |
N3 |
H6 |
114.077 |
|
H7 |
N4 |
H8 |
114.077 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.375 |
|
|
|
| 2 |
C |
0.375 |
|
|
|
| 3 |
N |
-0.959 |
|
|
|
| 4 |
N |
-0.959 |
|
|
|
| 5 |
H |
0.292 |
|
|
|
| 6 |
H |
0.292 |
|
|
|
| 7 |
H |
0.292 |
|
|
|
| 8 |
H |
0.292 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.003 |
0.003 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.319 |
0.013 |
0.000 |
| y |
0.013 |
-10.071 |
0.000 |
| z |
0.000 |
0.000 |
-23.319 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.624 |
0.013 |
0.000 |
| y |
0.013 |
13.249 |
0.000 |
| z |
0.000 |
0.000 |
-6.625 |
|
| Polar |
|---|
| 3z2-r2 | -13.249 |
|---|
| x2-y2 | -13.248 |
|---|
| xy | 0.013 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.930 |
0.000 |
0.000 |
| y |
0.000 |
8.788 |
0.000 |
| z |
0.000 |
0.000 |
1.930 |
<r2> (average value of r
2) Å
2
| <r2> |
96.461 |
| (<r2>)1/2 |
9.821 |
Jump to
S1C1
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -187.010818 |
| Energy at 298.15K | -187.015309 |
| HF Energy | -187.010818 |
| Nuclear repulsion energy | 102.197269 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3485 |
3363 |
0.00 |
|
|
|
| 2 |
A1 |
2378 |
2295 |
0.00 |
|
|
|
| 3 |
A1 |
1702 |
1642 |
0.00 |
|
|
|
| 4 |
A1 |
860 |
830 |
0.00 |
|
|
|
| 5 |
B1 |
584 |
564 |
0.00 |
|
|
|
| 6 |
B2 |
3493 |
3370 |
2.09 |
|
|
|
| 7 |
B2 |
1711 |
1651 |
31.65 |
|
|
|
| 8 |
B2 |
1461 |
1410 |
116.47 |
|
|
|
| 9 |
E |
3570 |
3445 |
10.17 |
|
|
|
| 9 |
E |
3570 |
3445 |
10.17 |
|
|
|
| 10 |
E |
1191 |
1149 |
2.59 |
|
|
|
| 10 |
E |
1191 |
1149 |
2.59 |
|
|
|
| 11 |
E |
731 |
706 |
45.43 |
|
|
|
| 11 |
E |
731 |
706 |
45.43 |
|
|
|
| 12 |
E |
399 |
385 |
325.84 |
|
|
|
| 12 |
E |
399 |
385 |
325.84 |
|
|
|
| 13 |
E |
278 |
269 |
44.31 |
|
|
|
| 13 |
E |
278 |
269 |
44.31 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14005.8 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 13514.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is D2d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.609 |
| C2 |
0.000 |
0.000 |
-0.609 |
| N3 |
0.000 |
0.000 |
1.941 |
| N4 |
0.000 |
0.000 |
-1.941 |
| H5 |
0.000 |
0.863 |
2.474 |
| H6 |
0.000 |
-0.863 |
2.474 |
| H7 |
0.863 |
0.000 |
-2.474 |
| H8 |
-0.863 |
0.000 |
-2.474 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2178 | 1.3319 | 2.5496 | 2.0557 | 2.0557 | 3.2020 | 3.2020 |
C2 | 1.2178 | | 2.5496 | 1.3319 | 3.2020 | 3.2020 | 2.0557 | 2.0557 | N3 | 1.3319 | 2.5496 | | 3.8815 | 1.0150 | 1.0150 | 4.4989 | 4.4989 | N4 | 2.5496 | 1.3319 | 3.8815 | | 4.4989 | 4.4989 | 1.0150 | 1.0150 | H5 | 2.0557 | 3.2020 | 1.0150 | 4.4989 | | 1.7267 | 5.0974 | 5.0974 | H6 | 2.0557 | 3.2020 | 1.0150 | 4.4989 | 1.7267 | | 5.0974 | 5.0974 | H7 | 3.2020 | 2.0557 | 4.4989 | 1.0150 | 5.0974 | 5.0974 | | 1.7267 | H8 | 3.2020 | 2.0557 | 4.4989 | 1.0150 | 5.0974 | 5.0974 | 1.7267 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N4 |
180.000 |
|
C1 |
N3 |
H5 |
121.726 |
| C1 |
N3 |
H6 |
121.726 |
|
C2 |
C1 |
N3 |
180.000 |
| C2 |
N4 |
H7 |
121.726 |
|
C2 |
N4 |
H8 |
121.726 |
| H5 |
N3 |
H6 |
116.548 |
|
H7 |
N4 |
H8 |
116.548 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.375 |
|
|
|
| 2 |
C |
0.375 |
|
|
|
| 3 |
N |
-0.959 |
|
|
|
| 4 |
N |
-0.959 |
|
|
|
| 5 |
H |
0.292 |
|
|
|
| 6 |
H |
0.292 |
|
|
|
| 7 |
H |
0.292 |
|
|
|
| 8 |
H |
0.292 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.319 |
0.000 |
0.000 |
| y |
0.000 |
-23.319 |
0.000 |
| z |
0.000 |
0.000 |
-10.073 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.623 |
0.000 |
0.000 |
| y |
0.000 |
-6.623 |
0.000 |
| z |
0.000 |
0.000 |
13.246 |
|
| Polar |
|---|
| 3z2-r2 | 26.491 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.929 |
0.000 |
0.000 |
| y |
0.000 |
1.929 |
0.000 |
| z |
0.000 |
0.000 |
8.785 |
<r2> (average value of r
2) Å
2
| <r2> |
96.461 |
| (<r2>)1/2 |
9.821 |
