Jump to
S1C2
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -153.503820 |
| Energy at 298.15K | |
| HF Energy | -153.503820 |
| Nuclear repulsion energy | 73.620498 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3416 |
3296 |
6.26 |
|
|
|
| 2 |
A' |
3163 |
3052 |
8.44 |
|
|
|
| 3 |
A' |
2990 |
2885 |
54.55 |
|
|
|
| 4 |
A' |
1586 |
1531 |
1.74 |
|
|
|
| 5 |
A' |
1508 |
1455 |
0.99 |
|
|
|
| 6 |
A' |
1456 |
1405 |
4.38 |
|
|
|
| 7 |
A' |
1254 |
1210 |
46.93 |
|
|
|
| 8 |
A' |
1009 |
973 |
17.48 |
|
|
|
| 9 |
A' |
920 |
888 |
66.57 |
|
|
|
| 10 |
A' |
624 |
602 |
26.89 |
|
|
|
| 11 |
A' |
354 |
342 |
31.16 |
|
|
|
| 12 |
A" |
3270 |
3155 |
13.69 |
|
|
|
| 13 |
A" |
3015 |
2909 |
66.02 |
|
|
|
| 14 |
A" |
1311 |
1265 |
0.06 |
|
|
|
| 15 |
A" |
1163 |
1122 |
0.42 |
|
|
|
| 16 |
A" |
842 |
813 |
0.00 |
|
|
|
| 17 |
A" |
246 |
238 |
137.36 |
|
|
|
| 18 |
A" |
133i |
128i |
1.29 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13998.1 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 13506.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.118 |
-0.417 |
0.000 |
| C2 |
0.000 |
0.557 |
0.000 |
| C3 |
1.274 |
-0.219 |
0.000 |
| H4 |
-1.958 |
0.123 |
0.000 |
| H5 |
-0.041 |
1.205 |
0.889 |
| H6 |
-0.041 |
1.205 |
-0.889 |
| H7 |
1.668 |
-0.612 |
-0.930 |
| H8 |
1.668 |
-0.612 |
0.930 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| O1 | | 1.4829 | 2.4003 | 0.9992 | 2.1399 | 2.1399 | 2.9437 | 2.9437 |
C2 | 1.4829 | | 1.4925 | 2.0058 | 1.1005 | 1.1005 | 2.2397 | 2.2397 | C3 | 2.4003 | 1.4925 | | 3.2507 | 2.1328 | 2.1328 | 1.0836 | 1.0836 | H4 | 0.9992 | 2.0058 | 3.2507 | | 2.3737 | 2.3737 | 3.8155 | 3.8155 | H5 | 2.1399 | 1.1005 | 2.1328 | 2.3737 | | 1.7783 | 3.0874 | 2.4950 | H6 | 2.1399 | 1.1005 | 2.1328 | 2.3737 | 1.7783 | | 2.4950 | 3.0874 | H7 | 2.9437 | 2.2397 | 1.0836 | 3.8155 | 3.0874 | 2.4950 | | 1.8597 | H8 | 2.9437 | 2.2397 | 1.0836 | 3.8155 | 2.4950 | 3.0874 | 1.8597 | |
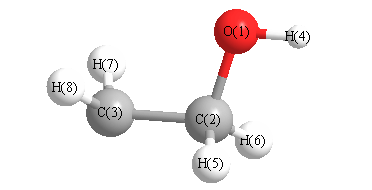 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
107.553 |
|
O1 |
C2 |
H5 |
110.990 |
| O1 |
C2 |
H6 |
110.990 |
|
C2 |
O1 |
H4 |
106.176 |
| C2 |
C3 |
H7 |
119.946 |
|
C2 |
C3 |
H8 |
119.946 |
| C3 |
C2 |
H5 |
109.758 |
|
C3 |
C2 |
H6 |
109.758 |
| H5 |
C2 |
H6 |
107.794 |
|
H7 |
C3 |
H8 |
118.212 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.533 |
|
|
|
| 2 |
C |
-0.210 |
|
|
|
| 3 |
C |
-0.327 |
|
|
|
| 4 |
H |
0.325 |
|
|
|
| 5 |
H |
0.179 |
|
|
|
| 6 |
H |
0.179 |
|
|
|
| 7 |
H |
0.194 |
|
|
|
| 8 |
H |
0.194 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.176 |
1.618 |
0.000 |
1.628 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.292 |
-2.842 |
0.000 |
| y |
-2.842 |
-19.946 |
0.000 |
| z |
0.000 |
0.000 |
-18.615 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.988 |
-2.842 |
0.000 |
| y |
-2.842 |
-2.492 |
0.000 |
| z |
0.000 |
0.000 |
-0.496 |
|
| Polar |
|---|
| 3z2-r2 | -0.992 |
|---|
| x2-y2 | 3.653 |
|---|
| xy | -2.842 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.156 |
-0.532 |
0.000 |
| y |
-0.532 |
2.881 |
0.000 |
| z |
0.000 |
0.000 |
3.062 |
<r2> (average value of r
2) Å
2
| <r2> |
51.086 |
| (<r2>)1/2 |
7.147 |
Jump to
S1C1
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -153.505537 |
| Energy at 298.15K | -153.510138 |
| HF Energy | -153.505537 |
| Nuclear repulsion energy | 74.057657 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3472 |
3350 |
1.01 |
|
|
|
| 2 |
A |
3294 |
3179 |
5.69 |
|
|
|
| 3 |
A |
3177 |
3065 |
4.98 |
|
|
|
| 4 |
A |
2941 |
2838 |
65.14 |
|
|
|
| 5 |
A |
2882 |
2781 |
69.91 |
|
|
|
| 6 |
A |
1561 |
1506 |
4.26 |
|
|
|
| 7 |
A |
1474 |
1423 |
4.63 |
|
|
|
| 8 |
A |
1455 |
1404 |
0.95 |
|
|
|
| 9 |
A |
1281 |
1236 |
54.77 |
|
|
|
| 10 |
A |
1213 |
1171 |
3.05 |
|
|
|
| 11 |
A |
1086 |
1048 |
5.32 |
|
|
|
| 12 |
A |
1039 |
1003 |
47.75 |
|
|
|
| 13 |
A |
969 |
935 |
5.03 |
|
|
|
| 14 |
A |
861 |
831 |
21.23 |
|
|
|
| 15 |
A |
429 |
414 |
34.36 |
|
|
|
| 16 |
A |
394 |
380 |
57.92 |
|
|
|
| 17 |
A |
293 |
282 |
100.63 |
|
|
|
| 18 |
A |
132 |
128 |
24.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13976.3 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 13485.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.125 |
-0.378 |
-0.056 |
| C2 |
0.004 |
0.536 |
0.032 |
| C3 |
1.251 |
-0.270 |
-0.015 |
| H4 |
-1.957 |
0.132 |
0.107 |
| H5 |
-0.015 |
1.127 |
0.974 |
| H6 |
0.023 |
1.283 |
-0.785 |
| H7 |
2.205 |
0.202 |
-0.219 |
| H8 |
1.212 |
-1.312 |
0.273 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| O1 | | 1.4559 | 2.3793 | 0.9899 | 2.1354 | 2.1470 | 3.3837 | 2.5385 |
C2 | 1.4559 | | 1.4858 | 2.0041 | 1.1121 | 1.1068 | 2.2396 | 2.2212 | C3 | 2.3793 | 1.4858 | | 3.2361 | 2.1293 | 2.1242 | 1.0829 | 1.0818 | H4 | 0.9899 | 2.0041 | 3.2361 | | 2.3482 | 2.4582 | 4.1751 | 3.4871 | H5 | 2.1354 | 1.1121 | 2.1293 | 2.3482 | | 1.7661 | 2.6841 | 2.8188 | H6 | 2.1470 | 1.1068 | 2.1242 | 2.4582 | 1.7661 | | 2.4993 | 3.0442 | H7 | 3.3837 | 2.2396 | 1.0829 | 4.1751 | 2.6841 | 2.4993 | | 1.8758 | H8 | 2.5385 | 2.2212 | 1.0818 | 3.4871 | 2.8188 | 3.0442 | 1.8758 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
107.951 |
|
O1 |
C2 |
H5 |
111.815 |
| O1 |
C2 |
H6 |
113.110 |
|
C2 |
O1 |
H4 |
108.549 |
| C2 |
C3 |
H7 |
120.544 |
|
C2 |
C3 |
H8 |
118.942 |
| C3 |
C2 |
H5 |
109.246 |
|
C3 |
C2 |
H6 |
109.161 |
| H5 |
C2 |
H6 |
105.489 |
|
H7 |
C3 |
H8 |
120.109 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.549 |
|
|
|
| 2 |
C |
-0.189 |
|
|
|
| 3 |
C |
-0.347 |
|
|
|
| 4 |
H |
0.332 |
|
|
|
| 5 |
H |
0.182 |
|
|
|
| 6 |
H |
0.182 |
|
|
|
| 7 |
H |
0.180 |
|
|
|
| 8 |
H |
0.209 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.280 |
1.800 |
0.397 |
1.864 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-15.627 |
-1.948 |
-0.459 |
| y |
-1.948 |
-18.935 |
-0.378 |
| z |
-0.459 |
-0.378 |
-20.154 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.918 |
-1.948 |
-0.459 |
| y |
-1.948 |
-1.044 |
-0.378 |
| z |
-0.459 |
-0.378 |
-2.873 |
|
| Polar |
|---|
| 3z2-r2 | -5.747 |
|---|
| x2-y2 | 3.308 |
|---|
| xy | -1.948 |
|---|
| xz | -0.459 |
|---|
| yz | -0.378 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.283 |
-0.196 |
-0.183 |
| y |
-0.196 |
3.491 |
-0.108 |
| z |
-0.183 |
-0.108 |
2.259 |
<r2> (average value of r
2) Å
2
| <r2> |
50.802 |
| (<r2>)1/2 |
7.128 |
