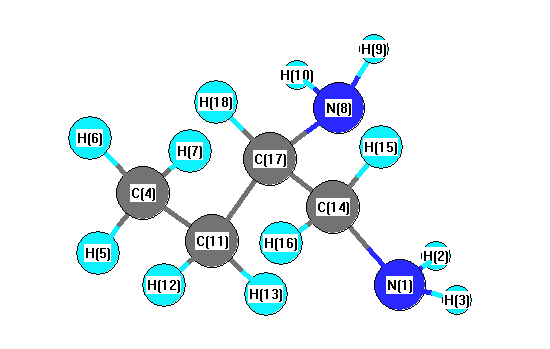
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3498 |
3376 |
0.25 |
|
|
|
| 2 |
A |
3495 |
3373 |
1.89 |
|
|
|
| 3 |
A |
3397 |
3278 |
3.78 |
|
|
|
| 4 |
A |
3382 |
3263 |
17.97 |
|
|
|
| 5 |
A |
3142 |
3032 |
19.65 |
|
|
|
| 6 |
A |
3107 |
2998 |
36.05 |
|
|
|
| 7 |
A |
3094 |
2986 |
30.07 |
|
|
|
| 8 |
A |
3065 |
2958 |
48.56 |
|
|
|
| 9 |
A |
3057 |
2949 |
2.87 |
|
|
|
| 10 |
A |
3041 |
2935 |
24.64 |
|
|
|
| 11 |
A |
2948 |
2844 |
89.91 |
|
|
|
| 12 |
A |
2895 |
2794 |
110.32 |
|
|
|
| 13 |
A |
1711 |
1651 |
29.09 |
|
|
|
| 14 |
A |
1676 |
1617 |
48.62 |
|
|
|
| 15 |
A |
1578 |
1523 |
2.38 |
|
|
|
| 16 |
A |
1576 |
1521 |
9.74 |
|
|
|
| 17 |
A |
1565 |
1511 |
5.11 |
|
|
|
| 18 |
A |
1548 |
1493 |
4.79 |
|
|
|
| 19 |
A |
1463 |
1411 |
8.75 |
|
|
|
| 20 |
A |
1446 |
1395 |
8.38 |
|
|
|
| 21 |
A |
1424 |
1374 |
6.20 |
|
|
|
| 22 |
A |
1404 |
1355 |
7.06 |
|
|
|
| 23 |
A |
1371 |
1323 |
2.08 |
|
|
|
| 24 |
A |
1337 |
1291 |
1.70 |
|
|
|
| 25 |
A |
1290 |
1245 |
3.08 |
|
|
|
| 26 |
A |
1278 |
1233 |
3.81 |
|
|
|
| 27 |
A |
1206 |
1164 |
6.62 |
|
|
|
| 28 |
A |
1153 |
1113 |
8.96 |
|
|
|
| 29 |
A |
1109 |
1070 |
0.17 |
|
|
|
| 30 |
A |
1084 |
1046 |
7.28 |
|
|
|
| 31 |
A |
1048 |
1011 |
6.07 |
|
|
|
| 32 |
A |
1019 |
984 |
2.34 |
|
|
|
| 33 |
A |
970 |
936 |
5.90 |
|
|
|
| 34 |
A |
905 |
873 |
1.92 |
|
|
|
| 35 |
A |
870 |
839 |
21.56 |
|
|
|
| 36 |
A |
804 |
776 |
121.57 |
|
|
|
| 37 |
A |
786 |
758 |
26.64 |
|
|
|
| 38 |
A |
742 |
716 |
153.95 |
|
|
|
| 39 |
A |
611 |
589 |
27.90 |
|
|
|
| 40 |
A |
448 |
432 |
8.53 |
|
|
|
| 41 |
A |
396 |
382 |
39.44 |
|
|
|
| 42 |
A |
382 |
369 |
54.66 |
|
|
|
| 43 |
A |
314 |
303 |
16.31 |
|
|
|
| 44 |
A |
267 |
258 |
4.09 |
|
|
|
| 45 |
A |
238 |
229 |
20.15 |
|
|
|
| 46 |
A |
218 |
210 |
6.13 |
|
|
|
| 47 |
A |
163 |
157 |
11.59 |
|
|
|
| 48 |
A |
83 |
80 |
2.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36801.6 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 35509.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.667 |
|
|
-0.947 |
| 2 |
H |
0.282 |
|
|
0.362 |
| 3 |
H |
0.253 |
|
|
0.381 |
| 4 |
C |
-0.569 |
|
|
-0.295 |
| 5 |
H |
0.193 |
|
|
0.082 |
| 6 |
H |
0.181 |
|
|
0.071 |
| 7 |
H |
0.175 |
|
|
0.069 |
| 8 |
N |
-0.661 |
|
|
-0.929 |
| 9 |
H |
0.261 |
|
|
0.366 |
| 10 |
H |
0.273 |
|
|
0.364 |
| 11 |
C |
-0.352 |
|
|
0.075 |
| 12 |
H |
0.184 |
|
|
0.001 |
| 13 |
H |
0.225 |
|
|
0.030 |
| 14 |
C |
-0.182 |
|
|
0.130 |
| 15 |
H |
0.141 |
|
|
0.007 |
| 16 |
H |
0.184 |
|
|
0.031 |
| 17 |
C |
-0.078 |
|
|
0.229 |
| 18 |
H |
0.158 |
|
|
-0.025 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.109 |
1.622 |
1.114 |
1.971 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.075 |
1.648 |
1.091 |
1.978 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.757 |
0.105 |
0.746 |
| y |
0.105 |
-37.764 |
-0.561 |
| z |
0.746 |
-0.561 |
-40.287 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.269 |
0.105 |
0.746 |
| y |
0.105 |
0.258 |
-0.561 |
| z |
0.746 |
-0.561 |
-3.527 |
|
| Polar |
|---|
| 3z2-r2 | -7.053 |
|---|
| x2-y2 | 2.008 |
|---|
| xy | 0.105 |
|---|
| xz | 0.746 |
|---|
| yz | -0.561 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.930 |
-0.077 |
0.169 |
| y |
-0.077 |
7.604 |
-0.123 |
| z |
0.169 |
-0.123 |
7.319 |
<r2> (average value of r
2) Å
2
| <r2> |
204.022 |
| (<r2>)1/2 |
14.284 |