Jump to
S1C2
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -356.469295 |
| Energy at 298.15K | -356.474666 |
| Nuclear repulsion energy | 230.364985 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3662 |
3534 |
65.64 |
|
|
|
| 2 |
A' |
3524 |
3400 |
52.45 |
|
|
|
| 3 |
A' |
3475 |
3353 |
28.88 |
|
|
|
| 4 |
A' |
1785 |
1722 |
145.11 |
|
|
|
| 5 |
A' |
1755 |
1694 |
187.43 |
|
|
|
| 6 |
A' |
1641 |
1584 |
178.81 |
|
|
|
| 7 |
A' |
1413 |
1364 |
9.80 |
|
|
|
| 8 |
A' |
1325 |
1279 |
68.80 |
|
|
|
| 9 |
A' |
1131 |
1091 |
224.38 |
|
|
|
| 10 |
A' |
1112 |
1073 |
47.83 |
|
|
|
| 11 |
A' |
744 |
718 |
5.21 |
|
|
|
| 12 |
A' |
605 |
584 |
63.14 |
|
|
|
| 13 |
A' |
535 |
516 |
2.96 |
|
|
|
| 14 |
A' |
421 |
406 |
10.01 |
|
|
|
| 15 |
A' |
266 |
257 |
13.36 |
|
|
|
| 16 |
A" |
859 |
829 |
1.64 |
|
|
|
| 17 |
A" |
683 |
659 |
423.56 |
|
|
|
| 18 |
A" |
668 |
645 |
31.02 |
|
|
|
| 19 |
A" |
619 |
597 |
34.56 |
|
|
|
| 20 |
A" |
435 |
419 |
42.58 |
|
|
|
| 21 |
A" |
118 |
114 |
6.49 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13388.0 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 12918.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.742 |
0.000 |
| C2 |
-0.053 |
-0.794 |
0.000 |
| O3 |
-1.097 |
-1.448 |
0.000 |
| O4 |
1.043 |
1.392 |
0.000 |
| O5 |
-1.247 |
1.282 |
0.000 |
| N6 |
1.218 |
-1.271 |
0.000 |
| H7 |
1.390 |
-2.268 |
0.000 |
| H8 |
1.985 |
-0.606 |
0.000 |
| H9 |
-1.170 |
2.277 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5367 | 2.4496 | 1.2293 | 1.3593 | 2.3528 | 3.3157 | 2.3997 | 1.9301 |
C2 | 1.5367 | | 1.2320 | 2.4457 | 2.3947 | 1.3583 | 2.0634 | 2.0472 | 3.2675 | O3 | 2.4496 | 1.2320 | | 3.5566 | 2.7343 | 2.3224 | 2.6190 | 3.1954 | 3.7257 | O4 | 1.2293 | 2.4457 | 3.5566 | | 2.2932 | 2.6689 | 3.6771 | 2.2094 | 2.3835 | O5 | 1.3593 | 2.3947 | 2.7343 | 2.2932 | | 3.5492 | 4.4227 | 3.7437 | 0.9979 | N6 | 2.3528 | 1.3583 | 2.3224 | 2.6689 | 3.5492 | | 1.0121 | 1.0149 | 4.2769 | H7 | 3.3157 | 2.0634 | 2.6190 | 3.6771 | 4.4227 | 1.0121 | | 1.7655 | 5.2167 | H8 | 2.3997 | 2.0472 | 3.1954 | 2.2094 | 3.7437 | 1.0149 | 1.7655 | | 4.2742 | H9 | 1.9301 | 3.2675 | 3.7257 | 2.3835 | 0.9979 | 4.2769 | 5.2167 | 4.2742 | |
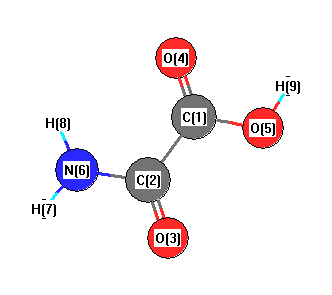 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
124.073 |
|
C1 |
C2 |
N6 |
108.571 |
| C1 |
O5 |
H9 |
108.975 |
|
C2 |
C1 |
O4 |
123.934 |
| C2 |
C1 |
O5 |
111.418 |
|
C2 |
N6 |
H7 |
120.327 |
| C2 |
N6 |
H8 |
118.521 |
|
O3 |
C2 |
N6 |
127.356 |
| O4 |
C1 |
O5 |
124.648 |
|
H7 |
N6 |
H8 |
121.152 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.609 |
|
|
0.627 |
| 2 |
C |
0.599 |
|
|
0.559 |
| 3 |
O |
-0.478 |
|
|
-0.460 |
| 4 |
O |
-0.489 |
|
|
-0.496 |
| 5 |
O |
-0.532 |
|
|
-0.625 |
| 6 |
N |
-0.750 |
|
|
-0.894 |
| 7 |
H |
0.329 |
|
|
0.431 |
| 8 |
H |
0.338 |
|
|
0.388 |
| 9 |
H |
0.374 |
|
|
0.471 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.234 |
0.910 |
0.000 |
2.412 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
2.237 |
0.914 |
0.000 |
2.416 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.757 |
-7.774 |
-0.004 |
| y |
-7.774 |
-38.671 |
0.002 |
| z |
-0.004 |
0.002 |
-32.885 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
12.021 |
-7.774 |
-0.004 |
| y |
-7.774 |
-10.350 |
0.002 |
| z |
-0.004 |
0.002 |
-1.671 |
|
| Polar |
|---|
| 3z2-r2 | -3.342 |
|---|
| x2-y2 | 14.914 |
|---|
| xy | -7.774 |
|---|
| xz | -0.004 |
|---|
| yz | 0.002 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.476 |
-0.341 |
0.000 |
| y |
-0.341 |
5.930 |
0.000 |
| z |
0.000 |
0.000 |
1.634 |
<r2> (average value of r
2) Å
2
| <r2> |
142.796 |
| (<r2>)1/2 |
11.950 |
Jump to
S1C1
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -356.477646 |
| Energy at 298.15K | -356.483253 |
| Nuclear repulsion energy | 232.098550 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3644 |
3516 |
73.27 |
|
|
|
| 2 |
A' |
3509 |
3386 |
63.32 |
|
|
|
| 3 |
A' |
3230 |
3116 |
108.48 |
|
|
|
| 4 |
A' |
1827 |
1763 |
155.82 |
|
|
|
| 5 |
A' |
1756 |
1694 |
168.51 |
|
|
|
| 6 |
A' |
1645 |
1587 |
90.87 |
|
|
|
| 7 |
A' |
1415 |
1365 |
149.77 |
|
|
|
| 8 |
A' |
1345 |
1298 |
419.63 |
|
|
|
| 9 |
A' |
1183 |
1142 |
15.46 |
|
|
|
| 10 |
A' |
1122 |
1083 |
3.72 |
|
|
|
| 11 |
A' |
764 |
737 |
10.92 |
|
|
|
| 12 |
A' |
631 |
609 |
7.59 |
|
|
|
| 13 |
A' |
559 |
539 |
0.89 |
|
|
|
| 14 |
A' |
410 |
396 |
8.21 |
|
|
|
| 15 |
A' |
274 |
265 |
49.72 |
|
|
|
| 16 |
A" |
852 |
822 |
38.10 |
|
|
|
| 17 |
A" |
807 |
779 |
120.35 |
|
|
|
| 18 |
A" |
712 |
687 |
201.73 |
|
|
|
| 19 |
A" |
669 |
646 |
140.96 |
|
|
|
| 20 |
A" |
456 |
440 |
1.56 |
|
|
|
| 21 |
A" |
164 |
159 |
7.31 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13485.9 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 13012.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.028 |
-0.791 |
0.000 |
| C2 |
0.000 |
0.748 |
0.000 |
| O3 |
-1.107 |
1.326 |
0.000 |
| O4 |
1.055 |
-1.456 |
0.000 |
| O5 |
-1.237 |
-1.272 |
0.000 |
| N6 |
1.228 |
1.284 |
0.000 |
| H7 |
1.369 |
2.287 |
0.000 |
| H8 |
2.018 |
0.644 |
0.000 |
| H9 |
-1.834 |
-0.452 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5393 | 2.4013 | 1.2240 | 1.3535 | 2.3968 | 3.3575 | 2.4539 | 1.8916 |
C2 | 1.5393 | | 1.2483 | 2.4438 | 2.3694 | 1.3395 | 2.0596 | 2.0209 | 2.1917 | O3 | 2.4013 | 1.2483 | | 3.5230 | 2.6015 | 2.3347 | 2.6557 | 3.1984 | 1.9212 | O4 | 1.2240 | 2.4438 | 3.5230 | | 2.2997 | 2.7455 | 3.7563 | 2.3105 | 3.0578 | O5 | 1.3535 | 2.3694 | 2.6015 | 2.2997 | | 3.5513 | 4.4117 | 3.7778 | 1.0137 | N6 | 2.3968 | 1.3395 | 2.3347 | 2.7455 | 3.5513 | | 1.0130 | 1.0171 | 3.5194 | H7 | 3.3575 | 2.0596 | 2.6557 | 3.7563 | 4.4117 | 1.0130 | | 1.7667 | 4.2144 | H8 | 2.4539 | 2.0209 | 3.1984 | 2.3105 | 3.7778 | 1.0171 | 1.7667 | | 4.0048 | H9 | 1.8916 | 2.1917 | 1.9212 | 3.0578 | 1.0137 | 3.5194 | 4.2144 | 4.0048 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
118.587 |
|
C1 |
C2 |
N6 |
112.547 |
| C1 |
O5 |
H9 |
105.178 |
|
C2 |
C1 |
O4 |
123.953 |
| C2 |
C1 |
O5 |
109.816 |
|
C2 |
N6 |
H7 |
121.588 |
| C2 |
N6 |
H8 |
117.438 |
|
O3 |
C2 |
N6 |
128.866 |
| O4 |
C1 |
O5 |
126.231 |
|
H7 |
N6 |
H8 |
120.975 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.649 |
|
|
|
| 2 |
C |
0.584 |
|
|
|
| 3 |
O |
-0.509 |
|
|
|
| 4 |
O |
-0.480 |
|
|
|
| 5 |
O |
-0.558 |
|
|
|
| 6 |
N |
-0.754 |
|
|
|
| 7 |
H |
0.338 |
|
|
|
| 8 |
H |
0.349 |
|
|
|
| 9 |
H |
0.382 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.709 |
2.978 |
0.000 |
3.434 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.366 |
6.748 |
0.000 |
| y |
6.748 |
-37.128 |
0.000 |
| z |
0.000 |
0.000 |
-32.845 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.620 |
6.748 |
0.000 |
| y |
6.748 |
-5.522 |
0.000 |
| z |
0.000 |
0.000 |
0.902 |
|
| Polar |
|---|
| 3z2-r2 | 1.805 |
|---|
| x2-y2 | 6.761 |
|---|
| xy | 6.748 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.485 |
0.001 |
0.000 |
| y |
0.001 |
4.770 |
0.000 |
| z |
0.000 |
0.000 |
1.612 |
<r2> (average value of r
2) Å
2
| <r2> |
140.181 |
| (<r2>)1/2 |
11.840 |
