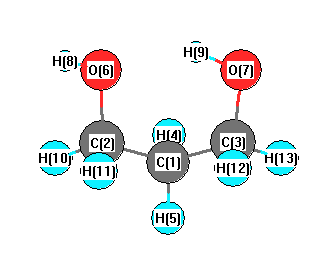
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3505 |
3382 |
2.10 |
|
|
|
| 2 |
A |
3304 |
3188 |
176.21 |
|
|
|
| 3 |
A |
3118 |
3009 |
26.80 |
|
|
|
| 4 |
A |
3081 |
2973 |
62.88 |
|
|
|
| 5 |
A |
3064 |
2957 |
36.59 |
|
|
|
| 6 |
A |
3048 |
2941 |
38.15 |
|
|
|
| 7 |
A |
3010 |
2904 |
45.57 |
|
|
|
| 8 |
A |
2933 |
2830 |
92.39 |
|
|
|
| 9 |
A |
1590 |
1535 |
9.95 |
|
|
|
| 10 |
A |
1574 |
1519 |
1.82 |
|
|
|
| 11 |
A |
1544 |
1490 |
109.28 |
|
|
|
| 12 |
A |
1521 |
1468 |
6.85 |
|
|
|
| 13 |
A |
1453 |
1402 |
6.09 |
|
|
|
| 14 |
A |
1423 |
1373 |
5.02 |
|
|
|
| 15 |
A |
1409 |
1360 |
1.81 |
|
|
|
| 16 |
A |
1327 |
1281 |
21.10 |
|
|
|
| 17 |
A |
1299 |
1254 |
2.98 |
|
|
|
| 18 |
A |
1230 |
1186 |
31.12 |
|
|
|
| 19 |
A |
1216 |
1173 |
22.15 |
|
|
|
| 20 |
A |
1137 |
1097 |
19.25 |
|
|
|
| 21 |
A |
1058 |
1021 |
74.29 |
|
|
|
| 22 |
A |
1046 |
1010 |
8.18 |
|
|
|
| 23 |
A |
962 |
928 |
3.08 |
|
|
|
| 24 |
A |
923 |
891 |
4.48 |
|
|
|
| 25 |
A |
900 |
869 |
31.24 |
|
|
|
| 26 |
A |
814 |
786 |
5.52 |
|
|
|
| 27 |
A |
733 |
708 |
155.07 |
|
|
|
| 28 |
A |
512 |
494 |
3.73 |
|
|
|
| 29 |
A |
381 |
368 |
2.11 |
|
|
|
| 30 |
A |
348 |
336 |
9.05 |
|
|
|
| 31 |
A |
291 |
280 |
95.99 |
|
|
|
| 32 |
A |
278 |
269 |
15.09 |
|
|
|
| 33 |
A |
111 |
107 |
0.35 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25072.8 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 24192.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.439 |
|
|
|
| 2 |
C |
-0.145 |
|
|
|
| 3 |
C |
-0.109 |
|
|
|
| 4 |
H |
0.220 |
|
|
|
| 5 |
H |
0.187 |
|
|
|
| 6 |
O |
-0.546 |
|
|
|
| 7 |
O |
-0.586 |
|
|
|
| 8 |
H |
0.354 |
|
|
|
| 9 |
H |
0.350 |
|
|
|
| 10 |
H |
0.191 |
|
|
|
| 11 |
H |
0.183 |
|
|
|
| 12 |
H |
0.147 |
|
|
|
| 13 |
H |
0.192 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.598 |
1.516 |
1.342 |
4.129 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.360 |
-5.019 |
1.579 |
| y |
-5.019 |
-30.290 |
-1.508 |
| z |
1.579 |
-1.508 |
-31.445 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.507 |
-5.019 |
1.579 |
| y |
-5.019 |
0.613 |
-1.508 |
| z |
1.579 |
-1.508 |
-1.120 |
|
| Polar |
|---|
| 3z2-r2 | -2.240 |
|---|
| x2-y2 | -0.070 |
|---|
| xy | -5.019 |
|---|
| xz | 1.579 |
|---|
| yz | -1.508 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.096 |
-0.313 |
0.201 |
| y |
-0.313 |
5.817 |
0.048 |
| z |
0.201 |
0.048 |
4.745 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |