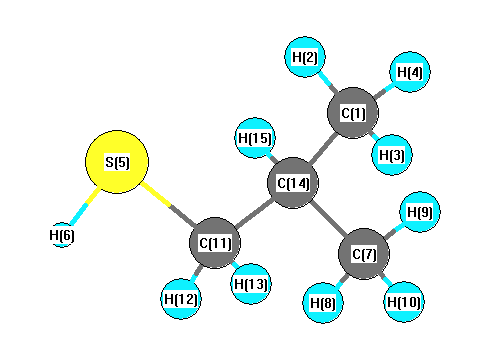
Vibrational Frequencies calculated at BLYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3072 |
3055 |
23.21 |
|
|
|
| 2 |
A |
3046 |
3029 |
23.71 |
|
|
|
| 3 |
A |
3036 |
3019 |
59.71 |
|
|
|
| 4 |
A |
3031 |
3014 |
23.09 |
|
|
|
| 5 |
A |
3026 |
3010 |
4.48 |
|
|
|
| 6 |
A |
3010 |
2993 |
17.06 |
|
|
|
| 7 |
A |
2978 |
2962 |
8.11 |
|
|
|
| 8 |
A |
2972 |
2955 |
12.13 |
|
|
|
| 9 |
A |
2967 |
2951 |
22.03 |
|
|
|
| 10 |
A |
2371 |
2358 |
48.61 |
|
|
|
| 11 |
A |
1539 |
1530 |
14.57 |
|
|
|
| 12 |
A |
1536 |
1527 |
4.64 |
|
|
|
| 13 |
A |
1525 |
1516 |
4.55 |
|
|
|
| 14 |
A |
1519 |
1510 |
0.06 |
|
|
|
| 15 |
A |
1490 |
1482 |
3.98 |
|
|
|
| 16 |
A |
1432 |
1424 |
7.78 |
|
|
|
| 17 |
A |
1415 |
1407 |
7.32 |
|
|
|
| 18 |
A |
1364 |
1357 |
3.03 |
|
|
|
| 19 |
A |
1340 |
1333 |
1.06 |
|
|
|
| 20 |
A |
1257 |
1250 |
33.50 |
|
|
|
| 21 |
A |
1234 |
1228 |
2.77 |
|
|
|
| 22 |
A |
1179 |
1173 |
7.40 |
|
|
|
| 23 |
A |
1114 |
1108 |
5.23 |
|
|
|
| 24 |
A |
1063 |
1057 |
3.08 |
|
|
|
| 25 |
A |
963 |
958 |
1.46 |
|
|
|
| 26 |
A |
948 |
943 |
0.52 |
|
|
|
| 27 |
A |
924 |
919 |
5.03 |
|
|
|
| 28 |
A |
857 |
852 |
2.56 |
|
|
|
| 29 |
A |
849 |
844 |
4.25 |
|
|
|
| 30 |
A |
745 |
741 |
5.85 |
|
|
|
| 31 |
A |
655 |
652 |
6.83 |
|
|
|
| 32 |
A |
414 |
412 |
0.36 |
|
|
|
| 33 |
A |
384 |
382 |
1.82 |
|
|
|
| 34 |
A |
319 |
318 |
0.73 |
|
|
|
| 35 |
A |
254 |
252 |
0.17 |
|
|
|
| 36 |
A |
230 |
229 |
0.07 |
|
|
|
| 37 |
A |
200 |
199 |
3.25 |
|
|
|
| 38 |
A |
136 |
135 |
8.03 |
|
|
|
| 39 |
A |
69 |
69 |
20.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28231.7 cm
-1
Scaled (by 0.9945) Zero Point Vibrational Energy (zpe) 28076.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.504 |
|
|
|
| 2 |
H |
0.203 |
|
|
|
| 3 |
H |
0.173 |
|
|
|
| 4 |
H |
0.175 |
|
|
|
| 5 |
S |
-0.020 |
|
|
|
| 6 |
H |
0.064 |
|
|
|
| 7 |
C |
-0.495 |
|
|
|
| 8 |
H |
0.174 |
|
|
|
| 9 |
H |
0.181 |
|
|
|
| 10 |
H |
0.175 |
|
|
|
| 11 |
C |
-0.519 |
|
|
|
| 12 |
H |
0.210 |
|
|
|
| 13 |
H |
0.209 |
|
|
|
| 14 |
C |
-0.220 |
|
|
|
| 15 |
H |
0.194 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.300 |
-1.349 |
0.691 |
1.996 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.510 |
2.935 |
-1.734 |
| y |
2.935 |
-40.239 |
-0.956 |
| z |
-1.734 |
-0.956 |
-41.908 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.436 |
2.935 |
-1.734 |
| y |
2.935 |
1.470 |
-0.956 |
| z |
-1.734 |
-0.956 |
-1.034 |
|
| Polar |
|---|
| 3z2-r2 | -2.068 |
|---|
| x2-y2 | -1.270 |
|---|
| xy | 2.935 |
|---|
| xz | -1.734 |
|---|
| yz | -0.956 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
210.006 |
| (<r2>)1/2 |
14.492 |