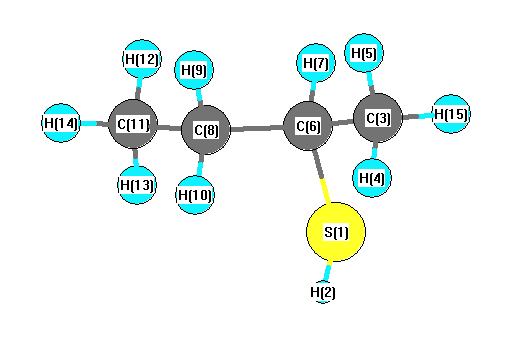
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3139 |
3028 |
15.82 |
|
|
|
| 2 |
A |
3135 |
3025 |
17.29 |
|
|
|
| 3 |
A |
3116 |
3007 |
43.12 |
|
|
|
| 4 |
A |
3110 |
3001 |
12.00 |
|
|
|
| 5 |
A |
3092 |
2984 |
3.44 |
|
|
|
| 6 |
A |
3065 |
2958 |
10.28 |
|
|
|
| 7 |
A |
3053 |
2945 |
17.81 |
|
|
|
| 8 |
A |
3044 |
2937 |
29.70 |
|
|
|
| 9 |
A |
3031 |
2925 |
12.90 |
|
|
|
| 10 |
A |
2455 |
2369 |
32.85 |
|
|
|
| 11 |
A |
1576 |
1521 |
6.60 |
|
|
|
| 12 |
A |
1563 |
1508 |
11.11 |
|
|
|
| 13 |
A |
1562 |
1507 |
2.45 |
|
|
|
| 14 |
A |
1558 |
1503 |
11.35 |
|
|
|
| 15 |
A |
1541 |
1487 |
2.81 |
|
|
|
| 16 |
A |
1465 |
1414 |
6.93 |
|
|
|
| 17 |
A |
1461 |
1410 |
13.50 |
|
|
|
| 18 |
A |
1399 |
1350 |
1.66 |
|
|
|
| 19 |
A |
1362 |
1315 |
8.50 |
|
|
|
| 20 |
A |
1347 |
1299 |
3.09 |
|
|
|
| 21 |
A |
1292 |
1247 |
7.44 |
|
|
|
| 22 |
A |
1205 |
1163 |
2.18 |
|
|
|
| 23 |
A |
1144 |
1104 |
9.06 |
|
|
|
| 24 |
A |
1125 |
1086 |
5.86 |
|
|
|
| 25 |
A |
1039 |
1003 |
11.09 |
|
|
|
| 26 |
A |
1031 |
995 |
1.92 |
|
|
|
| 27 |
A |
991 |
956 |
5.62 |
|
|
|
| 28 |
A |
879 |
849 |
4.97 |
|
|
|
| 29 |
A |
843 |
814 |
8.38 |
|
|
|
| 30 |
A |
813 |
785 |
12.91 |
|
|
|
| 31 |
A |
572 |
552 |
11.01 |
|
|
|
| 32 |
A |
460 |
444 |
0.37 |
|
|
|
| 33 |
A |
375 |
362 |
0.75 |
|
|
|
| 34 |
A |
327 |
316 |
1.35 |
|
|
|
| 35 |
A |
263 |
254 |
0.28 |
|
|
|
| 36 |
A |
245 |
237 |
11.00 |
|
|
|
| 37 |
A |
229 |
221 |
9.23 |
|
|
|
| 38 |
A |
222 |
214 |
7.84 |
|
|
|
| 39 |
A |
122 |
118 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29126.6 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 28104.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.005 |
|
|
-0.373 |
| 2 |
H |
0.068 |
|
|
0.212 |
| 3 |
C |
-0.530 |
|
|
-0.599 |
| 4 |
H |
0.192 |
|
|
0.173 |
| 5 |
H |
0.196 |
|
|
0.172 |
| 6 |
C |
-0.447 |
|
|
0.072 |
| 7 |
H |
0.236 |
|
|
0.176 |
| 8 |
C |
-0.357 |
|
|
-0.148 |
| 9 |
H |
0.198 |
|
|
0.071 |
| 10 |
H |
0.190 |
|
|
0.091 |
| 11 |
C |
-0.557 |
|
|
-0.196 |
| 12 |
H |
0.185 |
|
|
0.058 |
| 13 |
H |
0.220 |
|
|
0.055 |
| 14 |
H |
0.187 |
|
|
0.071 |
| 15 |
H |
0.213 |
|
|
0.166 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.752 |
1.918 |
0.680 |
2.169 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.731 |
1.932 |
0.656 |
2.167 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.534 |
1.290 |
1.379 |
| y |
1.290 |
-43.216 |
-1.604 |
| z |
1.379 |
-1.604 |
-38.964 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.444 |
1.290 |
1.379 |
| y |
1.290 |
-2.467 |
-1.604 |
| z |
1.379 |
-1.604 |
3.911 |
|
| Polar |
|---|
| 3z2-r2 | 7.821 |
|---|
| x2-y2 | 0.682 |
|---|
| xy | 1.290 |
|---|
| xz | 1.379 |
|---|
| yz | -1.604 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.312 |
-0.673 |
0.286 |
| y |
-0.673 |
8.973 |
-0.204 |
| z |
0.286 |
-0.204 |
7.550 |
<r2> (average value of r
2) Å
2
| <r2> |
191.550 |
| (<r2>)1/2 |
13.840 |