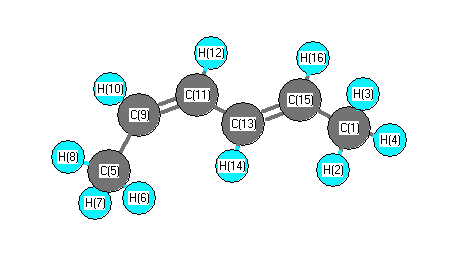
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3172 |
3061 |
59.61 |
|
|
|
| 2 |
A' |
3163 |
3052 |
4.25 |
|
|
|
| 3 |
A' |
3143 |
3033 |
15.20 |
|
|
|
| 4 |
A' |
3140 |
3030 |
5.54 |
|
|
|
| 5 |
A' |
3130 |
3020 |
2.70 |
|
|
|
| 6 |
A' |
3111 |
3001 |
8.35 |
|
|
|
| 7 |
A' |
3036 |
2929 |
25.85 |
|
|
|
| 8 |
A' |
3030 |
2924 |
34.25 |
|
|
|
| 9 |
A' |
1720 |
1660 |
1.67 |
|
|
|
| 10 |
A' |
1687 |
1628 |
3.18 |
|
|
|
| 11 |
A' |
1558 |
1503 |
9.47 |
|
|
|
| 12 |
A' |
1554 |
1500 |
14.30 |
|
|
|
| 13 |
A' |
1476 |
1424 |
3.42 |
|
|
|
| 14 |
A' |
1466 |
1414 |
11.64 |
|
|
|
| 15 |
A' |
1465 |
1413 |
0.32 |
|
|
|
| 16 |
A' |
1380 |
1331 |
0.82 |
|
|
|
| 17 |
A' |
1360 |
1312 |
1.13 |
|
|
|
| 18 |
A' |
1320 |
1274 |
0.91 |
|
|
|
| 19 |
A' |
1195 |
1153 |
2.10 |
|
|
|
| 20 |
A' |
1115 |
1076 |
1.41 |
|
|
|
| 21 |
A' |
1061 |
1023 |
2.24 |
|
|
|
| 22 |
A' |
956 |
923 |
22.37 |
|
|
|
| 23 |
A' |
915 |
883 |
10.29 |
|
|
|
| 24 |
A' |
619 |
597 |
5.75 |
|
|
|
| 25 |
A' |
476 |
460 |
0.14 |
|
|
|
| 26 |
A' |
332 |
320 |
1.89 |
|
|
|
| 27 |
A' |
157 |
151 |
0.48 |
|
|
|
| 28 |
A" |
3072 |
2964 |
22.56 |
|
|
|
| 29 |
A" |
3070 |
2962 |
20.35 |
|
|
|
| 30 |
A" |
1553 |
1499 |
9.26 |
|
|
|
| 31 |
A" |
1549 |
1495 |
7.22 |
|
|
|
| 32 |
A" |
1103 |
1065 |
1.09 |
|
|
|
| 33 |
A" |
1097 |
1059 |
0.00 |
|
|
|
| 34 |
A" |
1040 |
1004 |
19.71 |
|
|
|
| 35 |
A" |
995 |
960 |
21.98 |
|
|
|
| 36 |
A" |
865 |
835 |
14.70 |
|
|
|
| 37 |
A" |
735 |
710 |
21.80 |
|
|
|
| 38 |
A" |
447 |
431 |
5.81 |
|
|
|
| 39 |
A" |
212 |
204 |
0.01 |
|
|
|
| 40 |
A" |
199 |
192 |
1.36 |
|
|
|
| 41 |
A" |
133 |
128 |
0.00 |
|
|
|
| 42 |
A" |
107 |
103 |
1.23 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31455.9 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 30351.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.601 |
|
|
-0.341 |
| 2 |
H |
0.196 |
|
|
0.102 |
| 3 |
H |
0.203 |
|
|
0.117 |
| 4 |
H |
0.203 |
|
|
0.117 |
| 5 |
C |
-0.609 |
|
|
-0.227 |
| 6 |
H |
0.199 |
|
|
0.077 |
| 7 |
H |
0.203 |
|
|
0.087 |
| 8 |
H |
0.203 |
|
|
0.087 |
| 9 |
C |
-0.168 |
|
|
-0.066 |
| 10 |
H |
0.180 |
|
|
0.105 |
| 11 |
C |
-0.185 |
|
|
-0.276 |
| 12 |
H |
0.177 |
|
|
0.150 |
| 13 |
C |
-0.194 |
|
|
-0.152 |
| 14 |
H |
0.177 |
|
|
0.186 |
| 15 |
C |
-0.162 |
|
|
-0.103 |
| 16 |
H |
0.177 |
|
|
0.138 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.233 |
0.123 |
0.000 |
0.264 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.249 |
0.158 |
0.000 |
0.295 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.013 |
-0.572 |
0.000 |
| y |
-0.572 |
-33.924 |
0.000 |
| z |
0.000 |
0.000 |
-41.338 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.617 |
-0.572 |
0.000 |
| y |
-0.572 |
4.752 |
0.000 |
| z |
0.000 |
0.000 |
-6.369 |
|
| Polar |
|---|
| 3z2-r2 | -12.738 |
|---|
| x2-y2 | -2.090 |
|---|
| xy | -0.572 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.206 |
-1.187 |
0.000 |
| y |
-1.187 |
16.628 |
0.000 |
| z |
0.000 |
0.000 |
4.637 |
<r2> (average value of r
2) Å
2
| <r2> |
260.141 |
| (<r2>)1/2 |
16.129 |