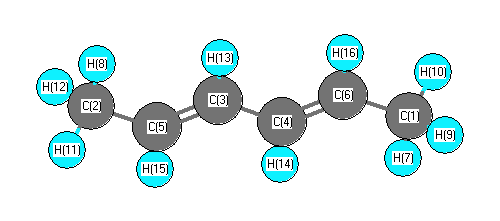
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3145 |
3035 |
0.00 |
|
|
|
| 2 |
Ag |
3129 |
3019 |
0.00 |
|
|
|
| 3 |
Ag |
3111 |
3002 |
0.00 |
|
|
|
| 4 |
Ag |
3031 |
2924 |
0.00 |
|
|
|
| 5 |
Ag |
1725 |
1664 |
0.00 |
|
|
|
| 6 |
Ag |
1559 |
1504 |
0.00 |
|
|
|
| 7 |
Ag |
1463 |
1411 |
0.00 |
|
|
|
| 8 |
Ag |
1401 |
1352 |
0.00 |
|
|
|
| 9 |
Ag |
1364 |
1316 |
0.00 |
|
|
|
| 10 |
Ag |
1207 |
1165 |
0.00 |
|
|
|
| 11 |
Ag |
1098 |
1059 |
0.00 |
|
|
|
| 12 |
Ag |
962 |
928 |
0.00 |
|
|
|
| 13 |
Ag |
444 |
428 |
0.00 |
|
|
|
| 14 |
Ag |
340 |
328 |
0.00 |
|
|
|
| 15 |
Au |
3070 |
2962 |
45.09 |
|
|
|
| 16 |
Au |
1548 |
1494 |
17.16 |
|
|
|
| 17 |
Au |
1101 |
1062 |
2.89 |
|
|
|
| 18 |
Au |
1040 |
1004 |
76.02 |
|
|
|
| 19 |
Au |
782 |
754 |
6.84 |
|
|
|
| 20 |
Au |
243 |
235 |
1.83 |
|
|
|
| 21 |
Au |
161 |
156 |
0.14 |
|
|
|
| 22 |
Au |
103 |
100 |
2.00 |
|
|
|
| 23 |
Bg |
3070 |
2962 |
0.00 |
|
|
|
| 24 |
Bg |
1548 |
1494 |
0.00 |
|
|
|
| 25 |
Bg |
1103 |
1064 |
0.00 |
|
|
|
| 26 |
Bg |
982 |
948 |
0.00 |
|
|
|
| 27 |
Bg |
885 |
854 |
0.00 |
|
|
|
| 28 |
Bg |
340 |
328 |
0.00 |
|
|
|
| 29 |
Bg |
187 |
180 |
0.00 |
|
|
|
| 30 |
Bu |
3151 |
3040 |
82.20 |
|
|
|
| 31 |
Bu |
3135 |
3025 |
3.18 |
|
|
|
| 32 |
Bu |
3111 |
3002 |
16.66 |
|
|
|
| 33 |
Bu |
3030 |
2924 |
69.66 |
|
|
|
| 34 |
Bu |
1698 |
1638 |
4.13 |
|
|
|
| 35 |
Bu |
1554 |
1500 |
21.83 |
|
|
|
| 36 |
Bu |
1462 |
1411 |
6.83 |
|
|
|
| 37 |
Bu |
1380 |
1332 |
2.86 |
|
|
|
| 38 |
Bu |
1317 |
1271 |
1.15 |
|
|
|
| 39 |
Bu |
1113 |
1074 |
9.51 |
|
|
|
| 40 |
Bu |
944 |
911 |
24.68 |
|
|
|
| 41 |
Bu |
562 |
542 |
0.08 |
|
|
|
| 42 |
Bu |
142 |
137 |
1.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31369.0 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 30268.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.601 |
|
|
-0.371 |
| 2 |
C |
-0.601 |
|
|
-0.376 |
| 3 |
C |
-0.181 |
|
|
-0.266 |
| 4 |
C |
-0.181 |
|
|
-0.280 |
| 5 |
C |
-0.167 |
|
|
-0.002 |
| 6 |
C |
-0.167 |
|
|
0.002 |
| 7 |
H |
0.196 |
|
|
0.117 |
| 8 |
H |
0.196 |
|
|
0.118 |
| 9 |
H |
0.202 |
|
|
0.117 |
| 10 |
H |
0.202 |
|
|
0.117 |
| 11 |
H |
0.202 |
|
|
0.118 |
| 12 |
H |
0.202 |
|
|
0.118 |
| 13 |
H |
0.173 |
|
|
0.182 |
| 14 |
H |
0.173 |
|
|
0.186 |
| 15 |
H |
0.177 |
|
|
0.109 |
| 16 |
H |
0.177 |
|
|
0.109 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.002 |
-0.001 |
0.000 |
0.002 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.195 |
-1.016 |
0.000 |
| y |
-1.016 |
-34.446 |
0.000 |
| z |
0.000 |
0.000 |
-41.348 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.702 |
-1.016 |
0.000 |
| y |
-1.016 |
3.826 |
0.000 |
| z |
0.000 |
0.000 |
-6.528 |
|
| Polar |
|---|
| 3z2-r2 | -13.055 |
|---|
| x2-y2 | -0.749 |
|---|
| xy | -1.016 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.840 |
-4.393 |
0.000 |
| y |
-4.393 |
14.564 |
0.000 |
| z |
0.000 |
0.000 |
4.654 |
<r2> (average value of r
2) Å
2
| <r2> |
291.509 |
| (<r2>)1/2 |
17.074 |