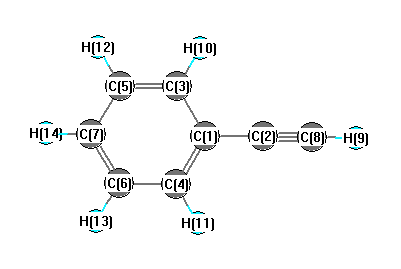
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3497 |
3374 |
78.25 |
|
|
|
| 2 |
A |
3227 |
3114 |
4.89 |
|
|
|
| 3 |
A |
3213 |
3100 |
13.52 |
|
|
|
| 4 |
A |
3194 |
3082 |
0.24 |
|
|
|
| 5 |
A |
2233 |
2154 |
0.74 |
|
|
|
| 6 |
A |
1631 |
1574 |
1.34 |
|
|
|
| 7 |
A |
1544 |
1489 |
19.38 |
|
|
|
| 8 |
A |
1242 |
1198 |
0.38 |
|
|
|
| 9 |
A |
1226 |
1183 |
1.09 |
|
|
|
| 10 |
A |
1063 |
1025 |
2.60 |
|
|
|
| 11 |
A |
1034 |
998 |
0.48 |
|
|
|
| 12 |
A |
786 |
759 |
3.25 |
|
|
|
| 13 |
A |
487 |
470 |
0.08 |
|
|
|
| 14 |
A |
1009 |
973 |
0.00 |
|
|
|
| 15 |
A |
870 |
840 |
0.00 |
|
|
|
| 16 |
A |
422 |
407 |
0.00 |
|
|
|
| 17 |
A |
1038 |
1002 |
0.13 |
|
|
|
| 18 |
A |
954 |
920 |
4.39 |
|
|
|
| 19 |
A |
790 |
762 |
39.56 |
|
|
|
| 20 |
A |
727 |
702 |
43.70 |
|
|
|
| 21 |
A |
699 |
674 |
60.56 |
|
|
|
| 22 |
A |
561 |
542 |
4.85 |
|
|
|
| 23 |
A |
375 |
362 |
3.40 |
|
|
|
| 24 |
A |
143 |
138 |
1.83 |
|
|
|
| 25 |
A |
3221 |
3108 |
15.15 |
|
|
|
| 26 |
A |
3202 |
3090 |
4.00 |
|
|
|
| 27 |
A |
1601 |
1545 |
2.06 |
|
|
|
| 28 |
A |
1499 |
1446 |
7.00 |
|
|
|
| 29 |
A |
1391 |
1343 |
0.45 |
|
|
|
| 30 |
A |
1300 |
1254 |
0.23 |
|
|
|
| 31 |
A |
1232 |
1189 |
0.01 |
|
|
|
| 32 |
A |
1115 |
1076 |
3.39 |
|
|
|
| 33 |
A |
741 |
715 |
43.86 |
|
|
|
| 34 |
A |
661 |
638 |
0.30 |
|
|
|
| 35 |
A |
542 |
523 |
4.66 |
|
|
|
| 36 |
A |
161 |
155 |
1.29 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24314.8 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 23461.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.106 |
|
|
|
| 2 |
C |
0.109 |
|
|
|
| 3 |
C |
-0.165 |
|
|
|
| 4 |
C |
-0.165 |
|
|
|
| 5 |
C |
-0.180 |
|
|
|
| 6 |
C |
-0.180 |
|
|
|
| 7 |
C |
-0.183 |
|
|
|
| 8 |
C |
-0.346 |
|
|
|
| 9 |
H |
0.251 |
|
|
|
| 10 |
H |
0.195 |
|
|
|
| 11 |
H |
0.195 |
|
|
|
| 12 |
H |
0.192 |
|
|
|
| 13 |
H |
0.192 |
|
|
|
| 14 |
H |
0.191 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.613 |
0.613 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-51.110 |
0.000 |
0.000 |
| y |
0.000 |
-41.559 |
0.000 |
| z |
0.000 |
0.000 |
-37.340 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-11.660 |
0.000 |
0.000 |
| y |
0.000 |
2.666 |
0.000 |
| z |
0.000 |
0.000 |
8.994 |
|
| Polar |
|---|
| 3z2-r2 | 17.988 |
|---|
| x2-y2 | -9.551 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.878 |
0.000 |
0.000 |
| y |
0.000 |
10.433 |
0.000 |
| z |
0.000 |
0.000 |
18.348 |
<r2> (average value of r
2) Å
2
| <r2> |
265.225 |
| (<r2>)1/2 |
16.286 |