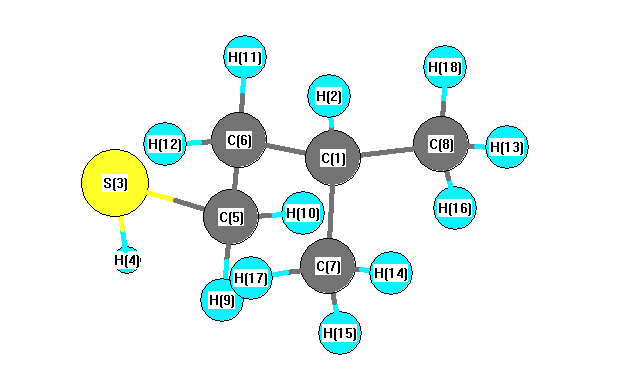
Vibrational Frequencies calculated at BLYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3038 |
3021 |
63.89 |
|
|
|
| 2 |
A' |
3030 |
3013 |
68.59 |
|
|
|
| 3 |
A' |
3027 |
3011 |
9.53 |
|
|
|
| 4 |
A' |
2976 |
2959 |
6.59 |
|
|
|
| 5 |
A' |
2970 |
2953 |
22.14 |
|
|
|
| 6 |
A' |
2951 |
2934 |
9.40 |
|
|
|
| 7 |
A' |
2372 |
2359 |
48.67 |
|
|
|
| 8 |
A' |
1541 |
1533 |
11.80 |
|
|
|
| 9 |
A' |
1537 |
1528 |
7.21 |
|
|
|
| 10 |
A' |
1514 |
1506 |
2.23 |
|
|
|
| 11 |
A' |
1500 |
1491 |
6.47 |
|
|
|
| 12 |
A' |
1429 |
1421 |
6.50 |
|
|
|
| 13 |
A' |
1386 |
1378 |
1.27 |
|
|
|
| 14 |
A' |
1312 |
1304 |
23.82 |
|
|
|
| 15 |
A' |
1246 |
1239 |
15.50 |
|
|
|
| 16 |
A' |
1186 |
1179 |
5.61 |
|
|
|
| 17 |
A' |
1131 |
1125 |
5.04 |
|
|
|
| 18 |
A' |
974 |
969 |
1.22 |
|
|
|
| 19 |
A' |
941 |
935 |
1.27 |
|
|
|
| 20 |
A' |
809 |
805 |
0.97 |
|
|
|
| 21 |
A' |
734 |
730 |
1.90 |
|
|
|
| 22 |
A' |
675 |
671 |
15.03 |
|
|
|
| 23 |
A' |
497 |
494 |
0.73 |
|
|
|
| 24 |
A' |
371 |
369 |
0.75 |
|
|
|
| 25 |
A' |
240 |
239 |
1.66 |
|
|
|
| 26 |
A' |
233 |
232 |
1.32 |
|
|
|
| 27 |
A' |
178 |
177 |
2.13 |
|
|
|
| 28 |
A" |
3093 |
3076 |
17.10 |
|
|
|
| 29 |
A" |
3027 |
3011 |
24.57 |
|
|
|
| 30 |
A" |
3025 |
3008 |
0.00 |
|
|
|
| 31 |
A" |
3007 |
2990 |
10.35 |
|
|
|
| 32 |
A" |
2966 |
2950 |
28.86 |
|
|
|
| 33 |
A" |
1529 |
1520 |
1.34 |
|
|
|
| 34 |
A" |
1517 |
1509 |
0.54 |
|
|
|
| 35 |
A" |
1413 |
1405 |
6.51 |
|
|
|
| 36 |
A" |
1364 |
1356 |
4.52 |
|
|
|
| 37 |
A" |
1320 |
1313 |
0.14 |
|
|
|
| 38 |
A" |
1206 |
1199 |
2.86 |
|
|
|
| 39 |
A" |
1075 |
1069 |
0.30 |
|
|
|
| 40 |
A" |
960 |
955 |
0.93 |
|
|
|
| 41 |
A" |
953 |
948 |
0.70 |
|
|
|
| 42 |
A" |
910 |
905 |
0.04 |
|
|
|
| 43 |
A" |
765 |
761 |
4.15 |
|
|
|
| 44 |
A" |
358 |
356 |
0.00 |
|
|
|
| 45 |
A" |
237 |
236 |
0.01 |
|
|
|
| 46 |
A" |
132 |
131 |
8.44 |
|
|
|
| 47 |
A" |
93 |
93 |
6.60 |
|
|
|
| 48 |
A" |
42 |
42 |
14.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34392.0 cm
-1
Scaled (by 0.9945) Zero Point Vibrational Energy (zpe) 34202.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.208 |
|
|
|
| 2 |
H |
0.181 |
|
|
|
| 3 |
S |
-0.014 |
|
|
|
| 4 |
H |
0.064 |
|
|
|
| 5 |
C |
-0.552 |
|
|
|
| 6 |
C |
-0.323 |
|
|
|
| 7 |
C |
-0.504 |
|
|
|
| 8 |
C |
-0.504 |
|
|
|
| 9 |
H |
0.216 |
|
|
|
| 10 |
H |
0.216 |
|
|
|
| 11 |
H |
0.191 |
|
|
|
| 12 |
H |
0.191 |
|
|
|
| 13 |
H |
0.176 |
|
|
|
| 14 |
H |
0.176 |
|
|
|
| 15 |
H |
0.173 |
|
|
|
| 16 |
H |
0.173 |
|
|
|
| 17 |
H |
0.175 |
|
|
|
| 18 |
H |
0.175 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.893 |
-1.003 |
0.000 |
2.142 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-52.331 |
3.496 |
0.000 |
| y |
3.496 |
-45.124 |
0.000 |
| z |
0.000 |
0.000 |
-49.083 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.227 |
3.496 |
0.000 |
| y |
3.496 |
5.583 |
0.000 |
| z |
0.000 |
0.000 |
-0.355 |
|
| Polar |
|---|
| 3z2-r2 | -0.710 |
|---|
| x2-y2 | -7.207 |
|---|
| xy | 3.496 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
323.116 |
| (<r2>)1/2 |
17.975 |