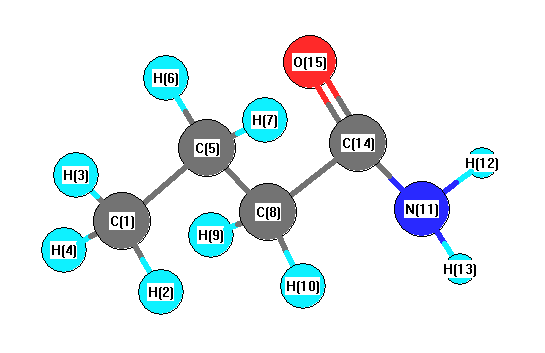
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3659 |
3531 |
19.92 |
|
|
|
| 2 |
A |
3529 |
3405 |
20.80 |
|
|
|
| 3 |
A |
3141 |
3031 |
24.66 |
|
|
|
| 4 |
A |
3115 |
3006 |
26.76 |
|
|
|
| 5 |
A |
3112 |
3003 |
37.92 |
|
|
|
| 6 |
A |
3084 |
2976 |
2.76 |
|
|
|
| 7 |
A |
3058 |
2951 |
28.96 |
|
|
|
| 8 |
A |
3052 |
2945 |
6.58 |
|
|
|
| 9 |
A |
3046 |
2940 |
20.08 |
|
|
|
| 10 |
A |
1757 |
1695 |
226.67 |
|
|
|
| 11 |
A |
1672 |
1613 |
76.35 |
|
|
|
| 12 |
A |
1572 |
1517 |
7.55 |
|
|
|
| 13 |
A |
1566 |
1511 |
8.45 |
|
|
|
| 14 |
A |
1556 |
1501 |
2.48 |
|
|
|
| 15 |
A |
1537 |
1483 |
4.32 |
|
|
|
| 16 |
A |
1465 |
1413 |
3.93 |
|
|
|
| 17 |
A |
1396 |
1347 |
2.91 |
|
|
|
| 18 |
A |
1377 |
1328 |
40.69 |
|
|
|
| 19 |
A |
1343 |
1296 |
90.37 |
|
|
|
| 20 |
A |
1331 |
1284 |
2.26 |
|
|
|
| 21 |
A |
1279 |
1234 |
28.45 |
|
|
|
| 22 |
A |
1165 |
1124 |
4.41 |
|
|
|
| 23 |
A |
1132 |
1092 |
12.50 |
|
|
|
| 24 |
A |
1070 |
1032 |
7.36 |
|
|
|
| 25 |
A |
1027 |
991 |
0.85 |
|
|
|
| 26 |
A |
917 |
885 |
1.18 |
|
|
|
| 27 |
A |
912 |
880 |
3.10 |
|
|
|
| 28 |
A |
865 |
835 |
3.40 |
|
|
|
| 29 |
A |
773 |
746 |
6.43 |
|
|
|
| 30 |
A |
730 |
704 |
36.50 |
|
|
|
| 31 |
A |
593 |
572 |
59.30 |
|
|
|
| 32 |
A |
550 |
531 |
26.62 |
|
|
|
| 33 |
A |
471 |
454 |
202.80 |
|
|
|
| 34 |
A |
426 |
411 |
7.29 |
|
|
|
| 35 |
A |
335 |
323 |
0.10 |
|
|
|
| 36 |
A |
246 |
237 |
0.05 |
|
|
|
| 37 |
A |
196 |
189 |
0.29 |
|
|
|
| 38 |
A |
95 |
92 |
1.91 |
|
|
|
| 39 |
A |
53 |
51 |
4.43 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29100.6 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 28079.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.550 |
|
|
-0.397 |
| 2 |
H |
0.194 |
|
|
0.111 |
| 3 |
H |
0.191 |
|
|
0.090 |
| 4 |
H |
0.185 |
|
|
0.103 |
| 5 |
C |
-0.376 |
|
|
0.285 |
| 6 |
H |
0.180 |
|
|
-0.040 |
| 7 |
H |
0.208 |
|
|
-0.009 |
| 8 |
C |
-0.455 |
|
|
-0.484 |
| 9 |
H |
0.193 |
|
|
0.122 |
| 10 |
H |
0.223 |
|
|
0.125 |
| 11 |
N |
-0.750 |
|
|
-1.011 |
| 12 |
H |
0.315 |
|
|
0.425 |
| 13 |
H |
0.310 |
|
|
0.430 |
| 14 |
C |
0.634 |
|
|
0.780 |
| 15 |
O |
-0.503 |
|
|
-0.530 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.579 |
3.292 |
0.637 |
3.706 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
1.573 |
3.290 |
0.660 |
3.706 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.207 |
-7.037 |
-0.463 |
| y |
-7.037 |
-36.045 |
-0.359 |
| z |
-0.463 |
-0.359 |
-37.512 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.429 |
-7.037 |
-0.463 |
| y |
-7.037 |
2.315 |
-0.359 |
| z |
-0.463 |
-0.359 |
0.114 |
|
| Polar |
|---|
| 3z2-r2 | 0.229 |
|---|
| x2-y2 | -3.162 |
|---|
| xy | -7.037 |
|---|
| xz | -0.463 |
|---|
| yz | -0.359 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.083 |
0.082 |
0.434 |
| y |
0.082 |
7.375 |
0.182 |
| z |
0.434 |
0.182 |
5.370 |
<r2> (average value of r
2) Å
2
| <r2> |
214.564 |
| (<r2>)1/2 |
14.648 |