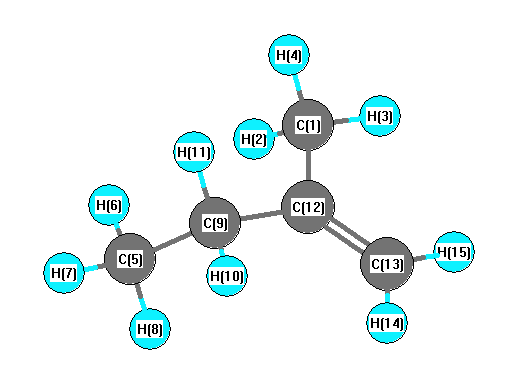
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3225 |
3112 |
19.07 |
|
|
|
| 2 |
A |
3149 |
3039 |
8.42 |
|
|
|
| 3 |
A |
3129 |
3020 |
34.97 |
|
|
|
| 4 |
A |
3123 |
3014 |
14.01 |
|
|
|
| 5 |
A |
3117 |
3008 |
40.04 |
|
|
|
| 6 |
A |
3086 |
2977 |
10.66 |
|
|
|
| 7 |
A |
3082 |
2974 |
20.90 |
|
|
|
| 8 |
A |
3055 |
2948 |
22.14 |
|
|
|
| 9 |
A |
3040 |
2934 |
26.61 |
|
|
|
| 10 |
A |
3036 |
2930 |
15.54 |
|
|
|
| 11 |
A |
1727 |
1666 |
11.23 |
|
|
|
| 12 |
A |
1572 |
1517 |
4.43 |
|
|
|
| 13 |
A |
1569 |
1514 |
8.04 |
|
|
|
| 14 |
A |
1561 |
1506 |
10.74 |
|
|
|
| 15 |
A |
1553 |
1498 |
13.98 |
|
|
|
| 16 |
A |
1540 |
1486 |
1.40 |
|
|
|
| 17 |
A |
1500 |
1448 |
0.25 |
|
|
|
| 18 |
A |
1463 |
1412 |
6.38 |
|
|
|
| 19 |
A |
1453 |
1402 |
6.75 |
|
|
|
| 20 |
A |
1372 |
1324 |
0.80 |
|
|
|
| 21 |
A |
1340 |
1293 |
1.49 |
|
|
|
| 22 |
A |
1291 |
1245 |
2.58 |
|
|
|
| 23 |
A |
1148 |
1108 |
0.09 |
|
|
|
| 24 |
A |
1126 |
1087 |
9.14 |
|
|
|
| 25 |
A |
1087 |
1049 |
5.19 |
|
|
|
| 26 |
A |
1042 |
1005 |
1.52 |
|
|
|
| 27 |
A |
1005 |
969 |
0.38 |
|
|
|
| 28 |
A |
944 |
911 |
51.35 |
|
|
|
| 29 |
A |
936 |
903 |
1.29 |
|
|
|
| 30 |
A |
822 |
793 |
2.80 |
|
|
|
| 31 |
A |
768 |
742 |
1.46 |
|
|
|
| 32 |
A |
731 |
706 |
0.10 |
|
|
|
| 33 |
A |
543 |
524 |
7.90 |
|
|
|
| 34 |
A |
438 |
422 |
1.30 |
|
|
|
| 35 |
A |
393 |
379 |
0.85 |
|
|
|
| 36 |
A |
260 |
251 |
0.59 |
|
|
|
| 37 |
A |
226 |
218 |
0.36 |
|
|
|
| 38 |
A |
185 |
178 |
0.43 |
|
|
|
| 39 |
A |
70 |
67 |
0.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30352.4 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 29287.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.595 |
|
|
-0.384 |
| 2 |
H |
0.201 |
|
|
0.123 |
| 3 |
H |
0.196 |
|
|
0.120 |
| 4 |
H |
0.198 |
|
|
0.114 |
| 5 |
C |
-0.547 |
|
|
-0.170 |
| 6 |
H |
0.188 |
|
|
0.051 |
| 7 |
H |
0.188 |
|
|
0.050 |
| 8 |
H |
0.196 |
|
|
0.062 |
| 9 |
C |
-0.427 |
|
|
-0.050 |
| 10 |
H |
0.195 |
|
|
0.043 |
| 11 |
H |
0.193 |
|
|
0.046 |
| 12 |
C |
0.049 |
|
|
0.165 |
| 13 |
C |
-0.380 |
|
|
-0.576 |
| 14 |
H |
0.173 |
|
|
0.199 |
| 15 |
H |
0.172 |
|
|
0.207 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.308 |
0.364 |
0.063 |
0.481 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.265 |
0.366 |
0.052 |
0.455 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.725 |
0.482 |
-0.936 |
| y |
0.482 |
-32.126 |
0.153 |
| z |
-0.936 |
0.153 |
-34.423 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.550 |
0.482 |
-0.936 |
| y |
0.482 |
1.448 |
0.153 |
| z |
-0.936 |
0.153 |
-1.998 |
|
| Polar |
|---|
| 3z2-r2 | -3.996 |
|---|
| x2-y2 | -0.599 |
|---|
| xy | 0.482 |
|---|
| xz | -0.936 |
|---|
| yz | 0.153 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.369 |
-0.813 |
-0.751 |
| y |
-0.813 |
8.221 |
0.387 |
| z |
-0.751 |
0.387 |
5.503 |
<r2> (average value of r
2) Å
2
| <r2> |
143.686 |
| (<r2>)1/2 |
11.987 |