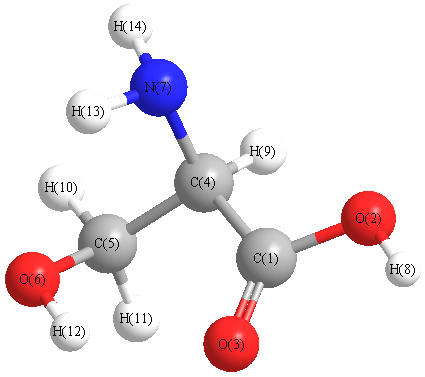
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3509 |
3386 |
0.83 |
|
|
|
| 2 |
A |
3467 |
3345 |
18.53 |
|
|
|
| 3 |
A |
3416 |
3296 |
0.33 |
|
|
|
| 4 |
A |
3374 |
3256 |
78.83 |
|
|
|
| 5 |
A |
3140 |
3030 |
24.39 |
|
|
|
| 6 |
A |
3074 |
2966 |
7.67 |
|
|
|
| 7 |
A |
2967 |
2863 |
72.87 |
|
|
|
| 8 |
A |
1732 |
1672 |
79.14 |
|
|
|
| 9 |
A |
1721 |
1661 |
90.11 |
|
|
|
| 10 |
A |
1575 |
1519 |
0.51 |
|
|
|
| 11 |
A |
1480 |
1428 |
92.70 |
|
|
|
| 12 |
A |
1409 |
1360 |
3.51 |
|
|
|
| 13 |
A |
1397 |
1348 |
9.06 |
|
|
|
| 14 |
A |
1377 |
1329 |
37.03 |
|
|
|
| 15 |
A |
1347 |
1300 |
9.67 |
|
|
|
| 16 |
A |
1257 |
1213 |
15.97 |
|
|
|
| 17 |
A |
1200 |
1157 |
6.14 |
|
|
|
| 18 |
A |
1153 |
1113 |
21.59 |
|
|
|
| 19 |
A |
1116 |
1077 |
203.72 |
|
|
|
| 20 |
A |
1040 |
1004 |
106.08 |
|
|
|
| 21 |
A |
1031 |
994 |
24.33 |
|
|
|
| 22 |
A |
963 |
929 |
9.40 |
|
|
|
| 23 |
A |
800 |
772 |
37.68 |
|
|
|
| 24 |
A |
737 |
712 |
232.21 |
|
|
|
| 25 |
A |
720 |
694 |
77.53 |
|
|
|
| 26 |
A |
670 |
647 |
138.45 |
|
|
|
| 27 |
A |
650 |
627 |
166.07 |
|
|
|
| 28 |
A |
557 |
538 |
3.78 |
|
|
|
| 29 |
A |
531 |
512 |
56.99 |
|
|
|
| 30 |
A |
406 |
392 |
8.90 |
|
|
|
| 31 |
A |
314 |
303 |
3.58 |
|
|
|
| 32 |
A |
294 |
283 |
9.41 |
|
|
|
| 33 |
A |
250 |
241 |
4.92 |
|
|
|
| 34 |
A |
224 |
216 |
48.63 |
|
|
|
| 35 |
A |
170 |
164 |
1.04 |
|
|
|
| 36 |
A |
53 |
51 |
0.78 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24560.9 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 23698.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.655 |
|
|
|
| 2 |
O |
-0.551 |
|
|
|
| 3 |
O |
-0.494 |
|
|
|
| 4 |
C |
-0.211 |
|
|
|
| 5 |
C |
-0.099 |
|
|
|
| 6 |
O |
-0.570 |
|
|
|
| 7 |
N |
-0.646 |
|
|
|
| 8 |
H |
0.369 |
|
|
|
| 9 |
H |
0.262 |
|
|
|
| 10 |
H |
0.161 |
|
|
|
| 11 |
H |
0.227 |
|
|
|
| 12 |
H |
0.345 |
|
|
|
| 13 |
H |
0.282 |
|
|
|
| 14 |
H |
0.270 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
4.024 |
0.024 |
0.756 |
4.094 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.515 |
-3.197 |
-0.457 |
| y |
-3.197 |
-43.906 |
0.597 |
| z |
-0.457 |
0.597 |
-38.513 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.306 |
-3.197 |
-0.457 |
| y |
-3.197 |
-3.391 |
0.597 |
| z |
-0.457 |
0.597 |
4.697 |
|
| Polar |
|---|
| 3z2-r2 | 9.395 |
|---|
| x2-y2 | 1.390 |
|---|
| xy | -3.197 |
|---|
| xz | -0.457 |
|---|
| yz | 0.597 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.351 |
0.221 |
0.137 |
| y |
0.221 |
6.829 |
0.048 |
| z |
0.137 |
0.048 |
4.721 |
<r2> (average value of r
2) Å
2
| <r2> |
217.506 |
| (<r2>)1/2 |
14.748 |