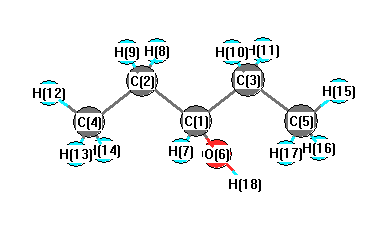
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3498 |
3376 |
1.34 |
|
|
|
| 2 |
A |
3150 |
3039 |
17.20 |
|
|
|
| 3 |
A |
3123 |
3013 |
34.87 |
|
|
|
| 4 |
A |
3116 |
3006 |
72.69 |
|
|
|
| 5 |
A |
3112 |
3003 |
18.24 |
|
|
|
| 6 |
A |
3095 |
2986 |
6.20 |
|
|
|
| 7 |
A |
3087 |
2978 |
4.06 |
|
|
|
| 8 |
A |
3058 |
2951 |
36.76 |
|
|
|
| 9 |
A |
3051 |
2944 |
1.90 |
|
|
|
| 10 |
A |
3050 |
2943 |
29.03 |
|
|
|
| 11 |
A |
3045 |
2938 |
21.91 |
|
|
|
| 12 |
A |
2983 |
2878 |
36.98 |
|
|
|
| 13 |
A |
1582 |
1527 |
6.95 |
|
|
|
| 14 |
A |
1581 |
1526 |
7.76 |
|
|
|
| 15 |
A |
1567 |
1512 |
8.51 |
|
|
|
| 16 |
A |
1562 |
1507 |
6.63 |
|
|
|
| 17 |
A |
1542 |
1488 |
6.03 |
|
|
|
| 18 |
A |
1531 |
1478 |
0.20 |
|
|
|
| 19 |
A |
1455 |
1404 |
5.39 |
|
|
|
| 20 |
A |
1450 |
1399 |
18.38 |
|
|
|
| 21 |
A |
1426 |
1376 |
2.83 |
|
|
|
| 22 |
A |
1409 |
1359 |
8.41 |
|
|
|
| 23 |
A |
1404 |
1355 |
0.33 |
|
|
|
| 24 |
A |
1361 |
1313 |
6.94 |
|
|
|
| 25 |
A |
1347 |
1300 |
2.18 |
|
|
|
| 26 |
A |
1326 |
1279 |
0.76 |
|
|
|
| 27 |
A |
1275 |
1231 |
23.67 |
|
|
|
| 28 |
A |
1177 |
1135 |
16.74 |
|
|
|
| 29 |
A |
1159 |
1119 |
0.92 |
|
|
|
| 30 |
A |
1109 |
1070 |
12.00 |
|
|
|
| 31 |
A |
1074 |
1036 |
11.82 |
|
|
|
| 32 |
A |
1036 |
999 |
14.70 |
|
|
|
| 33 |
A |
1026 |
990 |
4.23 |
|
|
|
| 34 |
A |
955 |
922 |
52.23 |
|
|
|
| 35 |
A |
929 |
896 |
17.13 |
|
|
|
| 36 |
A |
881 |
850 |
6.49 |
|
|
|
| 37 |
A |
795 |
767 |
2.53 |
|
|
|
| 38 |
A |
782 |
754 |
1.16 |
|
|
|
| 39 |
A |
496 |
478 |
2.55 |
|
|
|
| 40 |
A |
451 |
435 |
8.82 |
|
|
|
| 41 |
A |
403 |
389 |
6.81 |
|
|
|
| 42 |
A |
327 |
315 |
10.07 |
|
|
|
| 43 |
A |
287 |
277 |
25.40 |
|
|
|
| 44 |
A |
242 |
233 |
7.19 |
|
|
|
| 45 |
A |
211 |
203 |
37.67 |
|
|
|
| 46 |
A |
175 |
169 |
53.29 |
|
|
|
| 47 |
A |
123 |
119 |
2.54 |
|
|
|
| 48 |
A |
90 |
87 |
4.22 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36454.3 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 35174.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.033 |
|
|
|
| 2 |
C |
-0.372 |
|
|
|
| 3 |
C |
-0.378 |
|
|
|
| 4 |
C |
-0.556 |
|
|
|
| 5 |
C |
-0.567 |
|
|
|
| 6 |
O |
-0.547 |
|
|
|
| 7 |
H |
0.166 |
|
|
|
| 8 |
H |
0.180 |
|
|
|
| 9 |
H |
0.200 |
|
|
|
| 10 |
H |
0.200 |
|
|
|
| 11 |
H |
0.181 |
|
|
|
| 12 |
H |
0.180 |
|
|
|
| 13 |
H |
0.174 |
|
|
|
| 14 |
H |
0.211 |
|
|
|
| 15 |
H |
0.194 |
|
|
|
| 16 |
H |
0.183 |
|
|
|
| 17 |
H |
0.180 |
|
|
|
| 18 |
H |
0.337 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.027 |
-0.255 |
1.207 |
1.605 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.768 |
1.847 |
0.243 |
| y |
1.847 |
-38.540 |
2.025 |
| z |
0.243 |
2.025 |
-39.906 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.545 |
1.847 |
0.243 |
| y |
1.847 |
1.297 |
2.025 |
| z |
0.243 |
2.025 |
-0.752 |
|
| Polar |
|---|
| 3z2-r2 | -1.504 |
|---|
| x2-y2 | -1.228 |
|---|
| xy | 1.847 |
|---|
| xz | 0.243 |
|---|
| yz | 2.025 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.302 |
0.209 |
0.125 |
| y |
0.209 |
7.747 |
0.082 |
| z |
0.125 |
0.082 |
7.209 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |