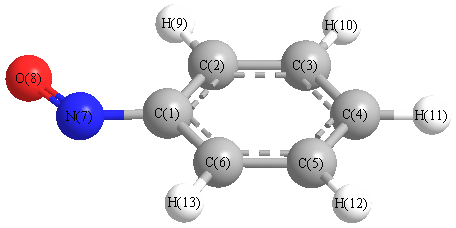
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3240 |
3126 |
4.78 |
|
|
|
| 2 |
A' |
3230 |
3116 |
6.34 |
|
|
|
| 3 |
A' |
3218 |
3105 |
7.29 |
|
|
|
| 4 |
A' |
3211 |
3098 |
6.30 |
|
|
|
| 5 |
A' |
3199 |
3087 |
0.45 |
|
|
|
| 6 |
A' |
1626 |
1569 |
7.36 |
|
|
|
| 7 |
A' |
1618 |
1561 |
8.47 |
|
|
|
| 8 |
A' |
1530 |
1476 |
5.74 |
|
|
|
| 9 |
A' |
1514 |
1461 |
13.68 |
|
|
|
| 10 |
A' |
1381 |
1333 |
3.37 |
|
|
|
| 11 |
A' |
1344 |
1297 |
9.65 |
|
|
|
| 12 |
A' |
1320 |
1273 |
64.66 |
|
|
|
| 13 |
A' |
1229 |
1186 |
0.50 |
|
|
|
| 14 |
A' |
1226 |
1183 |
0.37 |
|
|
|
| 15 |
A' |
1114 |
1074 |
66.01 |
|
|
|
| 16 |
A' |
1107 |
1068 |
68.71 |
|
|
|
| 17 |
A' |
1050 |
1013 |
15.17 |
|
|
|
| 18 |
A' |
1032 |
996 |
0.77 |
|
|
|
| 19 |
A' |
804 |
776 |
37.27 |
|
|
|
| 20 |
A' |
677 |
653 |
13.42 |
|
|
|
| 21 |
A' |
651 |
629 |
0.15 |
|
|
|
| 22 |
A' |
452 |
436 |
0.64 |
|
|
|
| 23 |
A' |
262 |
253 |
2.65 |
|
|
|
| 24 |
A" |
1073 |
1036 |
2.29 |
|
|
|
| 25 |
A" |
1048 |
1011 |
0.00 |
|
|
|
| 26 |
A" |
1004 |
969 |
6.33 |
|
|
|
| 27 |
A" |
897 |
865 |
0.00 |
|
|
|
| 28 |
A" |
812 |
784 |
42.17 |
|
|
|
| 29 |
A" |
732 |
706 |
50.47 |
|
|
|
| 30 |
A" |
498 |
480 |
3.40 |
|
|
|
| 31 |
A" |
438 |
422 |
0.02 |
|
|
|
| 32 |
A" |
259 |
250 |
0.11 |
|
|
|
| 33 |
A" |
137 |
132 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21464.9 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 20711.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.163 |
|
|
|
| 2 |
C |
-0.151 |
|
|
|
| 3 |
C |
-0.192 |
|
|
|
| 4 |
C |
-0.167 |
|
|
|
| 5 |
C |
-0.191 |
|
|
|
| 6 |
C |
-0.143 |
|
|
|
| 7 |
N |
-0.096 |
|
|
|
| 8 |
O |
-0.268 |
|
|
|
| 9 |
H |
0.219 |
|
|
|
| 10 |
H |
0.203 |
|
|
|
| 11 |
H |
0.204 |
|
|
|
| 12 |
H |
0.203 |
|
|
|
| 13 |
H |
0.217 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.340 |
-3.491 |
0.000 |
3.740 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.527 |
2.677 |
0.000 |
| y |
2.677 |
-47.664 |
0.000 |
| z |
0.000 |
0.000 |
-47.959 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.284 |
2.677 |
0.000 |
| y |
2.677 |
-2.921 |
0.000 |
| z |
0.000 |
0.000 |
-3.364 |
|
| Polar |
|---|
| 3z2-r2 | -6.727 |
|---|
| x2-y2 | 6.137 |
|---|
| xy | 2.677 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.614 |
-1.809 |
0.000 |
| y |
-1.809 |
13.573 |
0.000 |
| z |
0.000 |
0.000 |
2.735 |
<r2> (average value of r
2) Å
2
| <r2> |
250.838 |
| (<r2>)1/2 |
15.838 |