Jump to
S1C2
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -613.767242 |
| Energy at 298.15K | -613.774294 |
| HF Energy | -613.767242 |
| Nuclear repulsion energy | 202.553640 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3187 |
3075 |
3.83 |
|
|
|
| 2 |
A' |
3130 |
3020 |
9.05 |
|
|
|
| 3 |
A' |
3116 |
3006 |
14.42 |
|
|
|
| 4 |
A' |
3106 |
2997 |
10.26 |
|
|
|
| 5 |
A' |
3039 |
2933 |
20.64 |
|
|
|
| 6 |
A' |
1754 |
1692 |
1.48 |
|
|
|
| 7 |
A' |
1559 |
1504 |
14.19 |
|
|
|
| 8 |
A' |
1524 |
1471 |
2.43 |
|
|
|
| 9 |
A' |
1468 |
1417 |
4.13 |
|
|
|
| 10 |
A' |
1378 |
1330 |
0.43 |
|
|
|
| 11 |
A' |
1359 |
1311 |
14.75 |
|
|
|
| 12 |
A' |
1297 |
1252 |
19.87 |
|
|
|
| 13 |
A' |
1132 |
1092 |
3.68 |
|
|
|
| 14 |
A' |
1046 |
1010 |
9.58 |
|
|
|
| 15 |
A' |
923 |
890 |
12.25 |
|
|
|
| 16 |
A' |
671 |
647 |
18.05 |
|
|
|
| 17 |
A' |
556 |
537 |
14.40 |
|
|
|
| 18 |
A' |
336 |
325 |
1.65 |
|
|
|
| 19 |
A' |
180 |
174 |
1.47 |
|
|
|
| 20 |
A" |
3160 |
3049 |
4.23 |
|
|
|
| 21 |
A" |
3083 |
2975 |
17.73 |
|
|
|
| 22 |
A" |
1550 |
1496 |
8.96 |
|
|
|
| 23 |
A" |
1213 |
1170 |
1.19 |
|
|
|
| 24 |
A" |
1113 |
1074 |
0.44 |
|
|
|
| 25 |
A" |
1012 |
976 |
47.89 |
|
|
|
| 26 |
A" |
934 |
901 |
1.02 |
|
|
|
| 27 |
A" |
751 |
724 |
1.00 |
|
|
|
| 28 |
A" |
240 |
231 |
6.52 |
|
|
|
| 29 |
A" |
204 |
196 |
1.45 |
|
|
|
| 30 |
A" |
104 |
100 |
2.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22061.0 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 21286.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.331 |
0.368 |
0.000 |
| C2 |
0.000 |
1.058 |
0.000 |
| C3 |
1.203 |
0.491 |
0.000 |
| C4 |
2.503 |
1.257 |
0.000 |
| Cl5 |
-1.185 |
-1.536 |
0.000 |
| H6 |
-1.916 |
0.587 |
0.894 |
| H7 |
-1.916 |
0.587 |
-0.894 |
| H8 |
-0.091 |
2.146 |
0.000 |
| H9 |
1.274 |
-0.593 |
0.000 |
| H10 |
2.331 |
2.338 |
0.000 |
| H11 |
3.103 |
1.001 |
0.883 |
| H12 |
3.103 |
1.001 |
-0.883 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4992 | 2.5368 | 3.9356 | 1.9094 | 1.0912 | 1.0912 | 2.1676 | 2.7757 | 4.1584 | 4.5644 | 4.5644 |
C2 | 1.4992 | | 1.3301 | 2.5110 | 2.8521 | 2.1665 | 2.1665 | 1.0910 | 2.0852 | 2.6595 | 3.2263 | 3.2263 | C3 | 2.5368 | 1.3301 | | 1.5087 | 3.1325 | 3.2466 | 3.2466 | 2.1000 | 1.0864 | 2.1641 | 2.1558 | 2.1558 | C4 | 3.9356 | 2.5110 | 1.5087 | | 4.6262 | 4.5585 | 4.5585 | 2.7416 | 2.2212 | 1.0947 | 1.0977 | 1.0977 | Cl5 | 1.9094 | 2.8521 | 3.1325 | 4.6262 | | 2.4174 | 2.4174 | 3.8408 | 2.6331 | 5.2319 | 5.0594 | 5.0594 | H6 | 1.0912 | 2.1665 | 3.2466 | 4.5585 | 2.4174 | | 1.7881 | 2.5614 | 3.5168 | 4.6806 | 5.0359 | 5.3403 | H7 | 1.0912 | 2.1665 | 3.2466 | 4.5585 | 2.4174 | 1.7881 | | 2.5614 | 3.5168 | 4.6806 | 5.3403 | 5.0359 | H8 | 2.1676 | 1.0910 | 2.1000 | 2.7416 | 3.8408 | 2.5614 | 2.5614 | | 3.0593 | 2.4295 | 3.5051 | 3.5051 | H9 | 2.7757 | 2.0852 | 1.0864 | 2.2212 | 2.6331 | 3.5168 | 3.5168 | 3.0593 | | 3.1160 | 2.5817 | 2.5817 | H10 | 4.1584 | 2.6595 | 2.1641 | 1.0947 | 5.2319 | 4.6806 | 4.6806 | 2.4295 | 3.1160 | | 1.7784 | 1.7784 | H11 | 4.5644 | 3.2263 | 2.1558 | 1.0977 | 5.0594 | 5.0359 | 5.3403 | 3.5051 | 2.5817 | 1.7784 | | 1.7666 | H12 | 4.5644 | 3.2263 | 2.1558 | 1.0977 | 5.0594 | 5.3403 | 5.0359 | 3.5051 | 2.5817 | 1.7784 | 1.7666 | |
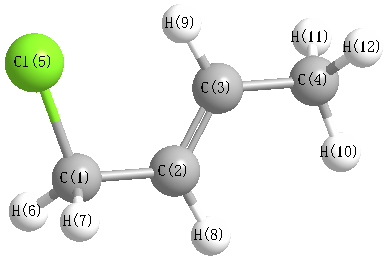 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.342 |
|
C1 |
C2 |
H8 |
112.668 |
| C2 |
C1 |
Cl5 |
113.045 |
|
C2 |
C1 |
H6 |
112.566 |
| C2 |
C1 |
H7 |
112.566 |
|
C2 |
C3 |
C4 |
124.271 |
| C2 |
C3 |
H9 |
118.945 |
|
C3 |
C2 |
H8 |
119.990 |
| C3 |
C4 |
H10 |
111.469 |
|
C3 |
C4 |
H11 |
110.616 |
| C3 |
C4 |
H12 |
110.616 |
|
C4 |
C3 |
H9 |
116.783 |
| Cl5 |
C1 |
H6 |
103.988 |
|
Cl5 |
C1 |
H7 |
103.988 |
| H6 |
C1 |
H7 |
110.038 |
|
H10 |
C4 |
H11 |
108.422 |
| H10 |
C4 |
H12 |
108.422 |
|
H11 |
C4 |
H12 |
107.155 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.564 |
|
|
|
| 2 |
C |
-0.160 |
|
|
|
| 3 |
C |
-0.158 |
|
|
|
| 4 |
C |
-0.603 |
|
|
|
| 5 |
Cl |
-0.062 |
|
|
|
| 6 |
H |
0.261 |
|
|
|
| 7 |
H |
0.261 |
|
|
|
| 8 |
H |
0.196 |
|
|
|
| 9 |
H |
0.217 |
|
|
|
| 10 |
H |
0.195 |
|
|
|
| 11 |
H |
0.208 |
|
|
|
| 12 |
H |
0.208 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.431 |
2.498 |
0.000 |
2.535 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.149 |
-3.057 |
0.000 |
| y |
-3.057 |
-39.333 |
0.000 |
| z |
0.000 |
0.000 |
-39.665 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.349 |
-3.057 |
0.000 |
| y |
-3.057 |
-1.926 |
0.000 |
| z |
0.000 |
0.000 |
-2.423 |
|
| Polar |
|---|
| 3z2-r2 | -4.847 |
|---|
| x2-y2 | 4.184 |
|---|
| xy | -3.057 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.974 |
0.344 |
0.000 |
| y |
0.344 |
8.290 |
0.000 |
| z |
0.000 |
0.000 |
4.096 |
<r2> (average value of r
2) Å
2
| <r2> |
213.038 |
| (<r2>)1/2 |
14.596 |
Jump to
S1C1
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -613.770626 |
| Energy at 298.15K | -613.777627 |
| HF Energy | -613.770626 |
| Nuclear repulsion energy | 197.936833 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3211 |
3098 |
5.44 |
|
|
|
| 2 |
A |
3176 |
3064 |
9.34 |
|
|
|
| 3 |
A |
3148 |
3037 |
17.91 |
|
|
|
| 4 |
A |
3137 |
3027 |
7.74 |
|
|
|
| 5 |
A |
3121 |
3011 |
5.04 |
|
|
|
| 6 |
A |
3082 |
2974 |
14.68 |
|
|
|
| 7 |
A |
3039 |
2933 |
15.70 |
|
|
|
| 8 |
A |
1726 |
1666 |
21.75 |
|
|
|
| 9 |
A |
1554 |
1499 |
16.62 |
|
|
|
| 10 |
A |
1547 |
1493 |
10.90 |
|
|
|
| 11 |
A |
1533 |
1479 |
5.45 |
|
|
|
| 12 |
A |
1465 |
1414 |
5.87 |
|
|
|
| 13 |
A |
1385 |
1336 |
1.26 |
|
|
|
| 14 |
A |
1373 |
1324 |
0.09 |
|
|
|
| 15 |
A |
1268 |
1223 |
40.92 |
|
|
|
| 16 |
A |
1217 |
1174 |
6.79 |
|
|
|
| 17 |
A |
1128 |
1088 |
1.54 |
|
|
|
| 18 |
A |
1108 |
1069 |
0.76 |
|
|
|
| 19 |
A |
1084 |
1046 |
11.49 |
|
|
|
| 20 |
A |
1016 |
981 |
41.07 |
|
|
|
| 21 |
A |
964 |
931 |
12.66 |
|
|
|
| 22 |
A |
883 |
852 |
3.75 |
|
|
|
| 23 |
A |
822 |
793 |
7.63 |
|
|
|
| 24 |
A |
599 |
578 |
91.50 |
|
|
|
| 25 |
A |
507 |
489 |
2.80 |
|
|
|
| 26 |
A |
337 |
325 |
10.56 |
|
|
|
| 27 |
A |
285 |
275 |
2.48 |
|
|
|
| 28 |
A |
206 |
199 |
1.88 |
|
|
|
| 29 |
A |
153 |
148 |
2.91 |
|
|
|
| 30 |
A |
90 |
87 |
0.60 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22081.1 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 21306.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.733 |
0.922 |
0.031 |
| C2 |
0.545 |
0.300 |
0.448 |
| C3 |
1.599 |
0.181 |
-0.365 |
| C4 |
2.920 |
-0.427 |
0.028 |
| Cl5 |
-2.120 |
-0.444 |
-0.063 |
| H6 |
-1.147 |
1.620 |
0.755 |
| H7 |
-0.710 |
1.345 |
-0.971 |
| H8 |
0.594 |
-0.065 |
1.470 |
| H9 |
1.522 |
0.540 |
-1.391 |
| H10 |
2.906 |
-0.767 |
1.068 |
| H11 |
3.155 |
-1.284 |
-0.615 |
| H12 |
3.734 |
0.301 |
-0.090 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4807 | 2.4785 | 3.8939 | 1.9490 | 1.0880 | 1.0879 | 2.1916 | 2.6932 | 4.1433 | 4.5169 | 4.5114 |
C2 | 1.4807 | | 1.3364 | 2.5191 | 2.8137 | 2.1680 | 2.1628 | 1.0867 | 2.0960 | 2.6642 | 3.2333 | 3.2341 | C3 | 2.4785 | 1.3364 | | 1.5065 | 3.7832 | 3.2966 | 2.6555 | 2.1069 | 1.0896 | 2.1589 | 2.1520 | 2.1562 | C4 | 3.8939 | 2.5191 | 1.5065 | | 5.0413 | 4.6109 | 4.1608 | 2.7609 | 2.2143 | 1.0941 | 1.0978 | 1.0978 | Cl5 | 1.9490 | 2.8137 | 3.7832 | 5.0413 | | 2.4243 | 2.4523 | 3.1398 | 4.0000 | 5.1621 | 5.3706 | 5.9014 | H6 | 1.0880 | 2.1680 | 3.2966 | 4.6109 | 2.4243 | | 1.8016 | 2.5260 | 3.5913 | 4.7141 | 5.3689 | 5.1264 | H7 | 1.0879 | 2.1628 | 2.6555 | 4.1608 | 2.4523 | 1.8016 | | 3.1054 | 2.4095 | 4.6572 | 4.6881 | 4.6488 | H8 | 2.1916 | 1.0867 | 2.1069 | 2.7609 | 3.1398 | 2.5260 | 3.1054 | | 3.0682 | 2.4498 | 3.5211 | 3.5257 | H9 | 2.6932 | 2.0960 | 1.0896 | 2.2143 | 4.0000 | 3.5913 | 2.4095 | 3.0682 | | 3.1097 | 2.5686 | 2.5770 | H10 | 4.1433 | 2.6642 | 2.1589 | 1.0941 | 5.1621 | 4.7141 | 4.6572 | 2.4498 | 3.1097 | | 1.7788 | 1.7795 | H11 | 4.5169 | 3.2333 | 2.1520 | 1.0978 | 5.3706 | 5.3689 | 4.6881 | 3.5211 | 2.5686 | 1.7788 | | 1.7671 | H12 | 4.5114 | 3.2341 | 2.1562 | 1.0978 | 5.9014 | 5.1264 | 4.6488 | 3.5257 | 2.5770 | 1.7795 | 1.7671 | |
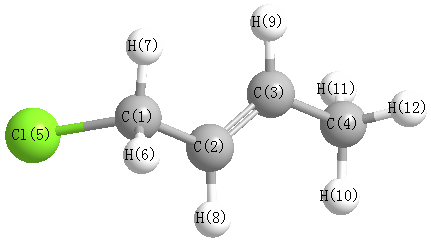 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.162 |
|
C1 |
C2 |
H8 |
116.377 |
| C2 |
C1 |
Cl5 |
109.494 |
|
C2 |
C1 |
H6 |
114.265 |
| C2 |
C1 |
H7 |
113.828 |
|
C2 |
C3 |
C4 |
124.670 |
| C2 |
C3 |
H9 |
119.189 |
|
C3 |
C2 |
H8 |
120.461 |
| C3 |
C4 |
H10 |
111.238 |
|
C3 |
C4 |
H11 |
110.461 |
| C3 |
C4 |
H12 |
110.793 |
|
C4 |
C3 |
H9 |
116.141 |
| Cl5 |
C1 |
H6 |
102.184 |
|
Cl5 |
C1 |
H7 |
104.081 |
| H6 |
C1 |
H7 |
111.780 |
|
H10 |
C4 |
H11 |
108.487 |
| H10 |
C4 |
H12 |
108.548 |
|
H11 |
C4 |
H12 |
107.187 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.537 |
|
|
|
| 2 |
C |
-0.145 |
|
|
|
| 3 |
C |
-0.141 |
|
|
|
| 4 |
C |
-0.604 |
|
|
|
| 5 |
Cl |
-0.104 |
|
|
|
| 6 |
H |
0.259 |
|
|
|
| 7 |
H |
0.250 |
|
|
|
| 8 |
H |
0.202 |
|
|
|
| 9 |
H |
0.191 |
|
|
|
| 10 |
H |
0.205 |
|
|
|
| 11 |
H |
0.215 |
|
|
|
| 12 |
H |
0.210 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.955 |
1.614 |
0.027 |
3.367 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.646 |
-3.139 |
-0.933 |
| y |
-3.139 |
-38.167 |
-0.701 |
| z |
-0.933 |
-0.701 |
-36.819 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.153 |
-3.139 |
-0.933 |
| y |
-3.139 |
0.566 |
-0.701 |
| z |
-0.933 |
-0.701 |
2.587 |
|
| Polar |
|---|
| 3z2-r2 | 5.175 |
|---|
| x2-y2 | -2.479 |
|---|
| xy | -3.139 |
|---|
| xz | -0.933 |
|---|
| yz | -0.701 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.632 |
0.416 |
-0.865 |
| y |
0.416 |
5.537 |
-0.199 |
| z |
-0.865 |
-0.199 |
5.811 |
<r2> (average value of r
2) Å
2
| <r2> |
235.383 |
| (<r2>)1/2 |
15.342 |
