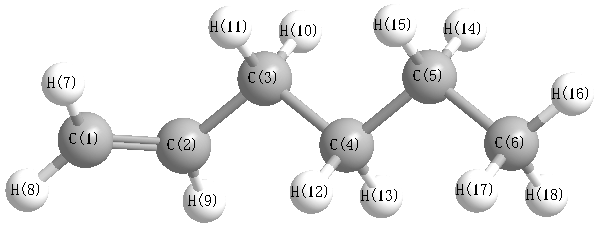
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3227 |
3114 |
20.64 |
|
|
|
| 2 |
A |
3150 |
3040 |
4.58 |
|
|
|
| 3 |
A |
3137 |
3027 |
25.44 |
|
|
|
| 4 |
A |
3113 |
3003 |
68.52 |
|
|
|
| 5 |
A |
3110 |
3001 |
36.92 |
|
|
|
| 6 |
A |
3093 |
2984 |
16.54 |
|
|
|
| 7 |
A |
3071 |
2964 |
3.57 |
|
|
|
| 8 |
A |
3059 |
2952 |
9.04 |
|
|
|
| 9 |
A |
3046 |
2939 |
42.62 |
|
|
|
| 10 |
A |
3042 |
2935 |
29.81 |
|
|
|
| 11 |
A |
3034 |
2928 |
0.38 |
|
|
|
| 12 |
A |
3024 |
2918 |
12.84 |
|
|
|
| 13 |
A |
1718 |
1658 |
9.17 |
|
|
|
| 14 |
A |
1572 |
1517 |
7.02 |
|
|
|
| 15 |
A |
1567 |
1512 |
8.16 |
|
|
|
| 16 |
A |
1559 |
1504 |
3.51 |
|
|
|
| 17 |
A |
1547 |
1492 |
0.26 |
|
|
|
| 18 |
A |
1538 |
1484 |
2.32 |
|
|
|
| 19 |
A |
1508 |
1455 |
1.19 |
|
|
|
| 20 |
A |
1462 |
1411 |
4.39 |
|
|
|
| 21 |
A |
1400 |
1351 |
1.04 |
|
|
|
| 22 |
A |
1383 |
1335 |
0.80 |
|
|
|
| 23 |
A |
1373 |
1325 |
0.53 |
|
|
|
| 24 |
A |
1363 |
1315 |
0.08 |
|
|
|
| 25 |
A |
1350 |
1303 |
0.41 |
|
|
|
| 26 |
A |
1318 |
1272 |
0.19 |
|
|
|
| 27 |
A |
1285 |
1240 |
0.37 |
|
|
|
| 28 |
A |
1221 |
1178 |
0.44 |
|
|
|
| 29 |
A |
1142 |
1102 |
4.91 |
|
|
|
| 30 |
A |
1085 |
1047 |
7.87 |
|
|
|
| 31 |
A |
1049 |
1012 |
9.01 |
|
|
|
| 32 |
A |
1040 |
1004 |
1.15 |
|
|
|
| 33 |
A |
1000 |
965 |
0.36 |
|
|
|
| 34 |
A |
972 |
938 |
35.93 |
|
|
|
| 35 |
A |
961 |
927 |
16.37 |
|
|
|
| 36 |
A |
945 |
912 |
4.53 |
|
|
|
| 37 |
A |
899 |
868 |
1.53 |
|
|
|
| 38 |
A |
822 |
793 |
1.19 |
|
|
|
| 39 |
A |
752 |
726 |
5.22 |
|
|
|
| 40 |
A |
656 |
633 |
12.74 |
|
|
|
| 41 |
A |
467 |
451 |
1.07 |
|
|
|
| 42 |
A |
365 |
352 |
0.55 |
|
|
|
| 43 |
A |
359 |
347 |
0.06 |
|
|
|
| 44 |
A |
244 |
236 |
0.00 |
|
|
|
| 45 |
A |
180 |
173 |
0.01 |
|
|
|
| 46 |
A |
125 |
121 |
0.08 |
|
|
|
| 47 |
A |
88 |
85 |
0.01 |
|
|
|
| 48 |
A |
71 |
69 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36746.9 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 35457.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.378 |
|
|
|
| 2 |
C |
-0.145 |
|
|
|
| 3 |
C |
-0.426 |
|
|
|
| 4 |
C |
-0.350 |
|
|
|
| 5 |
C |
-0.378 |
|
|
|
| 6 |
C |
-0.551 |
|
|
|
| 7 |
H |
0.177 |
|
|
|
| 8 |
H |
0.181 |
|
|
|
| 9 |
H |
0.179 |
|
|
|
| 10 |
H |
0.197 |
|
|
|
| 11 |
H |
0.191 |
|
|
|
| 12 |
H |
0.193 |
|
|
|
| 13 |
H |
0.183 |
|
|
|
| 14 |
H |
0.185 |
|
|
|
| 15 |
H |
0.186 |
|
|
|
| 16 |
H |
0.186 |
|
|
|
| 17 |
H |
0.187 |
|
|
|
| 18 |
H |
0.186 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.354 |
0.032 |
0.085 |
0.366 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.566 |
-0.926 |
-0.498 |
| y |
-0.926 |
-40.551 |
-1.076 |
| z |
-0.498 |
-1.076 |
-39.059 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.761 |
-0.926 |
-0.498 |
| y |
-0.926 |
-0.738 |
-1.076 |
| z |
-0.498 |
-1.076 |
1.499 |
|
| Polar |
|---|
| 3z2-r2 | 2.999 |
|---|
| x2-y2 | -0.015 |
|---|
| xy | -0.926 |
|---|
| xz | -0.498 |
|---|
| yz | -1.076 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.688 |
-0.620 |
-1.145 |
| y |
-0.620 |
7.218 |
-0.527 |
| z |
-1.145 |
-0.527 |
8.072 |
<r2> (average value of r
2) Å
2
| <r2> |
286.758 |
| (<r2>)1/2 |
16.934 |