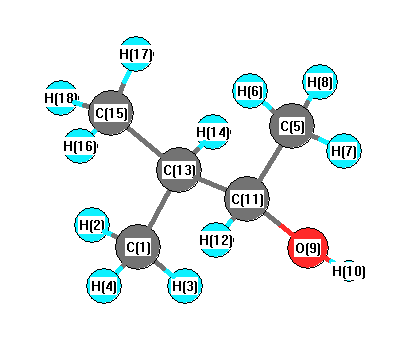
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3475 |
3353 |
1.73 |
|
|
|
| 2 |
A |
3162 |
3051 |
8.82 |
|
|
|
| 3 |
A |
3139 |
3029 |
23.51 |
|
|
|
| 4 |
A |
3119 |
3009 |
36.27 |
|
|
|
| 5 |
A |
3110 |
3001 |
78.45 |
|
|
|
| 6 |
A |
3107 |
2998 |
8.84 |
|
|
|
| 7 |
A |
3100 |
2991 |
5.90 |
|
|
|
| 8 |
A |
3056 |
2949 |
30.18 |
|
|
|
| 9 |
A |
3048 |
2941 |
12.37 |
|
|
|
| 10 |
A |
3045 |
2938 |
22.73 |
|
|
|
| 11 |
A |
3042 |
2935 |
13.92 |
|
|
|
| 12 |
A |
2978 |
2874 |
41.90 |
|
|
|
| 13 |
A |
1583 |
1528 |
11.29 |
|
|
|
| 14 |
A |
1575 |
1520 |
2.78 |
|
|
|
| 15 |
A |
1564 |
1509 |
13.02 |
|
|
|
| 16 |
A |
1561 |
1506 |
7.38 |
|
|
|
| 17 |
A |
1557 |
1503 |
0.93 |
|
|
|
| 18 |
A |
1553 |
1499 |
4.77 |
|
|
|
| 19 |
A |
1471 |
1419 |
5.86 |
|
|
|
| 20 |
A |
1455 |
1404 |
18.97 |
|
|
|
| 21 |
A |
1447 |
1397 |
16.26 |
|
|
|
| 22 |
A |
1428 |
1378 |
0.53 |
|
|
|
| 23 |
A |
1391 |
1342 |
2.21 |
|
|
|
| 24 |
A |
1380 |
1331 |
9.52 |
|
|
|
| 25 |
A |
1367 |
1319 |
1.25 |
|
|
|
| 26 |
A |
1292 |
1246 |
25.26 |
|
|
|
| 27 |
A |
1221 |
1178 |
6.89 |
|
|
|
| 28 |
A |
1193 |
1151 |
16.50 |
|
|
|
| 29 |
A |
1154 |
1114 |
3.36 |
|
|
|
| 30 |
A |
1106 |
1067 |
50.40 |
|
|
|
| 31 |
A |
1058 |
1021 |
22.54 |
|
|
|
| 32 |
A |
1027 |
991 |
10.27 |
|
|
|
| 33 |
A |
987 |
953 |
1.14 |
|
|
|
| 34 |
A |
970 |
936 |
0.53 |
|
|
|
| 35 |
A |
941 |
908 |
23.39 |
|
|
|
| 36 |
A |
884 |
853 |
8.50 |
|
|
|
| 37 |
A |
791 |
763 |
3.10 |
|
|
|
| 38 |
A |
512 |
494 |
3.18 |
|
|
|
| 39 |
A |
460 |
444 |
7.75 |
|
|
|
| 40 |
A |
404 |
390 |
1.02 |
|
|
|
| 41 |
A |
365 |
352 |
7.84 |
|
|
|
| 42 |
A |
363 |
350 |
5.21 |
|
|
|
| 43 |
A |
285 |
275 |
13.70 |
|
|
|
| 44 |
A |
279 |
269 |
107.61 |
|
|
|
| 45 |
A |
244 |
235 |
0.44 |
|
|
|
| 46 |
A |
229 |
221 |
0.15 |
|
|
|
| 47 |
A |
224 |
216 |
0.32 |
|
|
|
| 48 |
A |
85 |
82 |
1.45 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36392.1 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 35114.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.530 |
|
|
-0.479 |
| 2 |
H |
0.174 |
|
|
0.123 |
| 3 |
H |
0.216 |
|
|
0.132 |
| 4 |
H |
0.171 |
|
|
0.113 |
| 5 |
C |
-0.567 |
|
|
-0.496 |
| 6 |
H |
0.190 |
|
|
0.126 |
| 7 |
H |
0.175 |
|
|
0.123 |
| 8 |
H |
0.202 |
|
|
0.134 |
| 9 |
O |
-0.548 |
|
|
-0.632 |
| 10 |
H |
0.331 |
|
|
0.400 |
| 11 |
C |
0.050 |
|
|
0.301 |
| 12 |
H |
0.166 |
|
|
0.010 |
| 13 |
C |
-0.244 |
|
|
0.242 |
| 14 |
H |
0.196 |
|
|
0.040 |
| 15 |
C |
-0.541 |
|
|
-0.550 |
| 16 |
H |
0.181 |
|
|
0.139 |
| 17 |
H |
0.186 |
|
|
0.128 |
| 18 |
H |
0.191 |
|
|
0.147 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.970 |
0.953 |
0.916 |
1.640 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.964 |
0.963 |
0.923 |
1.645 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.954 |
-0.994 |
2.060 |
| y |
-0.994 |
-41.519 |
-1.367 |
| z |
2.060 |
-1.367 |
-39.490 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.550 |
-0.994 |
2.060 |
| y |
-0.994 |
-3.797 |
-1.367 |
| z |
2.060 |
-1.367 |
-0.754 |
|
| Polar |
|---|
| 3z2-r2 | -1.508 |
|---|
| x2-y2 | 5.565 |
|---|
| xy | -0.994 |
|---|
| xz | 2.060 |
|---|
| yz | -1.367 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.990 |
-0.084 |
0.195 |
| y |
-0.084 |
8.096 |
0.029 |
| z |
0.195 |
0.029 |
7.071 |
<r2> (average value of r
2) Å
2
| <r2> |
196.653 |
| (<r2>)1/2 |
14.023 |