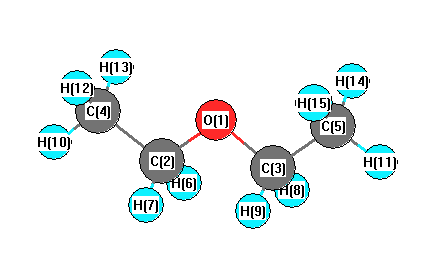
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3133 |
3023 |
30.30 |
|
|
|
| 2 |
A1 |
3066 |
2958 |
4.77 |
|
|
|
| 3 |
A1 |
2989 |
2884 |
85.37 |
|
|
|
| 4 |
A1 |
1601 |
1545 |
0.49 |
|
|
|
| 5 |
A1 |
1572 |
1516 |
5.16 |
|
|
|
| 6 |
A1 |
1462 |
1410 |
2.87 |
|
|
|
| 7 |
A1 |
1459 |
1408 |
6.30 |
|
|
|
| 8 |
A1 |
1176 |
1135 |
11.48 |
|
|
|
| 9 |
A1 |
1022 |
986 |
3.78 |
|
|
|
| 10 |
A1 |
834 |
805 |
6.52 |
|
|
|
| 11 |
A1 |
430 |
415 |
0.17 |
|
|
|
| 12 |
A1 |
184 |
177 |
0.75 |
|
|
|
| 13 |
A2 |
3140 |
3030 |
0.00 |
|
|
|
| 14 |
A2 |
3007 |
2901 |
0.00 |
|
|
|
| 15 |
A2 |
1551 |
1497 |
0.00 |
|
|
|
| 16 |
A2 |
1321 |
1275 |
0.00 |
|
|
|
| 17 |
A2 |
1174 |
1133 |
0.00 |
|
|
|
| 18 |
A2 |
860 |
829 |
0.00 |
|
|
|
| 19 |
A2 |
243 |
234 |
0.00 |
|
|
|
| 20 |
A2 |
108 |
104 |
0.00 |
|
|
|
| 21 |
B1 |
3140 |
3030 |
51.78 |
|
|
|
| 22 |
B1 |
3008 |
2902 |
121.22 |
|
|
|
| 23 |
B1 |
1552 |
1497 |
12.41 |
|
|
|
| 24 |
B1 |
1320 |
1273 |
0.02 |
|
|
|
| 25 |
B1 |
1208 |
1166 |
11.95 |
|
|
|
| 26 |
B1 |
870 |
840 |
2.85 |
|
|
|
| 27 |
B1 |
246 |
237 |
2.05 |
|
|
|
| 28 |
B1 |
97 |
93 |
5.27 |
|
|
|
| 29 |
B2 |
3133 |
3023 |
18.76 |
|
|
|
| 30 |
B2 |
3065 |
2958 |
23.99 |
|
|
|
| 31 |
B2 |
2978 |
2874 |
8.63 |
|
|
|
| 32 |
B2 |
1584 |
1528 |
3.58 |
|
|
|
| 33 |
B2 |
1568 |
1513 |
9.03 |
|
|
|
| 34 |
B2 |
1458 |
1407 |
12.09 |
|
|
|
| 35 |
B2 |
1388 |
1339 |
66.62 |
|
|
|
| 36 |
B2 |
1138 |
1098 |
79.75 |
|
|
|
| 37 |
B2 |
1079 |
1041 |
89.78 |
|
|
|
| 38 |
B2 |
942 |
909 |
3.73 |
|
|
|
| 39 |
B2 |
410 |
396 |
5.66 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30254.9 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 29193.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.487 |
|
|
-0.429 |
| 2 |
C |
-0.119 |
|
|
0.292 |
| 3 |
C |
-0.119 |
|
|
0.292 |
| 4 |
C |
-0.571 |
|
|
-0.373 |
| 5 |
C |
-0.571 |
|
|
-0.373 |
| 6 |
H |
0.171 |
|
|
-0.009 |
| 7 |
H |
0.171 |
|
|
-0.010 |
| 8 |
H |
0.171 |
|
|
-0.009 |
| 9 |
H |
0.171 |
|
|
-0.010 |
| 10 |
H |
0.182 |
|
|
0.092 |
| 11 |
H |
0.182 |
|
|
0.092 |
| 12 |
H |
0.204 |
|
|
0.111 |
| 13 |
H |
0.204 |
|
|
0.110 |
| 14 |
H |
0.204 |
|
|
0.110 |
| 15 |
H |
0.204 |
|
|
0.111 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.231 |
1.231 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.006 |
0.000 |
-1.223 |
1.223 |
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.976 |
0.000 |
0.000 |
| y |
0.000 |
8.344 |
0.000 |
| z |
0.000 |
0.000 |
6.187 |
<r2> (average value of r
2) Å
2
| <r2> |
179.290 |
| (<r2>)1/2 |
13.390 |