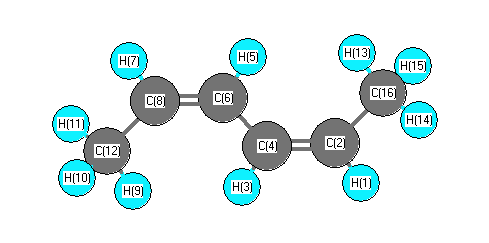
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3182 |
3070 |
60.68 |
|
|
|
| 2 |
A' |
3170 |
3059 |
0.00 |
|
|
|
| 3 |
A' |
3153 |
3043 |
0.00 |
|
|
|
| 4 |
A' |
3148 |
3038 |
26.25 |
|
|
|
| 5 |
A' |
3135 |
3025 |
4.58 |
|
|
|
| 6 |
A' |
3129 |
3019 |
0.00 |
|
|
|
| 7 |
A' |
3036 |
2929 |
0.09 |
|
|
|
| 8 |
A' |
3035 |
2929 |
53.20 |
|
|
|
| 9 |
A' |
1715 |
1655 |
0.00 |
|
|
|
| 10 |
A' |
1677 |
1618 |
7.26 |
|
|
|
| 11 |
A' |
1557 |
1502 |
0.00 |
|
|
|
| 12 |
A' |
1553 |
1499 |
22.03 |
|
|
|
| 13 |
A' |
1506 |
1453 |
0.00 |
|
|
|
| 14 |
A' |
1471 |
1420 |
15.55 |
|
|
|
| 15 |
A' |
1463 |
1412 |
0.00 |
|
|
|
| 16 |
A' |
1435 |
1384 |
3.22 |
|
|
|
| 17 |
A' |
1347 |
1300 |
0.00 |
|
|
|
| 18 |
A' |
1316 |
1270 |
1.39 |
|
|
|
| 19 |
A' |
1192 |
1150 |
0.00 |
|
|
|
| 20 |
A' |
1113 |
1074 |
5.08 |
|
|
|
| 21 |
A' |
1026 |
990 |
0.00 |
|
|
|
| 22 |
A' |
927 |
894 |
33.78 |
|
|
|
| 23 |
A' |
901 |
870 |
0.00 |
|
|
|
| 24 |
A' |
656 |
633 |
0.00 |
|
|
|
| 25 |
A' |
544 |
525 |
17.04 |
|
|
|
| 26 |
A' |
285 |
275 |
0.00 |
|
|
|
| 27 |
A' |
197 |
190 |
0.26 |
|
|
|
| 28 |
A" |
3071 |
2964 |
39.40 |
|
|
|
| 29 |
A" |
3071 |
2964 |
1.62 |
|
|
|
| 30 |
A" |
1554 |
1499 |
0.00 |
|
|
|
| 31 |
A" |
1554 |
1499 |
16.69 |
|
|
|
| 32 |
A" |
1098 |
1060 |
0.00 |
|
|
|
| 33 |
A" |
1097 |
1058 |
0.00 |
|
|
|
| 34 |
A" |
1037 |
1001 |
0.85 |
|
|
|
| 35 |
A" |
1007 |
972 |
0.00 |
|
|
|
| 36 |
A" |
820 |
791 |
0.00 |
|
|
|
| 37 |
A" |
715 |
690 |
82.78 |
|
|
|
| 38 |
A" |
475 |
459 |
0.00 |
|
|
|
| 39 |
A" |
315 |
304 |
0.00 |
|
|
|
| 40 |
A" |
129 |
125 |
0.00 |
|
|
|
| 41 |
A" |
124 |
120 |
0.56 |
|
|
|
| 42 |
A" |
65 |
63 |
0.60 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31500.6 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 30394.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.181 |
|
|
0.096 |
| 2 |
C |
-0.161 |
|
|
-0.086 |
| 3 |
H |
0.181 |
|
|
0.166 |
| 4 |
C |
-0.200 |
|
|
-0.221 |
| 5 |
H |
0.181 |
|
|
0.162 |
| 6 |
C |
-0.200 |
|
|
-0.220 |
| 7 |
H |
0.181 |
|
|
0.100 |
| 8 |
C |
-0.161 |
|
|
-0.099 |
| 9 |
H |
0.201 |
|
|
0.062 |
| 10 |
H |
0.204 |
|
|
0.063 |
| 11 |
H |
0.204 |
|
|
0.063 |
| 12 |
C |
-0.610 |
|
|
-0.134 |
| 13 |
H |
0.201 |
|
|
0.072 |
| 14 |
H |
0.204 |
|
|
0.068 |
| 15 |
H |
0.204 |
|
|
0.068 |
| 16 |
C |
-0.610 |
|
|
-0.159 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.001 |
-0.001 |
0.000 |
0.001 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.233 |
-1.091 |
0.000 |
| y |
-1.091 |
-35.545 |
0.000 |
| z |
0.000 |
0.000 |
-41.329 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.204 |
-1.091 |
0.000 |
| y |
-1.091 |
2.236 |
0.000 |
| z |
0.000 |
0.000 |
-6.440 |
|
| Polar |
|---|
| 3z2-r2 | -12.879 |
|---|
| x2-y2 | 1.312 |
|---|
| xy | -1.091 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.275 |
-4.192 |
0.000 |
| y |
-4.192 |
11.193 |
0.000 |
| z |
0.000 |
0.000 |
4.623 |
<r2> (average value of r
2) Å
2
| <r2> |
246.534 |
| (<r2>)1/2 |
15.701 |