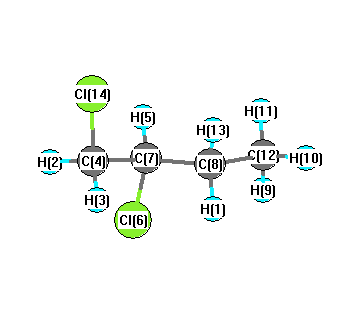
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3233 |
3120 |
0.47 |
|
|
|
| 2 |
A |
3166 |
3055 |
4.46 |
|
|
|
| 3 |
A |
3153 |
3042 |
0.70 |
|
|
|
| 4 |
A |
3141 |
3031 |
11.03 |
|
|
|
| 5 |
A |
3124 |
3014 |
24.30 |
|
|
|
| 6 |
A |
3098 |
2989 |
1.44 |
|
|
|
| 7 |
A |
3061 |
2954 |
12.11 |
|
|
|
| 8 |
A |
3059 |
2952 |
10.87 |
|
|
|
| 9 |
A |
1573 |
1518 |
9.45 |
|
|
|
| 10 |
A |
1564 |
1510 |
9.90 |
|
|
|
| 11 |
A |
1537 |
1483 |
4.12 |
|
|
|
| 12 |
A |
1527 |
1473 |
13.36 |
|
|
|
| 13 |
A |
1467 |
1416 |
9.35 |
|
|
|
| 14 |
A |
1418 |
1369 |
0.45 |
|
|
|
| 15 |
A |
1375 |
1327 |
3.34 |
|
|
|
| 16 |
A |
1353 |
1305 |
3.22 |
|
|
|
| 17 |
A |
1296 |
1251 |
8.33 |
|
|
|
| 18 |
A |
1256 |
1212 |
4.58 |
|
|
|
| 19 |
A |
1204 |
1162 |
20.05 |
|
|
|
| 20 |
A |
1134 |
1094 |
2.29 |
|
|
|
| 21 |
A |
1107 |
1069 |
3.53 |
|
|
|
| 22 |
A |
1084 |
1046 |
1.53 |
|
|
|
| 23 |
A |
1017 |
981 |
0.17 |
|
|
|
| 24 |
A |
958 |
925 |
12.48 |
|
|
|
| 25 |
A |
838 |
809 |
9.57 |
|
|
|
| 26 |
A |
824 |
795 |
16.55 |
|
|
|
| 27 |
A |
662 |
639 |
28.99 |
|
|
|
| 28 |
A |
612 |
590 |
77.21 |
|
|
|
| 29 |
A |
468 |
452 |
2.31 |
|
|
|
| 30 |
A |
369 |
356 |
2.59 |
|
|
|
| 31 |
A |
282 |
272 |
0.24 |
|
|
|
| 32 |
A |
244 |
236 |
0.31 |
|
|
|
| 33 |
A |
215 |
207 |
3.69 |
|
|
|
| 34 |
A |
187 |
180 |
6.05 |
|
|
|
| 35 |
A |
112 |
108 |
4.96 |
|
|
|
| 36 |
A |
103 |
100 |
2.66 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25411.5 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 24519.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.216 |
|
|
0.090 |
| 2 |
H |
0.279 |
|
|
0.245 |
| 3 |
H |
0.271 |
|
|
0.234 |
| 4 |
C |
-0.487 |
|
|
-0.392 |
| 5 |
H |
0.273 |
|
|
0.198 |
| 6 |
Cl |
-0.057 |
|
|
-0.209 |
| 7 |
C |
-0.352 |
|
|
-0.040 |
| 8 |
C |
-0.377 |
|
|
-0.001 |
| 9 |
H |
0.220 |
|
|
0.109 |
| 10 |
H |
0.200 |
|
|
0.112 |
| 11 |
H |
0.193 |
|
|
0.104 |
| 12 |
C |
-0.558 |
|
|
-0.358 |
| 13 |
H |
0.240 |
|
|
0.078 |
| 14 |
Cl |
-0.058 |
|
|
-0.170 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.358 |
0.765 |
0.262 |
0.884 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.319 |
0.739 |
0.232 |
0.838 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-60.835 |
4.588 |
-0.445 |
| y |
4.588 |
-51.144 |
-0.184 |
| z |
-0.445 |
-0.184 |
-50.620 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-9.953 |
4.588 |
-0.445 |
| y |
4.588 |
4.583 |
-0.184 |
| z |
-0.445 |
-0.184 |
5.369 |
|
| Polar |
|---|
| 3z2-r2 | 10.738 |
|---|
| x2-y2 | -9.691 |
|---|
| xy | 4.588 |
|---|
| xz | -0.445 |
|---|
| yz | -0.184 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.427 |
-1.797 |
0.373 |
| y |
-1.797 |
8.761 |
-0.220 |
| z |
0.373 |
-0.220 |
6.159 |
<r2> (average value of r
2) Å
2
| <r2> |
334.853 |
| (<r2>)1/2 |
18.299 |