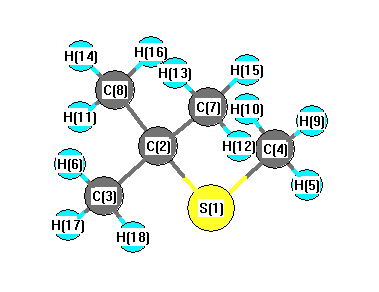
Vibrational Frequencies calculated at BLYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3094 |
3077 |
6.36 |
|
|
|
| 2 |
A |
3054 |
3037 |
26.94 |
|
|
|
| 3 |
A |
3033 |
3017 |
28.98 |
|
|
|
| 4 |
A |
3025 |
3008 |
30.55 |
|
|
|
| 5 |
A |
3003 |
2987 |
16.77 |
|
|
|
| 6 |
A |
2973 |
2957 |
31.95 |
|
|
|
| 7 |
A |
2964 |
2948 |
26.14 |
|
|
|
| 8 |
A |
1545 |
1537 |
9.76 |
|
|
|
| 9 |
A |
1529 |
1521 |
9.86 |
|
|
|
| 10 |
A |
1517 |
1509 |
1.98 |
|
|
|
| 11 |
A |
1504 |
1495 |
11.02 |
|
|
|
| 12 |
A |
1436 |
1429 |
2.39 |
|
|
|
| 13 |
A |
1412 |
1405 |
12.05 |
|
|
|
| 14 |
A |
1339 |
1332 |
2.79 |
|
|
|
| 15 |
A |
1226 |
1219 |
4.56 |
|
|
|
| 16 |
A |
1186 |
1179 |
64.40 |
|
|
|
| 17 |
A |
1047 |
1041 |
2.08 |
|
|
|
| 18 |
A |
962 |
957 |
11.40 |
|
|
|
| 19 |
A |
921 |
916 |
1.00 |
|
|
|
| 20 |
A |
771 |
767 |
1.51 |
|
|
|
| 21 |
A |
616 |
612 |
2.01 |
|
|
|
| 22 |
A |
526 |
523 |
6.32 |
|
|
|
| 23 |
A |
403 |
401 |
0.00 |
|
|
|
| 24 |
A |
343 |
342 |
1.64 |
|
|
|
| 25 |
A |
303 |
301 |
0.08 |
|
|
|
| 26 |
A |
276 |
275 |
0.01 |
|
|
|
| 27 |
A |
201 |
200 |
0.41 |
|
|
|
| 28 |
A |
3090 |
3073 |
15.32 |
|
|
|
| 29 |
A |
3053 |
3036 |
29.62 |
|
|
|
| 30 |
A |
3048 |
3031 |
1.54 |
|
|
|
| 31 |
A |
3020 |
3004 |
3.68 |
|
|
|
| 32 |
A |
2960 |
2943 |
13.09 |
|
|
|
| 33 |
A |
1533 |
1524 |
3.40 |
|
|
|
| 34 |
A |
1517 |
1509 |
1.49 |
|
|
|
| 35 |
A |
1508 |
1499 |
3.58 |
|
|
|
| 36 |
A |
1497 |
1488 |
7.12 |
|
|
|
| 37 |
A |
1412 |
1404 |
10.44 |
|
|
|
| 38 |
A |
1225 |
1218 |
9.35 |
|
|
|
| 39 |
A |
1043 |
1037 |
0.01 |
|
|
|
| 40 |
A |
986 |
981 |
0.10 |
|
|
|
| 41 |
A |
939 |
934 |
5.55 |
|
|
|
| 42 |
A |
916 |
911 |
0.20 |
|
|
|
| 43 |
A |
389 |
387 |
0.21 |
|
|
|
| 44 |
A |
289 |
288 |
0.95 |
|
|
|
| 45 |
A |
272 |
271 |
0.10 |
|
|
|
| 46 |
A |
238 |
237 |
0.00 |
|
|
|
| 47 |
A |
128 |
127 |
1.65 |
|
|
|
| 48 |
A |
38 |
38 |
1.44 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34654.5 cm
-1
Scaled (by 0.9945) Zero Point Vibrational Energy (zpe) 34463.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.198 |
|
|
|
| 2 |
C |
-0.293 |
|
|
|
| 3 |
C |
-0.485 |
|
|
|
| 4 |
C |
-0.751 |
|
|
|
| 5 |
H |
0.218 |
|
|
|
| 6 |
H |
0.180 |
|
|
|
| 7 |
C |
-0.482 |
|
|
|
| 8 |
C |
-0.482 |
|
|
|
| 9 |
H |
0.209 |
|
|
|
| 10 |
H |
0.209 |
|
|
|
| 11 |
H |
0.192 |
|
|
|
| 12 |
H |
0.192 |
|
|
|
| 13 |
H |
0.177 |
|
|
|
| 14 |
H |
0.177 |
|
|
|
| 15 |
H |
0.180 |
|
|
|
| 16 |
H |
0.180 |
|
|
|
| 17 |
H |
0.192 |
|
|
|
| 18 |
H |
0.192 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.720 |
0.882 |
0.000 |
1.933 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.331 |
-2.905 |
0.000 |
| y |
-2.905 |
-44.760 |
0.000 |
| z |
0.000 |
0.000 |
-48.182 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.860 |
-2.905 |
0.000 |
| y |
-2.905 |
3.497 |
0.000 |
| z |
0.000 |
0.000 |
-1.637 |
|
| Polar |
|---|
| 3z2-r2 | -3.273 |
|---|
| x2-y2 | -3.571 |
|---|
| xy | -2.905 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
237.153 |
| (<r2>)1/2 |
15.400 |