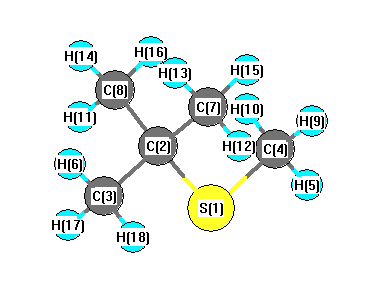
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3174 |
3063 |
4.64 |
|
|
|
| 2 |
A |
3133 |
3023 |
24.49 |
|
|
|
| 3 |
A |
3114 |
3005 |
27.73 |
|
|
|
| 4 |
A |
3106 |
2997 |
27.66 |
|
|
|
| 5 |
A |
3082 |
2974 |
15.02 |
|
|
|
| 6 |
A |
3050 |
2943 |
28.43 |
|
|
|
| 7 |
A |
3043 |
2936 |
20.65 |
|
|
|
| 8 |
A |
1581 |
1525 |
12.09 |
|
|
|
| 9 |
A |
1566 |
1511 |
12.61 |
|
|
|
| 10 |
A |
1552 |
1498 |
2.24 |
|
|
|
| 11 |
A |
1540 |
1486 |
11.00 |
|
|
|
| 12 |
A |
1477 |
1425 |
3.86 |
|
|
|
| 13 |
A |
1452 |
1401 |
16.39 |
|
|
|
| 14 |
A |
1385 |
1337 |
1.65 |
|
|
|
| 15 |
A |
1268 |
1224 |
3.50 |
|
|
|
| 16 |
A |
1227 |
1183 |
63.33 |
|
|
|
| 17 |
A |
1080 |
1042 |
2.58 |
|
|
|
| 18 |
A |
996 |
961 |
12.23 |
|
|
|
| 19 |
A |
956 |
922 |
1.31 |
|
|
|
| 20 |
A |
810 |
781 |
1.34 |
|
|
|
| 21 |
A |
653 |
630 |
1.94 |
|
|
|
| 22 |
A |
557 |
537 |
6.16 |
|
|
|
| 23 |
A |
417 |
402 |
0.02 |
|
|
|
| 24 |
A |
359 |
347 |
1.33 |
|
|
|
| 25 |
A |
316 |
305 |
0.07 |
|
|
|
| 26 |
A |
285 |
275 |
0.00 |
|
|
|
| 27 |
A |
211 |
204 |
0.46 |
|
|
|
| 28 |
A |
3173 |
3062 |
12.00 |
|
|
|
| 29 |
A |
3132 |
3022 |
25.15 |
|
|
|
| 30 |
A |
3128 |
3018 |
1.49 |
|
|
|
| 31 |
A |
3101 |
2992 |
2.74 |
|
|
|
| 32 |
A |
3039 |
2932 |
12.16 |
|
|
|
| 33 |
A |
1568 |
1513 |
3.63 |
|
|
|
| 34 |
A |
1552 |
1498 |
2.84 |
|
|
|
| 35 |
A |
1544 |
1489 |
5.15 |
|
|
|
| 36 |
A |
1533 |
1480 |
7.79 |
|
|
|
| 37 |
A |
1452 |
1401 |
14.51 |
|
|
|
| 38 |
A |
1267 |
1223 |
8.49 |
|
|
|
| 39 |
A |
1075 |
1037 |
0.02 |
|
|
|
| 40 |
A |
1014 |
979 |
0.14 |
|
|
|
| 41 |
A |
973 |
939 |
5.27 |
|
|
|
| 42 |
A |
952 |
918 |
0.51 |
|
|
|
| 43 |
A |
401 |
387 |
0.20 |
|
|
|
| 44 |
A |
303 |
292 |
0.93 |
|
|
|
| 45 |
A |
281 |
271 |
0.10 |
|
|
|
| 46 |
A |
247 |
238 |
0.01 |
|
|
|
| 47 |
A |
140 |
135 |
1.66 |
|
|
|
| 48 |
A |
41 |
39 |
1.60 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35652.8 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 34401.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.234 |
|
|
-0.306 |
| 2 |
C |
-0.350 |
|
|
0.416 |
| 3 |
C |
-0.512 |
|
|
-0.511 |
| 4 |
C |
-0.802 |
|
|
-0.332 |
| 5 |
H |
0.231 |
|
|
0.184 |
| 6 |
H |
0.192 |
|
|
0.146 |
| 7 |
C |
-0.508 |
|
|
-0.453 |
| 8 |
C |
-0.508 |
|
|
-0.453 |
| 9 |
H |
0.221 |
|
|
0.116 |
| 10 |
H |
0.221 |
|
|
0.116 |
| 11 |
H |
0.205 |
|
|
0.128 |
| 12 |
H |
0.205 |
|
|
0.128 |
| 13 |
H |
0.190 |
|
|
0.119 |
| 14 |
H |
0.190 |
|
|
0.119 |
| 15 |
H |
0.192 |
|
|
0.141 |
| 16 |
H |
0.192 |
|
|
0.141 |
| 17 |
H |
0.204 |
|
|
0.151 |
| 18 |
H |
0.204 |
|
|
0.151 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.766 |
0.879 |
0.000 |
1.973 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
1.787 |
0.853 |
0.000 |
1.980 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.437 |
-2.913 |
0.000 |
| y |
-2.913 |
-44.713 |
0.000 |
| z |
0.000 |
0.000 |
-48.129 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.016 |
-2.913 |
0.000 |
| y |
-2.913 |
3.570 |
0.000 |
| z |
0.000 |
0.000 |
-1.554 |
|
| Polar |
|---|
| 3z2-r2 | -3.108 |
|---|
| x2-y2 | -3.724 |
|---|
| xy | -2.913 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.500 |
-0.329 |
0.000 |
| y |
-0.329 |
12.134 |
0.000 |
| z |
0.000 |
0.000 |
8.228 |
<r2> (average value of r
2) Å
2
| <r2> |
232.900 |
| (<r2>)1/2 |
15.261 |