Jump to
S1C2
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -276.764821 |
| Energy at 298.15K | |
| HF Energy | -276.764821 |
| Nuclear repulsion energy | 123.972552 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3065 |
2957 |
0.00 |
179.61 |
0.10 |
0.18 |
| 2 |
Ag |
1583 |
1527 |
0.00 |
30.76 |
0.72 |
0.84 |
| 3 |
Ag |
1473 |
1422 |
0.00 |
5.25 |
0.75 |
0.86 |
| 4 |
Ag |
1058 |
1021 |
0.00 |
10.20 |
0.73 |
0.85 |
| 5 |
Ag |
1051 |
1014 |
0.00 |
6.21 |
0.18 |
0.31 |
| 6 |
Ag |
435 |
420 |
0.00 |
3.33 |
0.54 |
0.70 |
| 7 |
Au |
3126 |
3017 |
74.71 |
0.00 |
0.00 |
0.00 |
| 8 |
Au |
1231 |
1187 |
0.58 |
0.00 |
0.00 |
0.00 |
| 9 |
Au |
861 |
830 |
0.04 |
0.00 |
0.00 |
0.00 |
| 10 |
Au |
117 |
113 |
13.20 |
0.00 |
0.00 |
0.00 |
| 11 |
Bg |
3097 |
2988 |
0.00 |
122.12 |
0.75 |
0.86 |
| 12 |
Bg |
1348 |
1301 |
0.00 |
19.55 |
0.75 |
0.86 |
| 13 |
Bg |
1179 |
1138 |
0.00 |
12.18 |
0.75 |
0.86 |
| 14 |
Bu |
3074 |
2966 |
56.96 |
0.00 |
0.00 |
0.00 |
| 15 |
Bu |
1600 |
1543 |
1.51 |
0.00 |
0.00 |
0.00 |
| 16 |
Bu |
1398 |
1349 |
10.06 |
0.00 |
0.00 |
0.00 |
| 17 |
Bu |
1085 |
1047 |
99.28 |
0.00 |
0.00 |
0.00 |
| 18 |
Bu |
232 |
224 |
19.04 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 13505.3 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 13031.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.434 |
0.627 |
0.000 |
| C2 |
-0.434 |
-0.627 |
0.000 |
| F3 |
-0.434 |
1.751 |
0.000 |
| F4 |
0.434 |
-1.751 |
0.000 |
| H5 |
1.071 |
0.635 |
0.890 |
| H6 |
1.071 |
0.635 |
-0.890 |
| H7 |
-1.071 |
-0.635 |
0.890 |
| H8 |
-1.071 |
-0.635 |
-0.890 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5243 | 1.4202 | 2.3773 | 1.0941 | 1.0941 | 2.1558 | 2.1558 |
C2 | 1.5243 | | 2.3773 | 1.4202 | 2.1558 | 2.1558 | 1.0941 | 1.0941 | F3 | 1.4202 | 2.3773 | | 3.6074 | 2.0735 | 2.0735 | 2.6249 | 2.6249 | F4 | 2.3773 | 1.4202 | 3.6074 | | 2.6249 | 2.6249 | 2.0735 | 2.0735 | H5 | 1.0941 | 2.1558 | 2.0735 | 2.6249 | | 1.7794 | 2.4898 | 3.0603 | H6 | 1.0941 | 2.1558 | 2.0735 | 2.6249 | 1.7794 | | 3.0603 | 2.4898 | H7 | 2.1558 | 1.0941 | 2.6249 | 2.0735 | 2.4898 | 3.0603 | | 1.7794 | H8 | 2.1558 | 1.0941 | 2.6249 | 2.0735 | 3.0603 | 2.4898 | 1.7794 | |
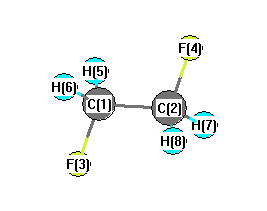 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
107.627 |
|
C1 |
C2 |
H7 |
109.747 |
| C1 |
C2 |
H8 |
109.747 |
|
C2 |
C1 |
F3 |
107.627 |
| C2 |
C1 |
H5 |
109.747 |
|
C2 |
C1 |
H6 |
109.747 |
| F3 |
C1 |
H5 |
110.443 |
|
F3 |
C1 |
H6 |
110.443 |
| F4 |
C2 |
H7 |
110.443 |
|
F4 |
C2 |
H8 |
110.443 |
| H5 |
C1 |
H6 |
108.823 |
|
H7 |
C2 |
H8 |
108.823 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.108 |
|
|
|
| 2 |
C |
-0.108 |
|
|
|
| 3 |
F |
-0.297 |
|
|
|
| 4 |
F |
-0.297 |
|
|
|
| 5 |
H |
0.203 |
|
|
|
| 6 |
H |
0.203 |
|
|
|
| 7 |
H |
0.203 |
|
|
|
| 8 |
H |
0.203 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.519 |
2.725 |
0.000 |
| y |
2.725 |
-28.829 |
0.000 |
| z |
0.000 |
0.000 |
-21.315 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.553 |
2.725 |
0.000 |
| y |
2.725 |
-7.412 |
0.000 |
| z |
0.000 |
0.000 |
3.859 |
|
| Polar |
|---|
| 3z2-r2 | 7.718 |
|---|
| x2-y2 | 7.310 |
|---|
| xy | 2.725 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.194 |
-0.096 |
0.000 |
| y |
-0.096 |
3.133 |
0.000 |
| z |
0.000 |
0.000 |
2.983 |
<r2> (average value of r
2) Å
2
| <r2> |
89.816 |
| (<r2>)1/2 |
9.477 |
Jump to
S1C1
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -276.765126 |
| Energy at 298.15K | |
| HF Energy | -276.765126 |
| Nuclear repulsion energy | 125.852513 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3092 |
2984 |
26.34 |
|
|
|
| 2 |
A |
3059 |
2952 |
30.88 |
|
|
|
| 3 |
A |
1563 |
1508 |
0.11 |
|
|
|
| 4 |
A |
1458 |
1406 |
12.13 |
|
|
|
| 5 |
A |
1335 |
1288 |
0.00 |
|
|
|
| 6 |
A |
1134 |
1095 |
3.53 |
|
|
|
| 7 |
A |
1102 |
1063 |
41.73 |
|
|
|
| 8 |
A |
872 |
841 |
14.52 |
|
|
|
| 9 |
A |
308 |
297 |
0.59 |
|
|
|
| 10 |
A |
145 |
140 |
3.74 |
|
|
|
| 11 |
B |
3107 |
2998 |
51.90 |
|
|
|
| 12 |
B |
3045 |
2938 |
16.09 |
|
|
|
| 13 |
B |
1564 |
1509 |
6.01 |
|
|
|
| 14 |
B |
1469 |
1418 |
8.70 |
|
|
|
| 15 |
B |
1273 |
1228 |
1.23 |
|
|
|
| 16 |
B |
1088 |
1050 |
19.92 |
|
|
|
| 17 |
B |
931 |
898 |
25.82 |
|
|
|
| 18 |
B |
455 |
439 |
15.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13499.1 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 13025.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.281 |
0.703 |
0.516 |
| C2 |
-0.281 |
-0.703 |
0.516 |
| F3 |
-0.281 |
1.451 |
-0.552 |
| F4 |
0.281 |
-1.451 |
-0.552 |
| H5 |
0.041 |
1.186 |
1.470 |
| H6 |
1.369 |
0.653 |
0.401 |
| H7 |
-0.041 |
-1.186 |
1.470 |
| H8 |
-1.369 |
-0.653 |
0.401 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5151 | 1.4193 | 2.4046 | 1.0963 | 1.0949 | 2.1415 | 2.1389 |
C2 | 1.5151 | | 2.4046 | 1.4193 | 2.1415 | 2.1389 | 1.0963 | 1.0949 | F3 | 1.4193 | 2.4046 | | 2.9563 | 2.0645 | 2.0657 | 3.3321 | 2.5533 | F4 | 2.4046 | 1.4193 | 2.9563 | | 3.3321 | 2.5533 | 2.0645 | 2.0657 | H5 | 1.0963 | 2.1415 | 2.0645 | 3.3321 | | 1.7860 | 2.3741 | 2.5520 | H6 | 1.0949 | 2.1389 | 2.0657 | 2.5533 | 1.7860 | | 2.5520 | 3.0330 | H7 | 2.1415 | 1.0963 | 3.3321 | 2.0645 | 2.3741 | 2.5520 | | 1.7860 | H8 | 2.1389 | 1.0949 | 2.5533 | 2.0657 | 2.5520 | 3.0330 | 1.7860 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.021 |
|
C1 |
C2 |
H7 |
109.137 |
| C1 |
C2 |
H8 |
109.009 |
|
C2 |
C1 |
F3 |
110.021 |
| C2 |
C1 |
H5 |
109.137 |
|
C2 |
C1 |
H6 |
109.009 |
| F3 |
C1 |
H5 |
109.637 |
|
F3 |
C1 |
H6 |
109.821 |
| F4 |
C2 |
H7 |
109.637 |
|
F4 |
C2 |
H8 |
109.821 |
| H5 |
C1 |
H6 |
109.196 |
|
H7 |
C2 |
H8 |
109.196 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.098 |
|
|
|
| 2 |
C |
-0.098 |
|
|
|
| 3 |
F |
-0.294 |
|
|
|
| 4 |
F |
-0.294 |
|
|
|
| 5 |
H |
0.185 |
|
|
|
| 6 |
H |
0.208 |
|
|
|
| 7 |
H |
0.185 |
|
|
|
| 8 |
H |
0.208 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.617 |
2.617 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.355 |
1.569 |
0.000 |
| y |
1.569 |
-25.997 |
0.000 |
| z |
0.000 |
0.000 |
-21.908 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.597 |
1.569 |
0.000 |
| y |
1.569 |
-4.365 |
0.000 |
| z |
0.000 |
0.000 |
1.768 |
|
| Polar |
|---|
| 3z2-r2 | 3.537 |
|---|
| x2-y2 | 4.642 |
|---|
| xy | 1.569 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.068 |
-0.023 |
0.000 |
| y |
-0.023 |
3.172 |
0.000 |
| z |
0.000 |
0.000 |
3.211 |
<r2> (average value of r
2) Å
2
| <r2> |
81.369 |
| (<r2>)1/2 |
9.020 |
